Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -2583.222587 |
| Energy at 298.15K | -2583.226700 |
| HF Energy | -2583.174741 |
| Nuclear repulsion energy | 79.736875 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3438 |
3056 |
0.50 |
|
|
|
| 2 |
A' |
1604 |
1426 |
34.05 |
|
|
|
| 3 |
A' |
820 |
729 |
16.33 |
|
|
|
| 4 |
A' |
643 |
572 |
11.86 |
|
|
|
| 5 |
A" |
3616 |
3214 |
0.66 |
|
|
|
| 6 |
A" |
1051 |
934 |
1.99 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5585.9 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 4965.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.013 |
1.503 |
0.000 |
| Br2 |
-0.013 |
-0.372 |
0.000 |
| H3 |
0.262 |
2.006 |
0.938 |
| H4 |
0.262 |
2.006 |
-0.938 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
H4 |
| C1 | | 1.8749 | 1.0993 | 1.0993 |
Br2 | 1.8749 | | 2.5713 | 2.5713 | H3 | 1.0993 | 2.5713 | | 1.8755 | H4 | 1.0993 | 2.5713 | 1.8755 | |
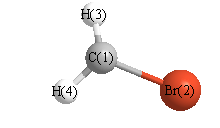 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.261 |
|
Br2 |
C1 |
H4 |
117.261 |
| H3 |
C1 |
H4 |
117.082 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -2583.221618 |
| Energy at 298.15K | |
| HF Energy | -2583.174307 |
| Nuclear repulsion energy | 79.878631 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3466 |
3081 |
0.04 |
37.11 |
0.12 |
0.22 |
| 2 |
A1 |
1590 |
1414 |
43.22 |
8.71 |
0.75 |
0.86 |
| 3 |
A1 |
824 |
732 |
14.80 |
6.90 |
0.34 |
0.51 |
| 4 |
B1 |
499i |
443i |
29.90 |
0.94 |
0.75 |
0.86 |
| 5 |
B2 |
3652 |
3246 |
0.02 |
26.31 |
0.75 |
0.86 |
| 6 |
B2 |
1015 |
903 |
1.87 |
4.86 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 5024.1 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 4466.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.494 |
| Br2 |
0.000 |
0.000 |
0.373 |
| H3 |
0.000 |
0.950 |
-2.040 |
| H4 |
0.000 |
-0.950 |
-2.040 |
Atom - Atom Distances (Å)
| |
C1 |
Br2 |
H3 |
H4 |
| C1 | | 1.8672 | 1.0953 | 1.0953 |
Br2 | 1.8672 | | 2.5928 | 2.5928 | H3 | 1.0953 | 2.5928 | | 1.8998 | H4 | 1.0953 | 2.5928 | 1.8998 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
119.858 |
|
Br2 |
C1 |
H4 |
119.858 |
| H3 |
C1 |
H4 |
120.283 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
