return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (Furan, 2,5-dihydro-)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-227.188740
Energy at 298.15K-227.195702
HF Energy-226.921817
Nuclear repulsion energy169.958295
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3561 3166 0.36      
2 A' 3434 3052 17.94      
3 A' 3329 2959 3.61      
4 A' 1779 1582 0.12      
5 A' 1726 1535 0.08      
6 A' 1519 1350 4.09      
7 A' 1284 1141 0.88      
8 A' 1192 1060 0.02      
9 A' 1102 980 9.86      
10 A' 1076 957 2.65      
11 A' 991 881 0.11      
12 A' 778 692 2.23      
13 A' 721 641 10.18      
14 A' 83 74 5.64      
15 A" 3532 3139 8.55      
16 A" 3431 3050 0.46      
17 A" 3325 2956 25.02      
18 A" 1718 1527 8.63      
19 A" 1490 1324 0.32      
20 A" 1430 1271 0.56      
21 A" 1306 1161 0.13      
22 A" 1279 1137 19.40      
23 A" 1126 1001 0.06      
24 A" 1015 902 0.01      
25 A" 1013 901 0.14      
26 A" 850 756 0.08      
27 A" 400 355 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 22245.3 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 19773.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.26098 0.24484 0.13331

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.156 -1.242 0.000
C2 -0.031 -0.377 1.181
C3 -0.031 1.080 0.673
C4 -0.031 1.080 -0.673
C5 -0.031 -0.377 -1.181
H6 0.793 -0.576 1.905
H7 -0.993 -0.622 1.697
H8 -0.993 -0.622 -1.697
H9 0.793 -0.576 -1.905
H10 -0.046 1.949 1.345
H11 -0.046 1.949 -1.345

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11
O11.47522.42482.42481.47522.11632.14092.14092.11633.46833.4683
C21.47521.54362.35822.36151.11531.11843.04433.20032.33223.4337
C32.42481.54361.34602.35822.22312.20733.07273.17351.09822.1968
C42.42482.35821.34601.54363.17353.07272.20732.22312.19681.0982
C51.47522.36152.35821.54363.20033.04431.11841.11533.43372.3322
H62.11631.11532.22313.17353.20031.79804.02113.81052.71924.2003
H72.14091.11842.20733.07273.04431.79803.39494.02112.76284.0943
H82.14093.04433.07272.20731.11844.02113.39491.79804.09432.7628
H92.11633.20033.17352.22311.11533.81054.02111.79804.20032.7192
H103.46832.33221.09822.19683.43372.71922.76284.09434.20032.6894
H113.46833.43372.19681.09822.33224.20034.09432.76282.71922.6894

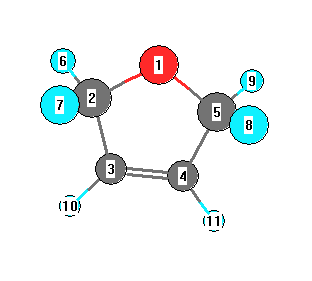
picture of Furan, 2,5-dihydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 106.859 O1 C2 H6 108.763
O1 C2 H7 110.516 O1 C5 C4 106.859
O1 C5 H8 110.516 O1 C5 H9 108.763
C2 O1 C5 106.333 C2 C3 C4 109.204
C2 C3 H10 123.079 C3 C2 H6 112.467
C3 C2 H7 111.028 C3 C4 C5 109.204
C3 C4 H11 127.710 C4 C3 H10 127.710
C4 C5 H8 111.028 C4 C5 H9 112.467
C5 C4 H11 123.079 H6 C2 H7 107.213
H8 C5 H9 107.213
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability