return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2NCH3 (N-methylmethanimine)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-131.587198
Energy at 298.15K-131.592528
HF Energy-131.400984
Nuclear repulsion energy69.927126
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3544 3150 4.97      
2 A' 3522 3131 4.21      
3 A' 3374 2999 0.94      
4 A' 3341 2970 3.82      
5 A' 1726 1534 12.40      
6 A' 1706 1517 3.23      
7 A' 1601 1423 8.09      
8 A' 1556 1383 0.31      
9 A' 1307 1162 2.35      
10 A' 1178 1047 2.24      
11 A' 1012 899 1.43      
12 A' 517 459 1.79      
13 A" 3537 3144 0.72      
14 A" 1704 1514 1.50      
15 A" 1213 1078 0.10      
16 A" 1035 920 10.72      
17 A" 755 671 0.26      
18 A" 162 144 4.28      

Unscaled Zero Point Vibrational Energy (zpe) 16393.5 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 14572.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
1.54482 0.34349 0.29704

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.100 -0.436 0.000
N2 0.000 0.624 0.000
C3 1.211 0.087 0.000
H4 -0.716 -1.477 0.000
H5 -1.731 -0.282 0.896
H6 -1.731 -0.282 -0.896
H7 1.435 -0.999 0.000
H8 2.080 0.770 0.000

Atom - Atom Distances (Å)
  C1 N2 C3 H4 H5 H6 H7 H8
C11.52772.36921.10901.10691.10692.59673.4007
N21.52771.32402.21882.14952.14952.16612.0848
C32.36921.32402.48113.09723.09721.10921.1049
H41.10902.21882.48111.80581.80582.20303.5859
H51.10692.14953.09721.80581.79253.36764.0535
H61.10692.14953.09721.80581.79253.36764.0535
H72.59672.16611.10922.20303.36763.36761.8822
H83.40072.08481.10493.58594.05354.05351.8822

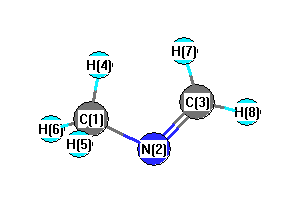
picture of N-methylmethanimine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 C3 112.169 N2 C1 H4 113.653
N2 C1 H5 108.289 N2 C1 H6 108.289
N2 C3 H7 125.569 N2 C3 H8 117.981
H4 C1 H5 109.166 H4 C1 H6 109.166
H5 C1 H6 108.136 H7 C3 H8 116.450
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability