Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Geometric Data calculated at MP2=FULL/STO-3G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -231.755479 |
| Energy at 298.15K | -231.768643 |
| Nuclear repulsion energy | 250.276043 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3549 |
3155 |
4.30 |
|
|
|
| 2 |
A' |
3547 |
3153 |
4.28 |
|
|
|
| 3 |
A' |
3542 |
3148 |
5.40 |
|
|
|
| 4 |
A' |
3533 |
3140 |
1.65 |
|
|
|
| 5 |
A' |
3417 |
3037 |
3.15 |
|
|
|
| 6 |
A' |
3406 |
3028 |
0.81 |
|
|
|
| 7 |
A' |
3365 |
2991 |
4.26 |
|
|
|
| 8 |
A' |
3364 |
2990 |
3.33 |
|
|
|
| 9 |
A' |
1737 |
1544 |
0.58 |
|
|
|
| 10 |
A' |
1732 |
1540 |
0.45 |
|
|
|
| 11 |
A' |
1716 |
1525 |
0.06 |
|
|
|
| 12 |
A' |
1696 |
1508 |
0.49 |
|
|
|
| 13 |
A' |
1621 |
1441 |
1.16 |
|
|
|
| 14 |
A' |
1613 |
1434 |
0.05 |
|
|
|
| 15 |
A' |
1511 |
1343 |
0.49 |
|
|
|
| 16 |
A' |
1465 |
1302 |
7.46 |
|
|
|
| 17 |
A' |
1385 |
1231 |
13.21 |
|
|
|
| 18 |
A' |
1345 |
1195 |
0.87 |
|
|
|
| 19 |
A' |
1121 |
996 |
1.12 |
|
|
|
| 20 |
A' |
1093 |
972 |
0.39 |
|
|
|
| 21 |
A' |
1071 |
952 |
0.14 |
|
|
|
| 22 |
A' |
1039 |
923 |
0.12 |
|
|
|
| 23 |
A' |
940 |
836 |
0.16 |
|
|
|
| 24 |
A' |
790 |
702 |
1.00 |
|
|
|
| 25 |
A' |
671 |
596 |
0.57 |
|
|
|
| 26 |
A' |
404 |
359 |
0.09 |
|
|
|
| 27 |
A' |
355 |
316 |
0.01 |
|
|
|
| 28 |
A' |
91 |
81 |
0.00 |
|
|
|
| 29 |
A" |
3545 |
3151 |
5.04 |
|
|
|
| 30 |
A" |
3544 |
3150 |
1.16 |
|
|
|
| 31 |
A" |
3533 |
3141 |
0.43 |
|
|
|
| 32 |
A" |
3406 |
3028 |
5.98 |
|
|
|
| 33 |
A" |
1740 |
1546 |
1.15 |
|
|
|
| 34 |
A" |
1731 |
1539 |
0.00 |
|
|
|
| 35 |
A" |
1695 |
1507 |
0.54 |
|
|
|
| 36 |
A" |
1459 |
1297 |
2.78 |
|
|
|
| 37 |
A" |
1416 |
1259 |
0.38 |
|
|
|
| 38 |
A" |
1372 |
1220 |
1.76 |
|
|
|
| 39 |
A" |
1360 |
1209 |
2.24 |
|
|
|
| 40 |
A" |
1211 |
1076 |
0.00 |
|
|
|
| 41 |
A" |
1113 |
990 |
0.00 |
|
|
|
| 42 |
A" |
1086 |
965 |
0.04 |
|
|
|
| 43 |
A" |
1005 |
893 |
0.46 |
|
|
|
| 44 |
A" |
878 |
781 |
0.28 |
|
|
|
| 45 |
A" |
406 |
361 |
0.06 |
|
|
|
| 46 |
A" |
317 |
282 |
0.01 |
|
|
|
| 47 |
A" |
252 |
224 |
0.02 |
|
|
|
| 48 |
A" |
222 |
198 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41204.0 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 36626.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.065 |
0.413 |
0.000 |
| C2 |
0.238 |
-1.796 |
0.000 |
| C3 |
-0.065 |
-0.722 |
1.109 |
| C4 |
-0.065 |
-0.722 |
-1.109 |
| C5 |
1.257 |
1.246 |
0.000 |
| C6 |
-1.309 |
1.354 |
0.000 |
| H7 |
1.293 |
-2.124 |
0.000 |
| H8 |
-0.414 |
-2.688 |
0.000 |
| H9 |
0.694 |
-0.618 |
1.906 |
| H10 |
0.694 |
-0.618 |
-1.906 |
| H11 |
-1.058 |
-0.860 |
1.574 |
| H12 |
-1.058 |
-0.860 |
-1.574 |
| H13 |
2.138 |
0.582 |
0.000 |
| H14 |
-2.244 |
0.768 |
0.000 |
| H15 |
1.310 |
1.891 |
-0.895 |
| H16 |
1.310 |
1.891 |
0.895 |
| H17 |
-1.307 |
2.002 |
0.894 |
| H18 |
-1.307 |
2.002 |
-0.894 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.2294 | 1.5874 | 1.5874 | 1.5624 | 1.5603 | 2.8770 | 3.1201 | 2.2961 | 2.2961 | 2.2548 | 2.2548 | 2.2099 | 2.2078 | 2.2080 | 2.2080 | 2.2064 | 2.2064 |
C2 | 2.2294 | | 1.5732 | 1.5732 | 3.2076 | 3.5098 | 1.1045 | 1.1047 | 2.2867 | 2.2867 | 2.2438 | 2.2438 | 3.0437 | 3.5685 | 3.9420 | 3.9420 | 4.1966 | 4.1966 | C3 | 1.5874 | 1.5732 | | 2.2189 | 2.6176 | 2.6631 | 2.2445 | 2.2835 | 1.1052 | 3.1114 | 1.1050 | 2.8649 | 2.7905 | 2.8636 | 3.5686 | 2.9606 | 3.0020 | 3.6029 | C4 | 1.5874 | 1.5732 | 2.2189 | | 2.6176 | 2.6631 | 2.2445 | 2.2835 | 3.1114 | 1.1052 | 2.8649 | 1.1050 | 2.7905 | 2.8636 | 2.9606 | 3.5686 | 3.6029 | 3.0020 | C5 | 1.5624 | 3.2076 | 2.6176 | 2.6176 | | 2.5684 | 3.3694 | 4.2735 | 2.7247 | 2.7247 | 3.5030 | 3.5030 | 1.1035 | 3.5334 | 1.1041 | 1.1041 | 2.8189 | 2.8189 | C6 | 1.5603 | 3.5098 | 2.6631 | 2.6631 | 2.5684 | | 4.3437 | 4.1401 | 3.3964 | 3.3964 | 2.7285 | 2.7285 | 3.5331 | 1.1036 | 2.8193 | 2.8193 | 1.1042 | 1.1042 | H7 | 2.8770 | 1.1045 | 2.2445 | 2.2445 | 3.3694 | 4.3437 | | 1.7976 | 2.5019 | 2.5019 | 3.0988 | 3.0988 | 2.8344 | 4.5685 | 4.1127 | 4.1127 | 4.9579 | 4.9579 | H8 | 3.1201 | 1.1047 | 2.2835 | 2.2835 | 4.2735 | 4.1401 | 1.7976 | | 3.0239 | 3.0239 | 2.4969 | 2.4969 | 4.1477 | 3.9103 | 4.9732 | 4.9732 | 4.8571 | 4.8571 | H9 | 2.2961 | 2.2867 | 1.1052 | 3.1114 | 2.7247 | 3.3964 | 2.5019 | 3.0239 | | 3.8125 | 1.7992 | 3.9040 | 2.6759 | 3.7662 | 3.8102 | 2.7742 | 3.4484 | 4.3256 | H10 | 2.2961 | 2.2867 | 3.1114 | 1.1052 | 2.7247 | 3.3964 | 2.5019 | 3.0239 | 3.8125 | | 3.9040 | 1.7992 | 2.6759 | 3.7662 | 2.7742 | 3.8102 | 4.3256 | 3.4484 | H11 | 2.2548 | 2.2438 | 1.1050 | 2.8649 | 3.5030 | 2.7285 | 3.0988 | 2.4969 | 1.7992 | 3.9040 | | 3.1487 | 3.8437 | 2.5563 | 4.3896 | 3.6926 | 2.9521 | 3.7877 | H12 | 2.2548 | 2.2438 | 2.8649 | 1.1050 | 3.5030 | 2.7285 | 3.0988 | 2.4969 | 3.9040 | 1.7992 | 3.1487 | | 3.8437 | 2.5563 | 3.6926 | 4.3896 | 3.7877 | 2.9521 | H13 | 2.2099 | 3.0437 | 2.7905 | 2.7905 | 1.1035 | 3.5331 | 2.8344 | 4.1477 | 2.6759 | 2.6759 | 3.8437 | 3.8437 | | 4.3864 | 1.7887 | 1.7887 | 3.8326 | 3.8326 | H14 | 2.2078 | 3.5685 | 2.8636 | 2.8636 | 3.5334 | 1.1036 | 4.5685 | 3.9103 | 3.7662 | 3.7662 | 2.5563 | 2.5563 | 4.3864 | | 3.8331 | 3.8331 | 1.7891 | 1.7891 | H15 | 2.2080 | 3.9420 | 3.5686 | 2.9606 | 1.1041 | 2.8193 | 4.1127 | 4.9732 | 3.8102 | 2.7742 | 4.3896 | 3.6926 | 1.7887 | 3.8331 | | 1.7892 | 3.1722 | 2.6196 | H16 | 2.2080 | 3.9420 | 2.9606 | 3.5686 | 1.1041 | 2.8193 | 4.1127 | 4.9732 | 2.7742 | 3.8102 | 3.6926 | 4.3896 | 1.7887 | 3.8331 | 1.7892 | | 2.6196 | 3.1722 | H17 | 2.2064 | 4.1966 | 3.0020 | 3.6029 | 2.8189 | 1.1042 | 4.9579 | 4.8571 | 3.4484 | 4.3256 | 2.9521 | 3.7877 | 3.8326 | 1.7891 | 3.1722 | 2.6196 | | 1.7886 | H18 | 2.2064 | 4.1966 | 3.6029 | 3.0020 | 2.8189 | 1.1042 | 4.9579 | 4.8571 | 4.3256 | 3.4484 | 3.7877 | 2.9521 | 3.8326 | 1.7891 | 2.6196 | 3.1722 | 1.7886 | |
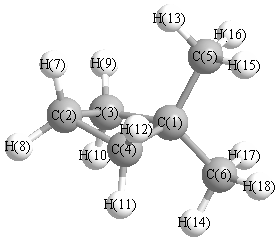 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
89.720 |
|
C1 |
C3 |
H9 |
115.865 |
| C1 |
C3 |
H11 |
112.517 |
|
C1 |
C4 |
C2 |
89.720 |
| C1 |
C4 |
H10 |
115.865 |
|
C1 |
C4 |
H12 |
112.517 |
| C1 |
C5 |
H13 |
110.804 |
|
C1 |
C5 |
H15 |
110.617 |
| C1 |
C5 |
H16 |
110.617 |
|
C1 |
C6 |
H14 |
110.777 |
| C1 |
C6 |
H17 |
110.638 |
|
C1 |
C6 |
H18 |
110.638 |
| C2 |
C3 |
H9 |
116.147 |
|
C2 |
C3 |
H11 |
112.646 |
| C2 |
C4 |
H10 |
116.147 |
|
C2 |
C4 |
H12 |
112.646 |
| C3 |
C1 |
C4 |
88.680 |
|
C3 |
C1 |
C5 |
112.411 |
| C3 |
C1 |
C6 |
115.569 |
|
C3 |
C2 |
C4 |
89.693 |
| C3 |
C2 |
H7 |
112.734 |
|
C3 |
C2 |
H8 |
115.918 |
| C4 |
C1 |
C5 |
112.411 |
|
C4 |
C1 |
C6 |
115.569 |
| C4 |
C2 |
H7 |
112.734 |
|
C4 |
C2 |
H8 |
115.918 |
| C5 |
C1 |
C6 |
110.670 |
|
H7 |
C2 |
H8 |
108.912 |
| H9 |
C3 |
H11 |
108.986 |
|
H10 |
C4 |
H12 |
108.986 |
| H13 |
C5 |
H15 |
108.236 |
|
H13 |
C5 |
H16 |
108.236 |
| H14 |
C6 |
H17 |
108.258 |
|
H14 |
C6 |
H18 |
108.258 |
| H15 |
C5 |
H16 |
108.237 |
|
H17 |
C6 |
H18 |
108.178 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
