Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -112.369653 |
| Energy at 298.15K | -112.371064 |
| HF Energy | -112.276516 |
| Nuclear repulsion energy | 29.824896 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3828 |
3403 |
11.83 |
|
|
|
| 2 |
A' |
3136 |
2787 |
90.16 |
|
|
|
| 3 |
A' |
1689 |
1502 |
28.29 |
|
|
|
| 4 |
A' |
1495 |
1328 |
24.37 |
|
|
|
| 5 |
A' |
1334 |
1186 |
71.27 |
|
|
|
| 6 |
A" |
1191 |
1058 |
49.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6336.0 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 5632.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.010 |
0.765 |
0.000 |
| O2 |
0.010 |
-0.584 |
0.000 |
| H3 |
-1.128 |
0.940 |
0.000 |
| H4 |
0.991 |
-0.859 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3493 | 1.1511 | 1.8977 |
O2 | 1.3493 | | 1.9020 | 1.0194 | H3 | 1.1511 | 1.9020 | | 2.7800 | H4 | 1.8977 | 1.0194 | 2.7800 | |
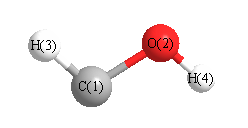 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
105.634 |
|
O2 |
C1 |
H3 |
98.742 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -112.360305 |
| Energy at 298.15K | -112.361704 |
| HF Energy | -112.267497 |
| Nuclear repulsion energy | 29.834408 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3716 |
3303 |
22.86 |
|
|
|
| 2 |
A' |
3082 |
2740 |
109.13 |
|
|
|
| 3 |
A' |
1661 |
1477 |
24.30 |
|
|
|
| 4 |
A' |
1497 |
1331 |
31.32 |
|
|
|
| 5 |
A' |
1351 |
1201 |
35.61 |
|
|
|
| 6 |
A" |
1103 |
980 |
24.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6204.7 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 5515.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.128 |
0.760 |
0.000 |
| O2 |
0.128 |
-0.580 |
0.000 |
| H3 |
-0.985 |
1.061 |
0.000 |
| H4 |
-0.813 |
-0.985 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
H3 |
H4 |
| C1 | | 1.3399 | 1.1530 | 1.9828 |
O2 | 1.3399 | | 1.9826 | 1.0251 | H3 | 1.1530 | 1.9826 | | 2.0528 | H4 | 1.9828 | 1.0251 | 2.0528 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
113.271 |
|
O2 |
C1 |
H3 |
105.115 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
