return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7 (n-Propyl radical)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 2A"
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-116.384595
Energy at 298.15K 
HF Energy-116.240756
Nuclear repulsion energy74.567915
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3624 3221 2.95      
2 A 3557 3162 3.65      
3 A 3555 3160 3.73      
4 A 3490 3102 3.69      
5 A 3455 3071 2.89      
6 A 3376 3001 3.90      
7 A 3374 3000 3.84      
8 A 1744 1550 0.91      
9 A 1736 1544 1.17      
10 A 1714 1524 0.02      
11 A 1655 1471 5.73      
12 A 1625 1444 0.48      
13 A 1544 1372 8.43      
14 A 1433 1274 0.13      
15 A 1312 1166 0.53      
16 A 1198 1065 0.32      
17 A 1197 1064 0.64      
18 A 1025 911 1.44      
19 A 983 874 0.18      
20 A 848 754 1.02      
21 A 597 530 9.06      
22 A 390 346 0.58      
23 A 241 214 0.03      
24 A 172 153 0.58      

Unscaled Zero Point Vibrational Energy (zpe) 21922.4 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 19486.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
1.05719 0.28802 0.25082

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.323 -0.314 -0.079
C2 0.077 0.567 0.064
C3 -1.248 -0.251 -0.041
H4 -2.124 0.410 0.077
H5 -1.323 -0.752 -1.021
H6 -1.291 -1.024 0.745
H7 0.095 1.095 1.041
H8 0.089 1.351 -0.718
H9 2.291 0.171 -0.261
H10 1.354 -1.267 0.465

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9 H10
C11.53252.57233.52512.84212.83102.17872.16881.09771.0978
C21.53251.56142.20642.20832.20661.11081.10712.27212.2708
C32.57231.56141.10331.10321.10342.18832.19403.57092.8391
H43.52512.20641.10331.78841.78832.51432.53214.43363.8805
H52.84212.20831.10321.78841.78703.11042.55093.80573.1041
H62.83102.20661.10341.78831.78702.54993.11233.90752.6707
H72.17871.11082.18832.51433.11042.54991.77742.71492.7382
H82.16881.10712.19402.53212.55093.11231.77742.53963.1393
H91.09772.27213.57094.43363.80573.90752.71492.53961.8635
H101.09782.27082.83913.88053.10412.67072.73823.13931.8635

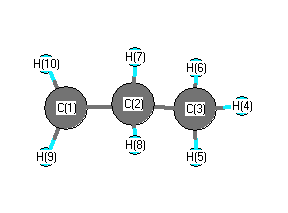
picture of n-Propyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.483 C1 C2 H7 109.991
C1 C2 H8 109.445 C2 C1 H9 118.566
C2 C1 H10 118.449 C2 C3 H4 110.610
C2 C3 H5 110.772 C2 C3 H6 110.627
C3 C2 H7 108.781 C3 C2 H8 109.432
H4 C3 H5 108.295 H4 C3 H6 108.275
H5 C3 H6 108.170 H7 C2 H8 106.530
H9 C1 H10 116.160
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability