Jump to
S2C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -75.895657 |
| Energy at 298.15K | -75.895436 |
| HF Energy | -75.785881 |
| Nuclear repulsion energy | 23.224676 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3449 |
3066 |
65.64 |
|
|
|
| 2 |
A1 |
1696 |
1508 |
59.90 |
|
|
|
| 3 |
A1 |
1374 |
1221 |
1.06 |
|
|
|
| 4 |
B1 |
849 |
755 |
49.15 |
|
|
|
| 5 |
B2 |
3594 |
3195 |
23.00 |
|
|
|
| 6 |
B2 |
473 |
420 |
0.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5716.9 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 5081.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.496 |
| C2 |
0.000 |
0.000 |
0.849 |
| H3 |
0.000 |
0.942 |
-1.058 |
| H4 |
0.000 |
-0.942 |
-1.058 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3444 | 1.0972 | 1.0972 |
C2 | 1.3444 | | 2.1267 | 2.1267 | H3 | 1.0972 | 2.1267 | | 1.8844 | H4 | 1.0972 | 2.1267 | 1.8844 | |
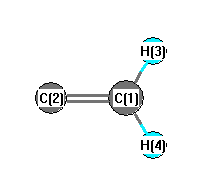 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C2 |
C1 |
H3 |
120.824 |
|
C2 |
C1 |
H4 |
120.824 |
| H3 |
C1 |
H4 |
118.351 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -75.845102 |
| Energy at 298.15K | -75.845175 |
| HF Energy | -75.784833 |
| Nuclear repulsion energy | 23.281674 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3405 |
3027 |
5.04 |
|
|
|
| 2 |
A1 |
1669 |
1483 |
0.02 |
|
|
|
| 3 |
A1 |
1469 |
1306 |
2.46 |
|
|
|
| 4 |
B1 |
900 |
800 |
14.18 |
|
|
|
| 5 |
B2 |
3544 |
3150 |
0.39 |
|
|
|
| 6 |
B2 |
1122 |
997 |
0.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6054.0 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 5381.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.488 |
| C2 |
0.000 |
0.000 |
0.847 |
| H3 |
0.000 |
0.932 |
-1.076 |
| H4 |
0.000 |
-0.932 |
-1.076 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3355 | 1.1020 | 1.1020 |
C2 | 1.3355 | | 2.1373 | 2.1373 | H3 | 1.1020 | 2.1373 | | 1.8643 | H4 | 1.1020 | 2.1373 | 1.8643 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
