return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2 (benzyl radical)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2B1
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-266.191934
Energy at 298.15K-266.198646
HF Energy-265.883792
Nuclear repulsion energy257.586059
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3549 3155 1.39      
2 A1 3535 3142 1.42      
3 A1 3522 3131 0.59      
4 A1 3472 3086 4.71      
5 A1 1682 1495 0.41      
6 A1 1649 1466 7.05      
7 A1 1633 1452 1.52      
8 A1 1343 1194 0.54      
9 A1 1258 1118 0.00      
10 A1 1112 989 0.47      
11 A1 1039 924 1.02      
12 A1 875 778 0.00      
13 A1 558 496 0.34      
14 A2 1106 983 0.00      
15 A2 939 835 0.00      
16 A2 582 517 0.00      
17 A2 429 381 0.00      
18 B1 1115 991 0.04      
19 B1 1004 893 0.64      
20 B1 831 739 14.96      
21 B1 762 677 5.60      
22 B1 662 588 11.64      
23 B1 515 457 3.09      
24 B1 224 199 0.36      
25 B2 3642 3237 0.09      
26 B2 3543 3149 1.12      
27 B2 3527 3135 0.01      
28 B2 1708 1518 0.97      
29 B2 1603 1425 5.56      
30 B2 1464 1301 0.77      
31 B2 1322 1176 0.00      
32 B2 1215 1080 0.07      
33 B2 1198 1065 0.09      
34 B2 1056 939 0.00      
35 B2 666 592 0.01      
36 B2 371 330 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 27353.9 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 24314.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.18123 0.08870 0.05955

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.000
C2 0.000 1.231 0.250
C3 0.000 1.221 -1.141
C4 0.000 0.000 -1.849
C5 0.000 -1.221 -1.141
C6 0.000 -1.231 0.250
C7 0.000 0.000 2.423
H8 0.000 2.184 0.799
H9 0.000 2.171 -1.694
H10 0.000 0.000 -2.948
H11 0.000 -2.171 -1.694
H12 0.000 -2.184 0.799
H13 0.000 0.938 2.991
H14 0.000 -0.938 2.991

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14
C11.44072.46442.84842.46441.44071.42342.19293.45983.94723.45982.19292.20112.2011
C21.44071.39142.43342.81912.46132.49691.09952.16023.42673.91823.45812.75623.4945
C32.46441.39141.41152.44242.81913.76732.16541.09922.18073.43683.91864.14174.6619
C42.84842.43341.41151.41152.43344.27183.43192.17631.09882.17633.43194.92984.9298
C52.46442.81912.44241.41151.39143.76733.91863.43682.18071.09922.16544.66194.1417
C61.44072.46132.81912.43341.39142.49693.45813.91823.42672.16021.09953.49452.7562
C71.42342.49693.76734.27183.76732.49692.72144.65465.37064.65462.72141.09621.0962
H82.19291.09952.16543.43193.91863.45812.72142.49334.33645.01774.36732.52173.8141
H93.45982.16021.09922.17633.43683.91824.65462.49332.50654.34165.01774.84515.6228
H103.94723.42672.18071.09882.18073.42675.37064.33642.50652.50654.33646.01226.0122
H113.45983.91823.43682.17631.09922.16024.65465.01774.34162.50652.49335.62284.8451
H122.19293.45813.91863.43192.16541.09952.72144.36735.01774.33642.49333.81412.5217
H132.20112.75624.14174.92984.66193.49451.09622.52174.84516.01225.62283.81411.8750
H142.20113.49454.66194.92984.14172.75621.09623.81415.62286.01224.84512.52171.8750

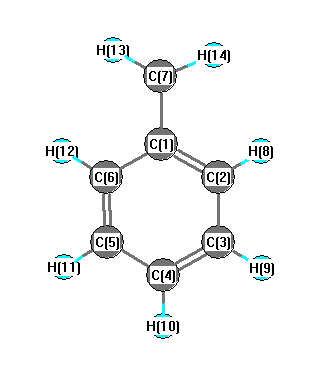
picture of benzyl radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.942 C1 C2 H8 118.751
C1 C6 C5 120.942 C1 C6 H12 118.751
C1 C7 H13 121.213 C1 C7 H14 121.213
C2 C1 C6 117.337 C2 C1 C7 121.332
C2 C3 C4 120.489 C2 C3 H9 119.850
C3 C2 H8 120.307 C3 C4 C5 119.802
C3 C4 H10 120.099 C4 C3 H9 119.662
C4 C5 C6 120.489 C4 C5 H11 119.662
C5 C1 C7 150.295 C5 C4 H10 120.099
C5 C6 H12 120.307 C6 C5 H11 119.850
H13 C7 H14 117.573
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability