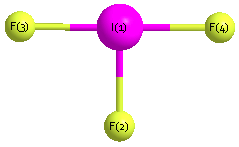
Vibrational Frequencies calculated at M06-2X/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
599 |
599 |
0.14 |
8.46 |
0.15 |
0.26 |
| 2 |
A1 |
570 |
570 |
3.19 |
12.20 |
0.41 |
0.58 |
| 3 |
A1 |
283 |
283 |
12.13 |
0.13 |
0.75 |
0.86 |
| 4 |
B1 |
147 |
147 |
3.54 |
0.01 |
0.75 |
0.86 |
| 5 |
B2 |
649 |
649 |
29.20 |
0.32 |
0.75 |
0.86 |
| 6 |
B2 |
340 |
340 |
7.02 |
2.78 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1294.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1294.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
I |
0.660 |
|
|
|
| 2 |
F |
-0.205 |
|
|
|
| 3 |
F |
-0.228 |
|
|
|
| 4 |
F |
-0.228 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.702 |
1.702 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.417 |
0.000 |
0.000 |
| y |
0.000 |
-43.225 |
0.000 |
| z |
0.000 |
0.000 |
-39.380 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.886 |
0.000 |
0.000 |
| y |
0.000 |
-4.827 |
0.000 |
| z |
0.000 |
0.000 |
0.941 |
|
| Polar |
|---|
| 3z2-r2 | 1.882 |
|---|
| x2-y2 | 5.808 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.350 |
0.000 |
0.000 |
| y |
0.000 |
4.750 |
0.000 |
| z |
0.000 |
0.000 |
1.981 |
<r2> (average value of r
2) Å
2
| <r2> |
125.604 |
| (<r2>)1/2 |
11.207 |