Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Geometric Data calculated at MP2=FULL/STO-3G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -231.755217 |
| Energy at 298.15K | -231.768380 |
| HF Energy | -231.433356 |
| Nuclear repulsion energy | 244.967588 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3550 |
3156 |
4.64 |
|
|
|
| 2 |
A' |
3549 |
3155 |
3.41 |
|
|
|
| 3 |
A' |
3534 |
3141 |
5.53 |
|
|
|
| 4 |
A' |
3445 |
3062 |
3.81 |
|
|
|
| 5 |
A' |
3444 |
3061 |
5.92 |
|
|
|
| 6 |
A' |
3406 |
3028 |
0.15 |
|
|
|
| 7 |
A' |
3368 |
2993 |
4.15 |
|
|
|
| 8 |
A' |
3367 |
2993 |
3.50 |
|
|
|
| 9 |
A' |
1735 |
1542 |
0.96 |
|
|
|
| 10 |
A' |
1734 |
1541 |
1.36 |
|
|
|
| 11 |
A' |
1707 |
1518 |
0.02 |
|
|
|
| 12 |
A' |
1620 |
1440 |
0.25 |
|
|
|
| 13 |
A' |
1619 |
1439 |
1.26 |
|
|
|
| 14 |
A' |
1551 |
1379 |
6.90 |
|
|
|
| 15 |
A' |
1539 |
1368 |
3.78 |
|
|
|
| 16 |
A' |
1447 |
1287 |
13.78 |
|
|
|
| 17 |
A' |
1385 |
1231 |
0.19 |
|
|
|
| 18 |
A' |
1329 |
1181 |
1.35 |
|
|
|
| 19 |
A' |
1249 |
1110 |
0.40 |
|
|
|
| 20 |
A' |
1171 |
1041 |
2.40 |
|
|
|
| 21 |
A' |
1073 |
954 |
0.26 |
|
|
|
| 22 |
A' |
1028 |
914 |
0.88 |
|
|
|
| 23 |
A' |
967 |
859 |
0.46 |
|
|
|
| 24 |
A' |
884 |
786 |
0.33 |
|
|
|
| 25 |
A' |
729 |
648 |
0.96 |
|
|
|
| 26 |
A' |
457 |
406 |
0.25 |
|
|
|
| 27 |
A' |
338 |
301 |
0.06 |
|
|
|
| 28 |
A' |
63 |
56 |
0.00 |
|
|
|
| 29 |
A" |
3546 |
3152 |
2.70 |
|
|
|
| 30 |
A" |
3546 |
3152 |
3.06 |
|
|
|
| 31 |
A" |
3533 |
3140 |
0.45 |
|
|
|
| 32 |
A" |
3405 |
3027 |
6.04 |
|
|
|
| 33 |
A" |
1737 |
1544 |
0.66 |
|
|
|
| 34 |
A" |
1736 |
1543 |
0.04 |
|
|
|
| 35 |
A" |
1694 |
1506 |
0.54 |
|
|
|
| 36 |
A" |
1472 |
1308 |
5.74 |
|
|
|
| 37 |
A" |
1429 |
1270 |
0.25 |
|
|
|
| 38 |
A" |
1403 |
1247 |
0.27 |
|
|
|
| 39 |
A" |
1287 |
1144 |
0.95 |
|
|
|
| 40 |
A" |
1265 |
1124 |
0.12 |
|
|
|
| 41 |
A" |
1119 |
995 |
1.41 |
|
|
|
| 42 |
A" |
1046 |
930 |
0.01 |
|
|
|
| 43 |
A" |
1025 |
911 |
0.68 |
|
|
|
| 44 |
A" |
897 |
798 |
0.29 |
|
|
|
| 45 |
A" |
405 |
360 |
0.01 |
|
|
|
| 46 |
A" |
272 |
241 |
0.08 |
|
|
|
| 47 |
A" |
235 |
209 |
0.00 |
|
|
|
| 48 |
A" |
219 |
194 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41277.7 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 36691.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.183 |
0.045 |
1.109 |
| C2 |
0.183 |
0.045 |
-1.109 |
| C3 |
-0.651 |
0.800 |
0.000 |
| C4 |
0.778 |
-0.910 |
0.000 |
| C5 |
-0.607 |
2.354 |
0.000 |
| C6 |
0.183 |
-2.349 |
0.000 |
| H7 |
0.953 |
0.689 |
1.571 |
| H8 |
-0.404 |
-0.445 |
1.907 |
| H9 |
0.953 |
0.689 |
-1.571 |
| H10 |
-0.404 |
-0.445 |
-1.907 |
| H11 |
-1.705 |
0.458 |
0.000 |
| H12 |
1.885 |
-0.968 |
0.000 |
| H13 |
0.436 |
2.715 |
0.000 |
| H14 |
-0.920 |
-2.316 |
0.000 |
| H15 |
-1.112 |
2.759 |
-0.894 |
| H16 |
-1.112 |
2.759 |
0.894 |
| H17 |
0.511 |
-2.906 |
-0.894 |
| H18 |
0.511 |
-2.906 |
0.894 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.2179 | 1.5795 | 1.5803 | 2.6803 | 2.6382 | 1.1052 | 1.1053 | 2.8621 | 3.1116 | 2.2280 | 2.2699 | 2.9019 | 2.8319 | 3.6130 | 3.0146 | 3.5823 | 2.9774 |
C2 | 2.2179 | | 1.5795 | 1.5803 | 2.6803 | 2.6382 | 2.8621 | 3.1116 | 1.1052 | 1.1053 | 2.2280 | 2.2699 | 2.9019 | 2.8319 | 3.0146 | 3.6130 | 2.9774 | 3.5823 | C3 | 1.5795 | 1.5795 | | 2.2289 | 1.5549 | 3.2570 | 2.2480 | 2.2911 | 2.2480 | 2.2911 | 1.1077 | 3.0912 | 2.2023 | 3.1269 | 2.2024 | 2.2024 | 3.9857 | 3.9857 | C4 | 1.5803 | 1.5803 | 2.2289 | | 3.5461 | 1.5568 | 2.2490 | 2.2915 | 2.2490 | 2.2915 | 2.8355 | 1.1078 | 3.6412 | 2.2047 | 4.2234 | 4.2234 | 2.2035 | 2.2035 | C5 | 2.6803 | 2.6803 | 1.5549 | 3.5461 | | 4.7685 | 2.7698 | 3.3934 | 2.7698 | 3.3934 | 2.1910 | 4.1524 | 1.1036 | 4.6802 | 1.1041 | 1.1041 | 5.4517 | 5.4517 | C6 | 2.6382 | 2.6382 | 3.2570 | 1.5568 | 4.7685 | | 3.5059 | 2.7576 | 3.5059 | 2.7576 | 3.3826 | 2.1917 | 5.0698 | 1.1035 | 5.3445 | 5.3445 | 1.1041 | 1.1041 | H7 | 1.1052 | 2.8621 | 2.2480 | 2.2490 | 2.7698 | 3.5059 | | 1.7998 | 3.1426 | 3.9022 | 3.0960 | 2.4666 | 2.6150 | 3.8738 | 3.8243 | 3.0009 | 4.3822 | 3.6853 | H8 | 1.1053 | 3.1116 | 2.2911 | 2.2915 | 3.3934 | 2.7576 | 1.7998 | | 3.9022 | 3.8146 | 2.4794 | 3.0244 | 3.7854 | 2.7208 | 4.3148 | 3.4344 | 3.8397 | 2.8141 | H9 | 2.8621 | 1.1052 | 2.2480 | 2.2490 | 2.7698 | 3.5059 | 3.1426 | 3.9022 | | 1.7998 | 3.0960 | 2.4666 | 2.6150 | 3.8738 | 3.0009 | 3.8243 | 3.6853 | 4.3822 | H10 | 3.1116 | 1.1053 | 2.2911 | 2.2915 | 3.3934 | 2.7576 | 3.9022 | 3.8146 | 1.7998 | | 2.4794 | 3.0244 | 3.7854 | 2.7208 | 3.4344 | 4.3148 | 2.8141 | 3.8397 | H11 | 2.2280 | 2.2280 | 1.1077 | 2.8355 | 2.1910 | 3.3826 | 3.0960 | 2.4794 | 3.0960 | 2.4794 | | 3.8626 | 3.1108 | 2.8826 | 2.5385 | 2.5385 | 4.1269 | 4.1269 | H12 | 2.2699 | 2.2699 | 3.0912 | 1.1078 | 4.1524 | 2.1917 | 2.4666 | 3.0244 | 2.4666 | 3.0244 | 3.8626 | | 3.9572 | 3.1121 | 4.8652 | 4.8652 | 2.5385 | 2.5385 | H13 | 2.9019 | 2.9019 | 2.2023 | 3.6412 | 1.1036 | 5.0698 | 2.6150 | 3.7854 | 2.6150 | 3.7854 | 3.1108 | 3.9572 | | 5.2101 | 1.7887 | 1.7887 | 5.6923 | 5.6923 | H14 | 2.8319 | 2.8319 | 3.1269 | 2.2047 | 4.6802 | 1.1035 | 3.8738 | 2.7208 | 3.8738 | 2.7208 | 2.8826 | 3.1121 | 5.2101 | | 5.1563 | 5.1563 | 1.7884 | 1.7884 | H15 | 3.6130 | 3.0146 | 2.2024 | 4.2234 | 1.1041 | 5.3445 | 3.8243 | 4.3148 | 3.0009 | 3.4344 | 2.5385 | 4.8652 | 1.7887 | 5.1563 | | 1.7884 | 5.8932 | 6.1587 | H16 | 3.0146 | 3.6130 | 2.2024 | 4.2234 | 1.1041 | 5.3445 | 3.0009 | 3.4344 | 3.8243 | 4.3148 | 2.5385 | 4.8652 | 1.7887 | 5.1563 | 1.7884 | | 6.1587 | 5.8932 | H17 | 3.5823 | 2.9774 | 3.9857 | 2.2035 | 5.4517 | 1.1041 | 4.3822 | 3.8397 | 3.6853 | 2.8141 | 4.1269 | 2.5385 | 5.6923 | 1.7884 | 5.8932 | 6.1587 | | 1.7890 | H18 | 2.9774 | 3.5823 | 3.9857 | 2.2035 | 5.4517 | 1.1041 | 3.6853 | 2.8141 | 4.3822 | 3.8397 | 4.1269 | 2.5385 | 5.6923 | 1.7884 | 6.1587 | 5.8932 | 1.7890 | |
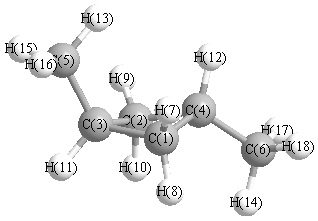 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
89.191 |
|
C1 |
C3 |
C5 |
117.543 |
| C1 |
C3 |
H11 |
110.793 |
|
C1 |
C4 |
C2 |
89.132 |
| C1 |
C4 |
C6 |
114.480 |
|
C1 |
C4 |
H12 |
114.065 |
| C2 |
C3 |
C5 |
117.543 |
|
C2 |
C3 |
H11 |
110.793 |
| C2 |
C4 |
C6 |
114.480 |
|
C2 |
C4 |
H12 |
114.065 |
| C3 |
C1 |
C4 |
89.723 |
|
C3 |
C1 |
H7 |
112.526 |
| C3 |
C1 |
H8 |
116.038 |
|
C3 |
C2 |
C4 |
89.723 |
| C3 |
C2 |
H9 |
112.526 |
|
C3 |
C2 |
H10 |
116.038 |
| C3 |
C5 |
H13 |
110.720 |
|
C3 |
C5 |
H15 |
110.695 |
| C3 |
C5 |
H16 |
110.695 |
|
C4 |
C1 |
H7 |
112.545 |
| C4 |
C1 |
H8 |
116.010 |
|
C4 |
C2 |
H9 |
112.545 |
| C4 |
C2 |
H10 |
116.010 |
|
C4 |
C6 |
H14 |
110.779 |
| C4 |
C6 |
H17 |
110.655 |
|
C4 |
C6 |
H18 |
110.655 |
| C5 |
C3 |
H11 |
109.602 |
|
C6 |
C4 |
H12 |
109.520 |
| H7 |
C1 |
H8 |
109.026 |
|
H9 |
C2 |
H10 |
109.026 |
| H13 |
C5 |
H15 |
108.231 |
|
H13 |
C5 |
H16 |
108.231 |
| H14 |
C6 |
H17 |
108.214 |
|
H14 |
C6 |
H18 |
108.214 |
| H15 |
C5 |
H16 |
108.171 |
|
H17 |
C6 |
H18 |
108.227 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
