return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (Pentalene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-303.091260
Energy at 298.15K-303.096631
HF Energy-302.591508
Nuclear repulsion energy308.806178
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3553 3158 0.00      
2 Ag 3530 3137 0.00      
3 Ag 3522 3131 0.00      
4 Ag 1760 1565 0.00      
5 Ag 1670 1484 0.00      
6 Ag 1490 1324 0.00      
7 Ag 1431 1272 0.00      
8 Ag 1273 1131 0.00      
9 Ag 1183 1051 0.00      
10 Ag 1036 921 0.00      
11 Ag 900 800 0.00      
12 Ag 756 672 0.00      
13 Ag 684 608 0.00      
14 Au 985 876 0.03      
15 Au 920 818 9.90      
16 Au 788 701 23.50      
17 Au 537 478 1.22      
18 Au 313 278 0.26      
19 Au 168 150 2.78      
20 Bg 995 884 0.00      
21 Bg 925 822 0.00      
22 Bg 822 730 0.00      
23 Bg 690 614 0.00      
24 Bg 562 500 0.00      
25 Bu 3554 3159 1.22      
26 Bu 3528 3136 18.02      
27 Bu 3522 3131 2.86      
28 Bu 1738 1545 14.45      
29 Bu 1635 1454 2.20      
30 Bu 1449 1288 8.14      
31 Bu 1353 1202 10.81      
32 Bu 1204 1070 4.07      
33 Bu 1107 984 10.68      
34 Bu 1017 904 4.67      
35 Bu 894 795 1.80      
36 Bu 471 419 0.62      

Unscaled Zero Point Vibrational Energy (zpe) 25981.7 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 23095.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.16724 0.06988 0.04929

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.372 1.116 0.000
C2 2.205 -0.174 0.000
C3 1.372 -1.269 0.000
C4 -1.372 -1.116 0.000
C5 -2.205 0.174 0.000
C6 -1.372 1.269 0.000
C7 0.047 0.752 0.000
C8 -0.047 -0.752 0.000
H9 1.811 2.130 0.000
H10 3.309 -0.185 0.000
H11 1.673 -2.329 0.000
H12 -1.811 -2.130 0.000
H13 -3.309 0.185 0.000
H14 -1.673 2.329 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.53562.38493.53773.69922.74881.37462.34571.10492.33293.45814.54674.77253.2778
C21.53561.37543.69924.42373.85732.34822.32482.33761.10362.21954.46725.52524.6155
C32.38491.37542.74883.85733.73792.41631.51063.42712.21891.10193.29804.90144.7134
C43.53773.69922.74881.53562.38492.34571.37464.54674.77253.27781.10492.33293.4581
C53.69924.42373.85731.53561.37542.32482.34824.46725.52524.61552.33761.10362.2195
C62.74883.85733.73792.38491.37541.51062.41633.29804.90144.71343.42712.21891.1019
C71.37462.34822.41632.34572.32481.51061.50602.23903.39343.48323.42883.40292.3337
C82.34572.32481.51061.37462.34822.41631.50603.42883.40292.33372.23903.39343.4832
H91.10492.33763.42714.54674.46723.29802.23903.42882.75714.46115.59215.47683.4899
H102.33291.10362.21894.77255.52524.90143.39343.40292.75712.69645.47686.62745.5799
H113.45812.21951.10193.27784.61554.71343.48322.33374.46112.69643.48995.57995.7349
H124.54674.46723.29801.10492.33763.42713.42882.23905.59215.47683.48992.75714.4611
H134.77255.52524.90142.33291.10362.21893.40293.39345.47686.62745.57992.75712.6964
H143.27784.61554.71343.45812.21951.10192.33373.48323.48995.57995.73494.46112.6964

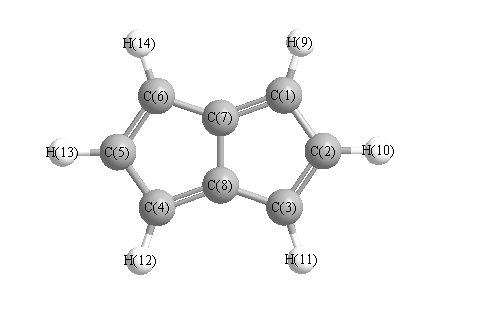
picture of Pentalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 109.902 C1 C2 H10 123.409
C1 C7 C6 144.592 C1 C7 C8 108.956
C2 C1 C7 107.456 C2 C1 H9 123.750
C2 C3 C8 107.234 C2 C3 H11 126.905
C3 C2 H10 126.689 C3 C8 C4 144.592
C3 C8 C7 106.452 C4 C5 C6 109.902
C4 C5 H13 123.409 C4 C8 C7 108.956
C5 C4 C8 107.456 C5 C4 H12 123.750
C5 C6 C7 107.234 C5 C6 H14 126.905
C6 C5 H13 126.689 C6 C7 C8 106.452
C7 C1 H9 128.794 C7 C6 H14 125.861
C8 C3 H11 125.861 C8 C4 H12 128.794
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability