return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (Cyclopentane)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-193.154632
Energy at 298.15K-193.166135
HF Energy-192.889072
Nuclear repulsion energy184.462477
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3535 3143 17.49      
2 A 3529 3137 1.10      
3 A 3529 3137 1.09      
4 A 3525 3133 0.38      
5 A 3523 3132 0.52      
6 A 3417 3038 1.60      
7 A 3412 3033 10.21      
8 A 3412 3033 8.49      
9 A 3408 3030 0.06      
10 A 3408 3029 3.93      
11 A 1740 1546 0.00      
12 A 1725 1533 0.35      
13 A 1725 1533 0.33      
14 A 1715 1524 0.01      
15 A 1712 1522 0.10      
16 A 1538 1367 5.89      
17 A 1538 1367 5.89      
18 A 1527 1357 0.02      
19 A 1491 1325 0.15      
20 A 1468 1305 0.00      
21 A 1429 1270 0.48      
22 A 1392 1238 0.49      
23 A 1379 1226 0.26      
24 A 1357 1206 0.57      
25 A 1346 1196 0.03      
26 A 1201 1067 0.02      
27 A 1173 1043 0.01      
28 A 1120 995 0.17      
29 A 1091 970 0.35      
30 A 1025 911 0.24      
31 A 1022 909 0.20      
32 A 982 873 0.00      
33 A 946 841 0.31      
34 A 901 801 0.22      
35 A 843 750 0.61      
36 A 683 607 0.08      
37 A 600 533 1.59      
38 A 239 213 0.00      
39 A 7 6 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 34805.2 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 30938.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.20966 0.20952 0.12030

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.043 -0.753 0.241
C2 -0.376 1.259 0.122
C3 -1.326 0.005 0.014
C4 -0.389 -1.249 -0.148
C5 1.058 0.737 -0.229
H6 -0.388 1.652 1.155
H7 -0.693 2.076 -0.550
H8 -2.010 0.097 -0.849
H9 -1.951 -0.085 0.921
H10 -0.391 -1.587 -1.201
H11 -0.720 -2.098 0.477
H12 1.840 -1.356 -0.230
H13 1.177 -0.801 1.338
H14 1.221 0.781 -1.322
H15 1.852 1.330 0.261

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C12.46462.49751.56481.56182.94413.41223.35103.14172.19782.23041.10521.10602.19682.2342
C22.46461.57602.52251.56641.10561.10502.22752.21873.13833.39293.44592.85162.20572.2338
C32.49751.57601.57432.50552.21242.23761.10511.10512.21072.23743.45512.94352.97903.4516
C41.56482.52251.57432.45883.18103.36362.22042.22201.10571.10512.23342.20442.84493.4411
C51.56181.56642.50552.45882.20092.22833.19513.32432.90603.42012.23422.19831.10601.1052
H62.94411.10562.21243.18102.20091.78373.01142.34884.00573.82573.99172.91563.07932.4330
H73.41221.10502.23763.36362.22831.78372.39622.90153.73294.29904.27813.91672.43672.7737
H83.35102.22751.10512.22043.19513.01142.39621.78042.36242.87034.16153.96753.33684.2032
H93.14172.21871.10512.22203.32432.34882.90151.78043.03202.40054.16073.23503.98034.1107
H102.19783.13832.21071.10572.90604.00573.73292.36243.03201.78502.44413.08522.86753.9591
H112.23043.39292.23741.10513.42013.82574.29902.87032.40051.78502.75812.45353.91104.2910
H121.10523.44593.45512.23342.23423.99174.27814.16154.16072.44412.75811.79052.47852.7304
H131.10602.85162.94352.20442.19832.91563.91673.96753.23503.08522.45351.79053.09482.4812
H142.19682.20572.97902.84491.10603.07932.43673.33683.98032.86753.91102.47853.09481.7897
H152.23422.23383.45163.44111.10522.43302.77374.20324.11073.95914.29102.73042.48121.7897

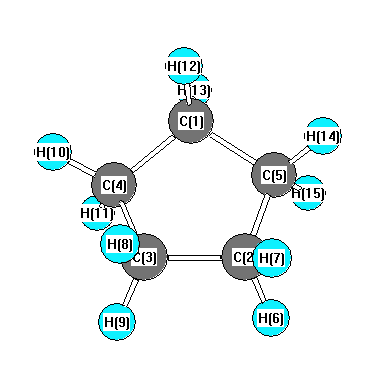
picture of Cyclopentane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 C3 105.427 C1 C4 H10 109.574
C1 C4 H11 112.167 C1 C5 C2 103.974
C1 C5 H14 109.687 C1 C5 H15 112.682
C2 C3 C4 106.398 C2 C3 H8 111.154
C2 C3 H9 110.463 C2 C5 H14 110.058
C2 C5 H15 112.319 C3 C2 C5 105.756
C3 C2 H6 109.944 C3 C2 H7 111.955
C3 C4 H10 109.924 C3 C4 H11 112.051
C4 C1 C5 103.704 C4 C1 H12 112.403
C4 C1 H13 110.068 C4 C3 H8 110.710
C4 C3 H9 110.835 C5 C1 H12 112.677
C5 C1 H13 109.798 C5 C2 H6 109.714
C5 C2 H7 111.894 H6 C2 H7 107.582
H8 C3 H9 107.325 H10 C4 H11 107.689
H12 C1 H13 108.142 H14 C5 H15 108.078
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability