Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -947.135238 |
| Energy at 298.15K | |
| HF Energy | -947.078103 |
| Nuclear repulsion energy | 120.568308 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3542 |
3149 |
5.26 |
|
|
|
| 2 |
A1 |
798 |
709 |
22.08 |
|
|
|
| 3 |
A1 |
299 |
265 |
3.83 |
|
|
|
| 4 |
B1 |
631i |
561i |
27.09 |
|
|
|
| 5 |
B2 |
1330 |
1183 |
112.22 |
|
|
|
| 6 |
B2 |
1054 |
937 |
54.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3195.4 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 2840.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.701 |
| H2 |
0.000 |
0.000 |
1.802 |
| Cl3 |
0.000 |
1.541 |
-0.177 |
| Cl4 |
0.000 |
-1.541 |
-0.177 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.1008 | 1.7732 | 1.7732 |
H2 | 1.1008 | | 2.5076 | 2.5076 | Cl3 | 1.7732 | 2.5076 | | 3.0816 | Cl4 | 1.7732 | 2.5076 | 3.0816 | |
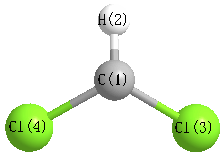 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
119.663 |
|
Cl3 |
C1 |
Cl4 |
120.674 |
| Cl4 |
C1 |
H2 |
119.663 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -947.139780 |
| Energy at 298.15K | -947.140627 |
| HF Energy | -947.080885 |
| Nuclear repulsion energy | 120.589640 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3480 |
3093 |
1.88 |
|
|
|
| 2 |
A' |
826 |
734 |
25.32 |
|
|
|
| 3 |
A' |
720 |
640 |
2.19 |
|
|
|
| 4 |
A' |
296 |
263 |
3.08 |
|
|
|
| 5 |
A" |
1337 |
1188 |
71.67 |
|
|
|
| 6 |
A" |
1016 |
903 |
90.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3837.0 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 3410.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.017 |
0.751 |
0.000 |
| H2 |
-0.669 |
1.622 |
0.000 |
| Cl3 |
0.017 |
-0.180 |
1.527 |
| Cl4 |
0.017 |
-0.180 |
-1.527 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
Cl3 |
Cl4 |
| C1 | | 1.1089 | 1.7888 | 1.7888 |
H2 | 1.1089 | | 2.4601 | 2.4601 | Cl3 | 1.7888 | 2.4601 | | 3.0547 | Cl4 | 1.7888 | 2.4601 | 3.0547 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Cl3 |
C1 |
H2 |
114.141 |
|
Cl3 |
C1 |
Cl4 |
117.259 |
| Cl4 |
C1 |
H2 |
114.141 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
