return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SSSCH3 (dimethyl trisulfide)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-1258.027873
Energy at 298.15K-1258.034020
Nuclear repulsion energy362.048461
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3533 3141 0.57      
2 A 3516 3126 5.37      
3 A 3344 2972 0.42      
4 A 1735 1543 2.23      
5 A 1721 1529 2.51      
6 A 1586 1410 3.14      
7 A 1096 974 0.70      
8 A 1093 971 0.47      
9 A 869 773 0.26      
10 A 598 532 0.84      
11 A 284 252 1.15      
12 A 199 177 1.28      
13 A 154 137 0.06      
14 A 52 46 0.45      
15 B 3533 3141 1.38      
16 B 3516 3126 3.87      
17 B 3344 2972 7.27      
18 B 1735 1542 5.38      
19 B 1721 1529 2.72      
20 B 1584 1408 34.43      
21 B 1095 974 9.15      
22 B 1092 970 2.74      
23 B 868 771 0.54      
24 B 618 549 3.56      
25 B 299 266 1.92      
26 B 164 146 0.12      
27 B 77 68 0.54      

Unscaled Zero Point Vibrational Energy (zpe) 19711.9 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 17521.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.14584 0.04613 0.04566

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 1.106
S2 0.000 1.640 -0.202
S3 0.000 -1.640 -0.202
C4 1.782 1.717 -0.592
C5 -1.782 -1.717 -0.592
H6 1.948 2.556 -1.290
H7 -1.948 -2.556 -1.290
H8 2.402 1.892 0.307
H9 2.141 0.792 -1.080
H10 -2.402 -1.892 0.307
H11 -2.141 -0.792 -1.080

Atom - Atom Distances (Å)
  S1 S2 S3 C4 C5 H6 H7 H8 H9 H10 H11
S12.09842.09843.00143.00144.00874.00873.16033.16113.16033.1611
S22.09843.28061.82563.82122.41144.75262.46812.46434.30173.3574
S32.09843.28063.82121.82564.75262.41144.30173.35742.46812.4643
C43.00141.82563.82124.94981.10405.71521.10541.10605.59814.6826
C53.00143.82121.82564.94985.71521.10405.59814.68261.10541.1060
H64.00872.41144.75261.10405.71526.42751.78811.78686.42305.2891
H74.00874.75262.41145.71521.10406.42756.42305.28911.78811.7868
H83.16032.46814.30171.10545.59811.78816.42301.78916.11505.4558
H93.16112.46433.35741.10604.68261.78685.28911.78915.45584.5657
H103.16034.30172.46815.59811.10546.42301.78816.11505.45581.7891
H113.16113.35742.46434.68261.10605.28911.78685.45584.56571.7891

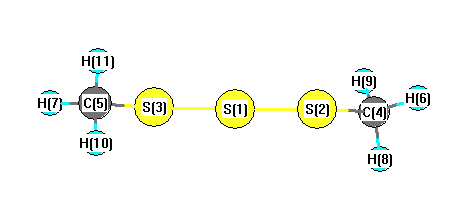
picture of dimethyl trisulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 C4 99.555 S1 S3 C5 99.555
S2 S1 S3 102.826 S2 C4 H6 108.264
S2 C4 H8 112.382 S2 C4 H9 112.060
S3 C5 H7 108.264 S3 C5 H10 112.382
S3 C5 H11 112.060 H6 C4 H8 108.059
H6 C4 H9 107.905 H7 C5 H10 108.059
H7 C5 H11 107.905 H8 C4 H9 108.007
H10 C5 H11 108.007
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability