return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4F2 (orthodifluorobenzene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-423.191369
Energy at 298.15K-423.195736
Nuclear repulsion energy339.352004
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3533 3140 3.60      
2 A1 3517 3127 0.24      
3 A1 1705 1516 3.14      
4 A1 1649 1466 88.14      
5 A1 1418 1261 22.22      
6 A1 1275 1133 2.62      
7 A1 1202 1069 3.42      
8 A1 1103 981 0.00      
9 A1 798 709 1.47      
10 A1 601 534 0.66      
11 A1 263 234 0.38      
12 A2 1065 947 0.00      
13 A2 934 830 0.00      
14 A2 751 668 0.00      
15 A2 575 511 0.00      
16 A2 195 173 0.00      
17 B1 1020 907 2.58      
18 B1 819 728 26.94      
19 B1 478 425 0.02      
20 B1 301 268 0.17      
21 B2 3524 3132 9.51      
22 B2 3511 3121 0.39      
23 B2 1751 1556 8.63      
24 B2 1593 1416 14.51      
25 B2 1423 1265 7.70      
26 B2 1320 1173 1.84      
27 B2 1189 1056 1.33      
28 B2 914 812 2.67      
29 B2 552 491 2.41      
30 B2 439 390 0.94      

Unscaled Zero Point Vibrational Energy (zpe) 19707.9 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 17518.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.10295 0.07091 0.04199

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.723 -0.558
C2 0.000 -0.723 -0.558
C3 0.000 -1.425 0.685
C4 0.000 -0.710 1.911
C5 0.000 0.710 1.911
C6 0.000 1.425 0.685
F7 0.000 1.407 -1.751
F8 0.000 -1.407 -1.751
H9 0.000 -2.524 0.669
H10 0.000 -1.261 2.863
H11 0.000 1.261 2.863
H12 0.000 2.524 0.669

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 F7 F8 H9 H10 H11 H12
C11.44552.48132.85482.46921.42781.37532.44113.47143.95473.46332.1800
C21.44551.42782.46922.85482.48132.44111.37532.18003.46333.95473.4714
C32.48131.42781.41882.46152.84913.73552.43661.10002.18393.45763.9490
C42.85482.46921.41881.42012.46154.23013.72802.19871.09992.18893.4647
C52.46922.85482.46151.42011.41883.72804.23013.46472.18891.09992.1987
C61.42782.48132.84912.46151.41882.43663.73553.94903.45762.18391.1000
F71.37532.44113.73554.23013.72802.43662.81384.61675.33004.61662.6659
F82.44111.37532.43663.72804.23013.73552.81382.66594.61665.33004.6167
H93.47142.18001.10002.19873.46473.94904.61672.66592.53174.37525.0489
H103.95473.46332.18391.09992.18893.45765.33004.61662.53172.52194.3752
H113.46333.95473.45762.18891.09992.18394.61665.33004.37522.52192.5317
H122.18003.47143.94903.46472.19871.10002.66594.61675.04894.37522.5317

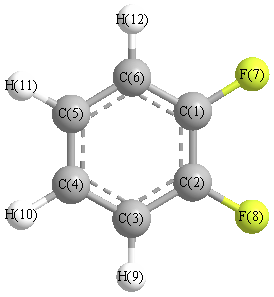
picture of orthodifluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.442 C1 C2 F8 119.833
C1 C6 C5 120.319 C1 C6 H12 118.602
C2 C1 C6 119.442 C2 C1 F7 119.833
C2 C3 C4 120.319 C2 C3 H9 118.602
C3 C2 F8 120.725 C3 C4 C5 120.239
C3 C4 H10 119.704 C4 C3 H9 121.079
C4 C5 C6 120.239 C4 C5 H11 120.057
C5 C4 H10 120.057 C5 C6 H12 121.079
C6 C1 F7 120.725 C6 C5 H11 119.704
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability