return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (benzocyclobutadiene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-303.074503
Energy at 298.15K-303.079835
HF Energy-302.577040
Nuclear repulsion energy307.103695
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3550 3156 0.95      
2 A1 3524 3132 1.99      
3 A1 3514 3123 0.66      
4 A1 1809 1608 0.10      
5 A1 1614 1435 0.77      
6 A1 1582 1407 1.48      
7 A1 1528 1358 10.58      
8 A1 1264 1124 0.34      
9 A1 1163 1034 2.13      
10 A1 1135 1009 3.28      
11 A1 1004 893 0.10      
12 A1 858 763 0.05      
13 A1 571 508 0.06      
14 A2 1055 938 0.00      
15 A2 967 860 0.00      
16 A2 936 832 0.00      
17 A2 803 714 0.00      
18 A2 613 545 0.00      
19 A2 309 275 0.00      
20 B1 984 875 3.29      
21 B1 807 717 27.58      
22 B1 766 681 5.83      
23 B1 373 332 0.54      
24 B1 248 220 3.31      
25 B2 3519 3128 2.11      
26 B2 3515 3124 3.87      
27 B2 3503 3114 3.14      
28 B2 1768 1571 1.88      
29 B2 1572 1397 3.41      
30 B2 1411 1254 1.75      
31 B2 1355 1204 8.70      
32 B2 1164 1035 0.16      
33 B2 1121 997 3.74      
34 B2 943 839 0.03      
35 B2 699 622 0.27      
36 B2 449 399 0.30      

Unscaled Zero Point Vibrational Energy (zpe) 25997.8 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 23109.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.15506 0.07100 0.04870

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.473 -0.631
C2 0.000 0.697 -1.883
C3 0.000 -0.697 -1.883
C4 0.000 -1.473 -0.631
C5 0.000 -0.689 2.087
C6 0.000 0.689 2.087
C7 0.000 0.734 0.533
C8 0.000 -0.734 0.533
H9 0.000 2.574 -0.665
H10 0.000 1.242 -2.840
H11 0.000 -1.242 -2.840
H12 0.000 -2.574 -0.665
H13 0.000 -1.472 2.866
H14 0.000 1.472 2.866

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.47332.50602.94673.47282.82871.37862.49531.10142.22133.50034.04764.57183.4970
C21.47331.39472.50604.20513.97012.41632.80822.23751.10092.16233.49105.22124.8121
C32.50601.39471.47333.97014.20512.80822.41633.49102.16231.10092.23754.81215.2212
C42.94672.50601.47332.82873.47282.49531.37864.04763.50032.22131.10143.49704.5718
C53.47284.20513.97012.82871.37752.10701.55474.26845.29164.95793.33581.10462.2966
C62.82873.97014.20513.47281.37751.55472.10703.33584.95795.29164.26842.29661.1046
C71.37862.41632.80822.49532.10701.55471.46812.19563.41093.90903.51843.21092.4471
C82.49532.80822.41631.37861.55472.10701.46813.51843.90903.41092.19562.44713.2109
H91.10142.23753.49104.04764.26843.33582.19563.51842.55084.39225.14835.37003.6993
H102.22131.10092.16233.50035.29164.95793.41093.90902.55082.48344.39226.31855.7109
H113.50032.16231.10092.22134.95795.29163.90903.41094.39222.48342.55085.71096.3185
H124.04763.49102.23751.10143.33584.26843.51842.19565.14834.39222.55083.69935.3700
H134.57185.22124.81213.49701.10462.29663.21092.44715.37006.31855.71093.69932.9432
H143.49704.81215.22124.57182.29661.10462.44713.21093.69935.71096.31855.37002.9432

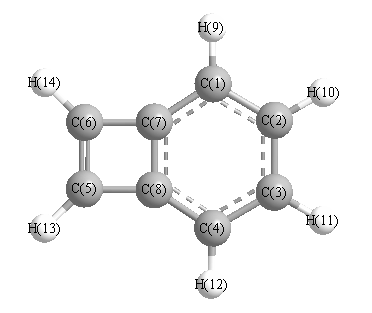
picture of benzocyclobutadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.784 C1 C2 H10 118.582
C1 C7 C6 149.243 C1 C7 N8 122.428
C2 C1 C7 115.788 C2 C1 H9 120.007
C2 C3 C4 121.784 C2 C3 H11 119.635
C3 C2 H10 119.635 C3 C4 N8 115.788
C3 C4 H12 120.007 C4 C3 H11 118.582
C4 N8 C5 149.243 C4 N8 C7 122.428
C5 C6 C7 91.671 C5 C6 H14 135.132
C5 N8 C7 88.329 C6 C5 N8 91.671
C6 C5 H13 135.132 C6 C7 N8 88.329
C7 C1 H9 124.204 C7 C6 H14 133.197
N8 C4 H12 124.204 N8 C5 H13 133.197
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability