return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHFCH3 (2-Fluoropropane)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-214.511740
Energy at 298.15K-214.519324
HF Energy-214.335199
Nuclear repulsion energy129.661091
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3563 3167 2.98      
2 A' 3560 3165 2.99      
3 A' 3377 3001 0.63      
4 A' 3361 2988 15.59      
5 A' 1736 1544 1.29      
6 A' 1724 1532 1.27      
7 A' 1613 1434 2.86      
8 A' 1551 1379 21.63      
9 A' 1342 1193 6.66      
10 A' 1297 1153 10.20      
11 A' 1092 970 0.49      
12 A' 902 802 0.03      
13 A' 493 438 0.65      
14 A' 357 317 0.18      
15 A' 216 192 0.02      
16 A" 3562 3166 0.20      
17 A" 3559 3164 0.96      
18 A" 3377 3002 1.43      
19 A" 1726 1534 0.00      
20 A" 1720 1529 0.05      
21 A" 1603 1425 0.98      
22 A" 1516 1348 21.53      
23 A" 1247 1108 9.98      
24 A" 1054 937 0.11      
25 A" 1029 914 0.19      
26 A" 407 362 2.22      
27 A" 187 166 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 23584.2 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 20964.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.27249 0.26004 0.15129

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.268 0.263 0.000
F2 -0.849 1.117 0.000
H3 1.198 0.886 0.000
C4 0.268 -0.623 1.298
C5 0.268 -0.623 -1.298
H6 1.170 -1.257 1.323
H7 1.170 -1.257 -1.323
H8 0.261 0.015 2.197
H9 0.261 0.015 -2.197
H10 -0.622 -1.274 1.321
H11 -0.622 -1.274 -1.321

Atom - Atom Distances (Å)
  C1 F2 H3 C4 C5 H6 H7 H8 H9 H10 H11
C11.40671.11991.57131.57132.20742.20742.21102.21102.21322.2132
F21.40672.06062.44172.44173.38583.38582.69692.69692.74052.7405
H31.11992.06062.19742.19742.51922.51922.54272.54273.11863.1186
C41.57132.44172.19742.59551.10252.84271.10313.55281.10302.8416
C51.57132.44172.19742.59552.84271.10253.55281.10312.84161.1030
H62.20743.38582.51921.10252.84272.64511.79203.85161.79243.1940
H72.20743.38582.51922.84271.10252.64513.85161.79203.19401.7924
H82.21102.69692.54271.10313.55281.79203.85164.39421.79123.8496
H92.21102.69692.54273.55281.10313.85161.79204.39423.84961.7912
H102.21322.74053.11861.10302.84161.79243.19401.79123.84962.6422
H112.21322.74053.11862.84161.10303.19401.79243.84961.79122.6422

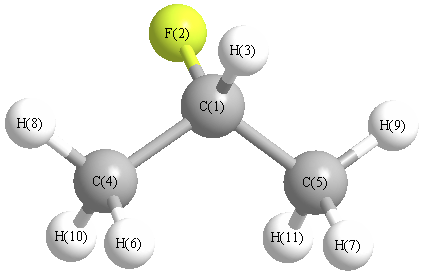
picture of 2-Fluoropropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H8 110.296 C1 C4 H10 110.475
C1 C5 H7 110.051 C1 C5 H9 110.296
C1 C5 H11 110.475 F2 C1 H3 108.754
F2 C1 C4 110.022 F2 C1 C5 110.022
H3 C1 C4 108.307 H3 C1 C5 108.307
C4 C1 C5 111.357 H7 C5 H9 108.675
H7 C5 H11 108.723 H8 C4 H10 108.572
H9 C5 H11 108.572
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability