Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -214.507800 |
| Energy at 298.15K | -214.515492 |
| HF Energy | -214.331099 |
| Nuclear repulsion energy | 125.426728 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3557 |
3161 |
2.57 |
|
|
|
| 2 |
A' |
3410 |
3031 |
3.78 |
|
|
|
| 3 |
A' |
3376 |
3001 |
2.99 |
|
|
|
| 4 |
A' |
3333 |
2963 |
12.30 |
|
|
|
| 5 |
A' |
1744 |
1550 |
0.59 |
|
|
|
| 6 |
A' |
1727 |
1535 |
0.61 |
|
|
|
| 7 |
A' |
1711 |
1521 |
0.16 |
|
|
|
| 8 |
A' |
1625 |
1445 |
1.76 |
|
|
|
| 9 |
A' |
1602 |
1424 |
5.55 |
|
|
|
| 10 |
A' |
1481 |
1316 |
17.45 |
|
|
|
| 11 |
A' |
1275 |
1133 |
4.44 |
|
|
|
| 12 |
A' |
1226 |
1090 |
2.32 |
|
|
|
| 13 |
A' |
1159 |
1030 |
0.36 |
|
|
|
| 14 |
A' |
992 |
882 |
4.75 |
|
|
|
| 15 |
A' |
460 |
409 |
1.48 |
|
|
|
| 16 |
A' |
284 |
253 |
0.86 |
|
|
|
| 17 |
A" |
3555 |
3160 |
4.36 |
|
|
|
| 18 |
A" |
3524 |
3132 |
3.01 |
|
|
|
| 19 |
A" |
3440 |
3058 |
12.45 |
|
|
|
| 20 |
A" |
1736 |
1543 |
1.81 |
|
|
|
| 21 |
A" |
1455 |
1293 |
0.00 |
|
|
|
| 22 |
A" |
1405 |
1249 |
1.09 |
|
|
|
| 23 |
A" |
1288 |
1145 |
3.02 |
|
|
|
| 24 |
A" |
988 |
879 |
1.84 |
|
|
|
| 25 |
A" |
819 |
728 |
1.35 |
|
|
|
| 26 |
A" |
233 |
207 |
0.00 |
|
|
|
| 27 |
A" |
112 |
99 |
0.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23757.8 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 21118.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.144 |
-0.804 |
0.000 |
| C2 |
0.000 |
0.762 |
0.000 |
| C3 |
-1.495 |
1.198 |
0.000 |
| F4 |
1.492 |
-1.195 |
0.000 |
| H5 |
-0.370 |
-1.221 |
0.900 |
| H6 |
-0.370 |
-1.221 |
-0.900 |
| H7 |
0.511 |
1.170 |
-0.892 |
| H8 |
0.511 |
1.170 |
0.892 |
| H9 |
-1.578 |
2.298 |
0.000 |
| H10 |
-2.015 |
0.812 |
-0.894 |
| H11 |
-2.015 |
0.812 |
0.894 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5721 | 2.5875 | 1.4038 | 1.1174 | 1.1174 | 2.1970 | 2.1970 | 3.5478 | 2.8411 | 2.8411 |
C2 | 1.5721 | | 1.5576 | 2.4611 | 2.2087 | 2.2087 | 1.1062 | 1.1062 | 2.2023 | 2.2046 | 2.2046 | C3 | 2.5875 | 1.5576 | | 3.8280 | 2.8154 | 2.8154 | 2.1959 | 2.1959 | 1.1031 | 1.1034 | 1.1034 | F4 | 1.4038 | 2.4611 | 3.8280 | | 2.0689 | 2.0689 | 2.7118 | 2.7118 | 4.6508 | 4.1387 | 4.1387 | H5 | 1.1174 | 2.2087 | 2.8154 | 2.0689 | | 1.7992 | 3.1154 | 2.5488 | 3.8278 | 3.1710 | 2.6150 | H6 | 1.1174 | 2.2087 | 2.8154 | 2.0689 | 1.7992 | | 2.5488 | 3.1154 | 3.8278 | 2.6150 | 3.1710 | H7 | 2.1970 | 1.1062 | 2.1959 | 2.7118 | 3.1154 | 2.5488 | | 1.7839 | 2.5361 | 2.5510 | 3.1140 | H8 | 2.1970 | 1.1062 | 2.1959 | 2.7118 | 2.5488 | 3.1154 | 1.7839 | | 2.5361 | 3.1140 | 2.5510 | H9 | 3.5478 | 2.2023 | 1.1031 | 4.6508 | 3.8278 | 3.8278 | 2.5361 | 2.5361 | | 1.7880 | 1.7880 | H10 | 2.8411 | 2.2046 | 1.1034 | 4.1387 | 3.1710 | 2.6150 | 2.5510 | 3.1140 | 1.7880 | | 1.7879 | H11 | 2.8411 | 2.2046 | 1.1034 | 4.1387 | 2.6150 | 3.1710 | 3.1140 | 2.5510 | 1.7880 | 1.7879 | |
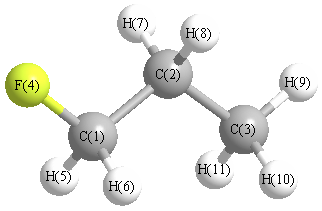 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.532 |
|
C1 |
C2 |
H7 |
108.983 |
| C1 |
C2 |
H8 |
108.983 |
|
C2 |
C1 |
F4 |
111.462 |
| C2 |
C1 |
H5 |
109.251 |
|
C2 |
C1 |
H6 |
109.251 |
| C2 |
C3 |
H9 |
110.567 |
|
C2 |
C3 |
H10 |
110.729 |
| C2 |
C3 |
H11 |
110.729 |
|
C3 |
C2 |
H7 |
109.887 |
| C3 |
C2 |
H8 |
109.887 |
|
F4 |
C1 |
H5 |
109.769 |
| F4 |
C1 |
H6 |
109.769 |
|
H5 |
C1 |
H6 |
107.241 |
| H7 |
C2 |
H8 |
107.473 |
|
H9 |
C3 |
H10 |
108.251 |
| H9 |
C3 |
H11 |
108.251 |
|
H10 |
C3 |
H11 |
108.222 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -214.507742 |
| Energy at 298.15K | |
| HF Energy | -214.331105 |
| Nuclear repulsion energy | 127.753515 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3558 |
3163 |
2.53 |
|
|
|
| 2 |
A |
3556 |
3161 |
3.68 |
|
|
|
| 3 |
A |
3524 |
3132 |
2.47 |
|
|
|
| 4 |
A |
3440 |
3058 |
14.07 |
|
|
|
| 5 |
A |
3409 |
3031 |
2.28 |
|
|
|
| 6 |
A |
3376 |
3001 |
3.02 |
|
|
|
| 7 |
A |
3333 |
2962 |
12.80 |
|
|
|
| 8 |
A |
1741 |
1548 |
1.06 |
|
|
|
| 9 |
A |
1734 |
1541 |
1.49 |
|
|
|
| 10 |
A |
1720 |
1529 |
0.97 |
|
|
|
| 11 |
A |
1705 |
1516 |
0.12 |
|
|
|
| 12 |
A |
1625 |
1444 |
0.34 |
|
|
|
| 13 |
A |
1583 |
1407 |
14.64 |
|
|
|
| 14 |
A |
1543 |
1372 |
10.52 |
|
|
|
| 15 |
A |
1436 |
1276 |
0.96 |
|
|
|
| 16 |
A |
1392 |
1238 |
2.38 |
|
|
|
| 17 |
A |
1291 |
1147 |
4.52 |
|
|
|
| 18 |
A |
1269 |
1128 |
2.22 |
|
|
|
| 19 |
A |
1215 |
1080 |
4.79 |
|
|
|
| 20 |
A |
1112 |
989 |
2.30 |
|
|
|
| 21 |
A |
1015 |
902 |
1.90 |
|
|
|
| 22 |
A |
953 |
847 |
1.55 |
|
|
|
| 23 |
A |
851 |
756 |
1.85 |
|
|
|
| 24 |
A |
497 |
441 |
0.94 |
|
|
|
| 25 |
A |
310 |
276 |
0.47 |
|
|
|
| 26 |
A |
210 |
186 |
0.33 |
|
|
|
| 27 |
A |
119 |
106 |
0.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23757.5 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 21118.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.823 |
0.510 |
0.298 |
| C2 |
-0.627 |
0.674 |
-0.288 |
| C3 |
-1.560 |
-0.504 |
0.122 |
| F4 |
1.439 |
-0.659 |
-0.175 |
| H5 |
1.437 |
1.401 |
0.015 |
| H6 |
0.769 |
0.483 |
1.413 |
| H7 |
-0.563 |
0.733 |
-1.391 |
| H8 |
-1.042 |
1.632 |
0.075 |
| H9 |
-2.561 |
-0.382 |
-0.325 |
| H10 |
-1.140 |
-1.465 |
-0.221 |
| H11 |
-1.672 |
-0.549 |
1.219 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5722 | 2.5955 | 1.4038 | 1.1175 | 1.1174 | 2.1963 | 2.1880 | 3.5543 | 2.8331 | 2.8633 |
C2 | 1.5722 | | 1.5578 | 2.4612 | 2.2085 | 2.2087 | 1.1063 | 1.1060 | 2.2038 | 2.2010 | 2.2047 | C3 | 2.5955 | 1.5578 | | 3.0173 | 3.5520 | 2.8392 | 2.1938 | 2.1988 | 1.1032 | 1.1033 | 1.1034 | F4 | 1.4038 | 2.4612 | 3.0173 | | 2.0686 | 2.0681 | 2.7253 | 3.3868 | 4.0121 | 2.7028 | 3.4113 | H5 | 1.1175 | 2.2085 | 3.5520 | 2.0686 | | 1.8008 | 2.5347 | 2.4902 | 4.3900 | 3.8613 | 3.8621 | H6 | 1.1174 | 2.2087 | 2.8392 | 2.0681 | 1.8008 | | 3.1148 | 2.5278 | 3.8539 | 3.1796 | 2.6572 | H7 | 2.1963 | 1.1063 | 2.1938 | 2.7253 | 2.5347 | 3.1148 | | 1.7857 | 2.5237 | 2.5559 | 3.1121 | H8 | 2.1880 | 1.1060 | 2.1988 | 3.3868 | 2.4902 | 2.5278 | 1.7857 | | 2.5543 | 3.1133 | 2.5423 | H9 | 3.5543 | 2.2038 | 1.1032 | 4.0121 | 4.3900 | 3.8539 | 2.5237 | 2.5543 | | 1.7891 | 1.7888 | H10 | 2.8331 | 2.2010 | 1.1033 | 2.7028 | 3.8613 | 3.1796 | 2.5559 | 3.1133 | 1.7891 | | 1.7879 | H11 | 2.8633 | 2.2047 | 1.1034 | 3.4113 | 3.8621 | 2.6572 | 3.1121 | 2.5423 | 1.7888 | 1.7879 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.041 |
|
C1 |
C2 |
H7 |
108.926 |
| C1 |
C2 |
H8 |
108.305 |
|
C2 |
C1 |
F4 |
111.466 |
| C2 |
C1 |
H5 |
109.228 |
|
C2 |
C1 |
H6 |
109.244 |
| C2 |
C3 |
H9 |
110.667 |
|
C2 |
C3 |
H10 |
110.436 |
| C2 |
C3 |
H11 |
110.722 |
|
C3 |
C2 |
H7 |
109.704 |
| C3 |
C2 |
H8 |
110.111 |
|
F4 |
C1 |
H5 |
109.744 |
| F4 |
C1 |
H6 |
109.699 |
|
H5 |
C1 |
H6 |
107.367 |
| H7 |
C2 |
H8 |
107.639 |
|
H9 |
C3 |
H10 |
108.363 |
| H9 |
C3 |
H11 |
108.327 |
|
H10 |
C3 |
H11 |
108.242 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
