return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C7H10 (Norbornene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-268.044840
Energy at 298.15K-268.056888
HF Energy-267.653096
Nuclear repulsion energy303.141136
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3551 3157 0.15      
2 A' 3539 3146 5.81      
3 A' 3531 3138 5.91      
4 A' 3478 3092 1.20      
5 A' 3412 3033 5.21      
6 A' 3409 3030 2.98      
7 A' 1736 1543 0.16      
8 A' 1722 1531 0.32      
9 A' 1703 1514 0.50      
10 A' 1500 1333 7.30      
11 A' 1469 1306 1.80      
12 A' 1367 1215 1.20      
13 A' 1261 1121 1.09      
14 A' 1201 1068 0.55      
15 A' 1165 1035 0.73      
16 A' 1086 966 0.08      
17 A' 1056 939 0.09      
18 A' 1020 906 0.27      
19 A' 986 877 0.67      
20 A' 896 797 0.48      
21 A' 840 747 1.28      
22 A' 776 689 13.09      
23 A' 490 436 0.56      
24 A' 407 362 0.42      
25 A" 3526 3134 0.01      
26 A" 3521 3130 2.95      
27 A" 3474 3088 6.63      
28 A" 3406 3027 3.53      
29 A" 1712 1522 0.34      
30 A" 1526 1356 0.72      
31 A" 1490 1324 0.15      
32 A" 1435 1275 0.00      
33 A" 1414 1257 0.05      
34 A" 1396 1241 0.00      
35 A" 1353 1203 0.01      
36 A" 1252 1113 0.02      
37 A" 1166 1036 0.30      
38 A" 1077 957 0.03      
39 A" 1055 938 0.19      
40 A" 1005 893 0.65      
41 A" 935 831 0.09      
42 A" 877 779 0.12      
43 A" 715 636 0.16      
44 A" 513 456 0.03      
45 A" 262 233 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 37355.3 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 33205.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.12780 0.11058 0.09711

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.149 0.800 0.000
C2 0.226 0.248 1.144
C3 0.226 0.248 -1.144
C4 -1.145 0.806 0.678
C5 -1.145 0.806 -0.678
C6 0.226 -1.297 0.790
C7 0.226 -1.297 -0.790
H8 1.223 1.902 0.000
H9 2.162 0.355 0.000
H10 0.515 0.478 2.186
H11 0.515 0.478 -2.186
H12 -1.975 1.061 1.350
H13 -1.975 1.061 -1.350
H14 1.135 -1.786 1.186
H15 1.135 -1.786 -1.186
H16 -0.654 -1.817 1.211
H17 -0.654 -1.817 -1.211

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.57041.57042.39172.39172.42382.42381.10501.10592.29922.29923.41343.41342.84472.84473.40073.4007
C21.57042.28811.55122.34691.58492.47552.24522.25121.10593.35092.35523.42422.22793.22372.24513.2531
C31.57042.28812.34691.55122.47551.58492.24522.25123.35091.10593.42422.35523.22372.22793.25312.2451
C42.39171.55122.34691.35572.51252.90782.69583.40492.26633.32631.09852.20633.48873.92272.72063.2689
C52.39172.34691.55121.35572.90782.51252.69583.40493.32632.26632.20631.09853.92273.48873.26892.7206
C62.42381.58492.47552.51252.90781.58033.44332.66492.27703.47783.27373.87101.10572.22991.10512.2470
C72.42382.47551.58492.90782.51251.58033.44332.66493.47782.27703.87103.27372.22991.10572.24701.1051
H81.10502.24522.24522.69582.69583.44333.44331.80982.70382.70383.57193.57193.87503.87504.33844.3384
H91.10592.25122.25123.40493.40492.66492.66491.80982.74002.74004.40834.40832.65392.65393.75663.7566
H102.29921.10593.35092.26633.32632.27703.47782.70382.74004.37282.69034.36422.55134.10912.75384.2632
H112.29923.35091.10593.32632.26633.47782.27702.70382.74004.37284.36422.69034.10912.55134.26322.7538
H123.41342.35523.42421.09852.20633.27373.87103.57194.40832.69034.36422.70034.21934.92033.16914.0724
H133.41343.42422.35522.20631.09853.87103.27373.57194.40834.36422.69032.70034.92034.21934.07243.1691
H142.84472.22793.22373.48873.92271.10572.22993.87502.65392.55134.10914.21934.92032.37271.78992.9918
H152.84473.22372.22793.92273.48872.22991.10573.87502.65394.10912.55134.92034.21932.37272.99181.7899
H163.40072.24513.25312.72063.26891.10512.24704.33843.75662.75384.26323.16914.07241.78992.99182.4222
H173.40073.25312.24513.26892.72062.24701.10514.33843.75664.26322.75384.07243.16912.99181.78992.4222

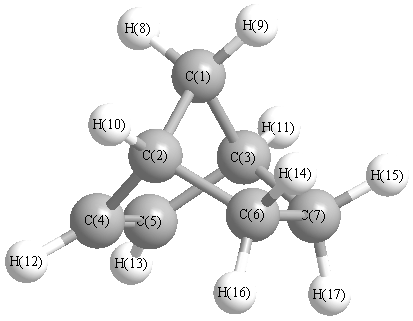
picture of Norbornene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 100.019 C1 C2 C6 100.373
C1 C2 H10 117.369 C1 C3 C5 100.019
C1 C3 C7 100.373 C1 C3 H11 117.369
C2 C1 C3 93.518 C2 C1 H8 112.962
C2 C1 H9 113.385 C2 C4 C5 107.489
C2 C4 H12 124.569 C2 C6 C7 102.901
C2 C6 H14 110.521 C2 C6 H16 111.907
C3 C1 H8 112.962 C3 C1 H9 113.385
C3 C5 C4 107.489 C3 C5 H13 124.569
C3 C7 C6 102.901 C3 C7 H15 110.521
C3 C7 H17 111.907 C4 C2 C6 106.473
C4 C2 H10 116.052 C4 C5 H13 127.734
C5 C3 C7 106.473 C5 C3 H11 116.052
C5 C4 H12 127.734 C6 C2 H10 114.420
C6 C7 H15 110.996 C6 C7 H17 112.389
C7 C3 H11 114.420 C7 C6 H14 110.996
C7 C6 H16 112.389 H8 C1 H9 109.882
H14 C6 H16 108.111 H15 C7 H17 108.111
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability