Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -130.933927 |
| Energy at 298.15K | -130.937953 |
| HF Energy | -130.810198 |
| Nuclear repulsion energy | 63.283662 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3650 |
3244 |
0.13 |
|
|
|
| 2 |
A' |
3584 |
3186 |
22.05 |
|
|
|
| 3 |
A' |
3482 |
3095 |
4.87 |
|
|
|
| 4 |
A' |
3468 |
3083 |
7.36 |
|
|
|
| 5 |
A' |
1659 |
1475 |
1.96 |
|
|
|
| 6 |
A' |
1573 |
1398 |
7.87 |
|
|
|
| 7 |
A' |
1458 |
1296 |
7.64 |
|
|
|
| 8 |
A' |
1245 |
1107 |
15.64 |
|
|
|
| 9 |
A' |
1124 |
999 |
0.91 |
|
|
|
| 10 |
A' |
999 |
888 |
0.80 |
|
|
|
| 11 |
A' |
475 |
423 |
5.66 |
|
|
|
| 12 |
A" |
1083 |
962 |
0.63 |
|
|
|
| 13 |
A" |
850 |
756 |
8.04 |
|
|
|
| 14 |
A" |
704 |
626 |
35.26 |
|
|
|
| 15 |
A" |
596 |
530 |
24.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12974.2 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 11532.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.428 |
0.000 |
| C2 |
1.188 |
-0.292 |
0.000 |
| N3 |
-1.224 |
-0.245 |
0.000 |
| H4 |
0.053 |
1.533 |
0.000 |
| H5 |
2.163 |
0.210 |
0.000 |
| H6 |
1.177 |
-1.389 |
0.000 |
| H7 |
-1.957 |
0.541 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3897 | 1.3966 | 1.1055 | 2.1744 | 2.1649 | 1.9605 |
C2 | 1.3897 | | 2.4127 | 2.1492 | 1.0966 | 1.0970 | 3.2541 | N3 | 1.3966 | 2.4127 | | 2.1881 | 3.4175 | 2.6591 | 1.0745 | H4 | 1.1055 | 2.1492 | 2.1881 | | 2.4909 | 3.1302 | 2.2416 | H5 | 2.1744 | 1.0966 | 3.4175 | 2.4909 | | 1.8789 | 4.1340 | H6 | 2.1649 | 1.0970 | 2.6591 | 3.1302 | 1.8789 | | 3.6802 | H7 | 1.9605 | 3.2541 | 1.0745 | 2.2416 | 4.1340 | 3.6802 | |
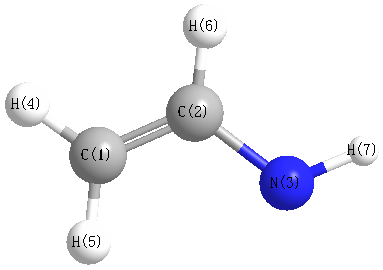 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.537 |
|
C1 |
C2 |
H6 |
120.600 |
| C1 |
N3 |
H7 |
104.250 |
|
C2 |
C1 |
N3 |
119.971 |
| C2 |
C1 |
H4 |
118.487 |
|
N3 |
C1 |
H4 |
121.542 |
| H5 |
C2 |
H6 |
117.863 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -130.935357 |
| Energy at 298.15K | -130.939417 |
| HF Energy | -130.810706 |
| Nuclear repulsion energy | 63.383720 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3638 |
3234 |
0.14 |
|
|
|
| 2 |
A' |
3561 |
3165 |
31.03 |
|
|
|
| 3 |
A' |
3517 |
3126 |
0.26 |
|
|
|
| 4 |
A' |
3467 |
3082 |
7.16 |
|
|
|
| 5 |
A' |
1655 |
1471 |
0.84 |
|
|
|
| 6 |
A' |
1577 |
1402 |
7.35 |
|
|
|
| 7 |
A' |
1463 |
1300 |
12.82 |
|
|
|
| 8 |
A' |
1248 |
1110 |
12.83 |
|
|
|
| 9 |
A' |
1100 |
978 |
1.60 |
|
|
|
| 10 |
A' |
1031 |
916 |
5.40 |
|
|
|
| 11 |
A' |
498 |
442 |
6.30 |
|
|
|
| 12 |
A" |
1126 |
1001 |
14.83 |
|
|
|
| 13 |
A" |
849 |
754 |
16.23 |
|
|
|
| 14 |
A" |
731 |
650 |
18.82 |
|
|
|
| 15 |
A" |
586 |
521 |
8.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13023.1 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 11576.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.461 |
0.000 |
| C2 |
1.161 |
-0.299 |
0.000 |
| N3 |
-1.307 |
-0.029 |
0.000 |
| H4 |
0.085 |
1.561 |
0.000 |
| H5 |
2.154 |
0.166 |
0.000 |
| H6 |
1.120 |
-1.397 |
0.000 |
| H7 |
-1.178 |
-1.098 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3881 | 1.3960 | 1.1029 | 2.1741 | 2.1695 | 1.9538 |
C2 | 1.3881 | | 2.4831 | 2.1492 | 1.0964 | 1.0982 | 2.4715 | N3 | 1.3960 | 2.4831 | | 2.1130 | 3.4666 | 2.7862 | 1.0767 | H4 | 1.1029 | 2.1492 | 2.1130 | | 2.4953 | 3.1336 | 2.9432 | H5 | 2.1741 | 1.0964 | 3.4666 | 2.4953 | | 1.8738 | 3.5633 | H6 | 2.1695 | 1.0982 | 2.7862 | 3.1336 | 1.8738 | | 2.3172 | H7 | 1.9538 | 2.4715 | 1.0767 | 2.9432 | 3.5633 | 2.3172 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.657 |
|
C1 |
C2 |
H6 |
121.079 |
| C1 |
N3 |
H7 |
103.648 |
|
C2 |
C1 |
N3 |
126.226 |
| C2 |
C1 |
H4 |
118.812 |
|
N3 |
C1 |
H4 |
114.962 |
| H5 |
C2 |
H6 |
117.264 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
