return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CCH (phenylacetylene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-303.118644
Energy at 298.15K-303.123201
HF Energy-302.608041
Nuclear repulsion energy293.574154
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3729 3314 123.80      
2 A 3530 3137 1.43      
3 A 3517 3127 3.16      
4 A 3506 3117 0.24      
5 A 2259 2008 24.44      
6 A 1739 1546 4.03      
7 A 1630 1449 16.78      
8 A 1284 1141 5.99      
9 A 1272 1131 0.11      
10 A 1106 983 0.00      
11 A 1067 948 1.07      
12 A 800 711 0.95      
13 A 483 429 0.09      
14 A 1059 942 0.00      
15 A 929 825 0.00      
16 A 435 386 0.00      
17 A 1076 956 0.01      
18 A 997 886 0.91      
19 A 817 726 14.30      
20 A 756 672 5.57      
21 A 722 642 12.55      
22 A 571 508 2.09      
23 A 376 334 0.38      
24 A 145 129 0.44      
25 A 3523 3131 6.80      
26 A 3511 3121 0.11      
27 A 1713 1523 0.81      
28 A 1573 1398 8.14      
29 A 1432 1273 0.56      
30 A 1290 1147 0.06      
31 A 1210 1076 0.02      
32 A 1159 1031 0.42      
33 A 748 665 12.21      
34 A 662 588 0.05      
35 A 542 482 1.04      
36 A 158 140 0.31      

Unscaled Zero Point Vibrational Energy (zpe) 25661.8 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 22810.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.18410 0.04901 0.03870

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.597
C2 0.000 0.000 2.072
C3 0.000 1.230 -0.126
C4 0.000 -1.230 -0.126
C5 0.000 1.226 -1.540
C6 0.000 -1.226 -1.540
C7 0.000 0.000 -2.250
C8 0.000 0.000 3.296
H9 0.000 0.000 4.377
H10 0.000 2.183 0.426
H11 0.000 -2.183 0.426
H12 0.000 2.181 -2.089
H13 0.000 -2.181 -2.089
H14 0.000 0.000 -3.351

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.47571.42641.42642.46352.46352.84672.69963.78032.18982.18983.45943.45943.9477
C21.47572.51872.51873.81483.81484.32241.22392.30462.73432.73434.69804.69805.4234
C31.42642.51872.46001.41462.83422.45493.63624.66751.10123.45742.18143.93533.4521
C41.42642.51872.46002.83421.41462.45493.63624.66753.45741.10123.93532.18143.4521
C52.46353.81481.41462.83422.45181.41664.98946.04272.18683.93541.10113.45042.1868
C62.46353.81482.83421.41462.45181.41664.98946.04273.93542.18683.45041.10112.1868
C72.84674.32242.45492.45491.41661.41665.54636.62703.45363.45362.18652.18651.1010
C82.69961.22393.63623.63624.98944.98945.54631.08073.60623.60625.80995.80996.6473
H93.78032.30464.66754.66756.04276.04276.62701.08074.51404.51406.82376.82377.7280
H102.18982.73431.10123.45742.18683.93543.45363.60624.51404.36632.51495.03654.3626
H112.18982.73433.45741.10123.93542.18683.45363.60624.51404.36635.03652.51494.3626
H123.45944.69802.18143.93531.10113.45042.18655.80996.82372.51495.03654.36112.5195
H133.45944.69803.93532.18143.45041.10112.18655.80996.82375.03652.51494.36112.5195
H143.94775.42343.45213.45212.18682.18681.10106.64737.72804.36264.36262.51952.5195

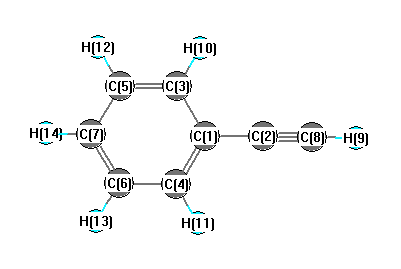
picture of phenylacetylene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C8 180.000 C1 C3 C5 120.255
C1 C3 H10 119.523 C1 C4 C6 120.255
C1 C4 H11 119.523 C2 C1 C3 120.421
C2 C1 C4 120.421 C2 C8 H9 180.000
C3 C1 C4 119.158 C3 C5 C7 120.240
C3 C5 H12 119.728 C4 C6 C7 120.240
C4 C6 H13 119.728 C5 C3 H10 120.222
C5 C7 C6 119.851 C5 C7 H14 120.074
C6 C4 H11 120.222 C6 C7 H14 120.074
C7 C5 H12 120.033 C7 C6 H13 120.033
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability