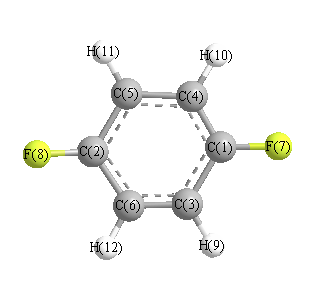
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3528 |
3136 |
0.00 |
132.55 |
0.19 |
0.31 |
| 2 |
Ag |
1774 |
1577 |
0.00 |
23.05 |
0.71 |
0.83 |
| 3 |
Ag |
1435 |
1275 |
0.00 |
7.94 |
0.20 |
0.34 |
| 4 |
Ag |
1242 |
1104 |
0.00 |
10.65 |
0.67 |
0.80 |
| 5 |
Ag |
895 |
796 |
0.00 |
21.94 |
0.10 |
0.18 |
| 6 |
Ag |
456 |
406 |
0.00 |
2.74 |
0.48 |
0.65 |
| 7 |
Au |
1044 |
928 |
0.00 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
442 |
393 |
0.00 |
0.00 |
0.72 |
0.84 |
| 9 |
B1g |
902 |
802 |
0.00 |
5.75 |
0.75 |
0.86 |
| 10 |
B1u |
3514 |
3123 |
1.63 |
0.00 |
0.00 |
0.00 |
| 11 |
B1u |
1677 |
1490 |
142.72 |
0.00 |
0.00 |
0.00 |
| 12 |
B1u |
1410 |
1253 |
33.40 |
0.00 |
0.00 |
0.00 |
| 13 |
B1u |
1073 |
954 |
1.85 |
0.00 |
0.00 |
0.00 |
| 14 |
B1u |
785 |
698 |
4.62 |
0.00 |
0.00 |
0.00 |
| 15 |
B2g |
1016 |
903 |
0.00 |
2.80 |
0.75 |
0.86 |
| 16 |
B2g |
752 |
668 |
0.00 |
0.00 |
0.75 |
0.86 |
| 17 |
B2g |
389 |
346 |
0.00 |
1.31 |
0.75 |
0.86 |
| 18 |
B2u |
3526 |
3134 |
16.43 |
0.00 |
0.00 |
0.00 |
| 19 |
B2u |
1520 |
1351 |
0.03 |
0.00 |
0.00 |
0.00 |
| 20 |
B2u |
1216 |
1081 |
8.23 |
0.00 |
0.00 |
0.00 |
| 21 |
B2u |
1166 |
1037 |
4.59 |
0.00 |
0.00 |
0.00 |
| 22 |
B2u |
340 |
302 |
2.64 |
0.00 |
0.00 |
0.00 |
| 23 |
B3g |
3515 |
3125 |
0.00 |
74.61 |
0.75 |
0.86 |
| 24 |
B3g |
1684 |
1497 |
0.00 |
6.13 |
0.75 |
0.86 |
| 25 |
B3g |
1391 |
1237 |
0.00 |
2.51 |
0.75 |
0.86 |
| 26 |
B3g |
676 |
601 |
0.00 |
5.75 |
0.75 |
0.86 |
| 27 |
B3g |
437 |
388 |
0.00 |
0.53 |
0.75 |
0.86 |
| 28 |
B3u |
876 |
778 |
36.43 |
0.00 |
0.00 |
0.00 |
| 29 |
B3u |
537 |
477 |
0.61 |
0.00 |
0.00 |
0.00 |
| 30 |
B3u |
164 |
146 |
0.53 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 19689.7 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 17502.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.