Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -571.055896 |
| Energy at 298.15K | -571.063442 |
| Nuclear repulsion energy | 155.303816 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3559 |
3164 |
1.56 |
|
|
|
| 2 |
A' |
3420 |
3040 |
1.83 |
|
|
|
| 3 |
A' |
3406 |
3028 |
0.86 |
|
|
|
| 4 |
A' |
3378 |
3003 |
2.31 |
|
|
|
| 5 |
A' |
1739 |
1546 |
0.79 |
|
|
|
| 6 |
A' |
1710 |
1520 |
0.20 |
|
|
|
| 7 |
A' |
1680 |
1493 |
0.89 |
|
|
|
| 8 |
A' |
1621 |
1441 |
0.92 |
|
|
|
| 9 |
A' |
1550 |
1378 |
2.02 |
|
|
|
| 10 |
A' |
1419 |
1261 |
34.04 |
|
|
|
| 11 |
A' |
1234 |
1097 |
0.02 |
|
|
|
| 12 |
A' |
1165 |
1036 |
0.81 |
|
|
|
| 13 |
A' |
1018 |
905 |
8.79 |
|
|
|
| 14 |
A' |
884 |
786 |
16.75 |
|
|
|
| 15 |
A' |
384 |
342 |
1.42 |
|
|
|
| 16 |
A' |
245 |
217 |
2.49 |
|
|
|
| 17 |
A" |
3559 |
3163 |
3.22 |
|
|
|
| 18 |
A" |
3536 |
3143 |
1.20 |
|
|
|
| 19 |
A" |
3525 |
3133 |
0.05 |
|
|
|
| 20 |
A" |
1732 |
1540 |
2.28 |
|
|
|
| 21 |
A" |
1453 |
1292 |
0.00 |
|
|
|
| 22 |
A" |
1385 |
1232 |
0.17 |
|
|
|
| 23 |
A" |
1207 |
1073 |
0.29 |
|
|
|
| 24 |
A" |
958 |
852 |
1.00 |
|
|
|
| 25 |
A" |
809 |
719 |
2.28 |
|
|
|
| 26 |
A" |
233 |
207 |
0.02 |
|
|
|
| 27 |
A" |
111 |
98 |
1.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23459.5 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 20853.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.602 |
0.000 |
| C2 |
0.921 |
-0.659 |
0.000 |
| C3 |
2.426 |
-0.247 |
0.000 |
| Cl4 |
-1.778 |
0.139 |
0.000 |
| H5 |
0.173 |
1.224 |
0.901 |
| H6 |
0.173 |
1.224 |
-0.901 |
| H7 |
0.697 |
-1.271 |
-0.893 |
| H8 |
0.697 |
-1.271 |
0.893 |
| H9 |
3.066 |
-1.145 |
0.000 |
| H10 |
2.668 |
0.351 |
-0.895 |
| H11 |
2.668 |
0.351 |
0.895 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5613 | 2.5700 | 1.8373 | 1.1086 | 1.1086 | 2.1891 | 2.1891 | 3.5289 | 2.8251 | 2.8251 |
C2 | 1.5613 | | 1.5598 | 2.8145 | 2.2168 | 2.2168 | 1.1059 | 1.1059 | 2.1992 | 2.2071 | 2.2071 | C3 | 2.5700 | 1.5598 | | 4.2213 | 2.8369 | 2.8369 | 2.1989 | 2.1989 | 1.1031 | 1.1033 | 1.1033 | Cl4 | 1.8373 | 2.8145 | 4.2213 | | 2.4077 | 2.4077 | 2.9850 | 2.9850 | 5.0112 | 4.5400 | 4.5400 | H5 | 1.1086 | 2.2168 | 2.8369 | 2.4077 | | 1.8029 | 3.1173 | 2.5491 | 3.8459 | 3.1952 | 2.6426 | H6 | 1.1086 | 2.2168 | 2.8369 | 2.4077 | 1.8029 | | 2.5491 | 3.1173 | 3.8459 | 2.6426 | 3.1952 | H7 | 2.1891 | 1.1059 | 2.1989 | 2.9850 | 3.1173 | 2.5491 | | 1.7860 | 2.5349 | 2.5530 | 3.1166 | H8 | 2.1891 | 1.1059 | 2.1989 | 2.9850 | 2.5491 | 3.1173 | 1.7860 | | 2.5349 | 3.1166 | 2.5530 | H9 | 3.5289 | 2.1992 | 1.1031 | 5.0112 | 3.8459 | 3.8459 | 2.5349 | 2.5349 | | 1.7885 | 1.7885 | H10 | 2.8251 | 2.2071 | 1.1033 | 4.5400 | 3.1952 | 2.6426 | 2.5530 | 3.1166 | 1.7885 | | 1.7893 | H11 | 2.8251 | 2.2071 | 1.1033 | 4.5400 | 2.6426 | 3.1952 | 3.1166 | 2.5530 | 1.7885 | 1.7893 | |
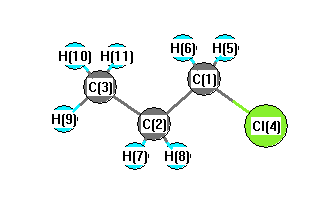 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
110.860 |
|
C1 |
C2 |
H7 |
109.122 |
| C1 |
C2 |
H8 |
109.122 |
|
C2 |
C1 |
Cl4 |
111.556 |
| C2 |
C1 |
H5 |
111.124 |
|
C2 |
C1 |
H6 |
111.124 |
| C2 |
C3 |
H9 |
110.181 |
|
C2 |
C3 |
H10 |
110.787 |
| C2 |
C3 |
H11 |
110.787 |
|
C3 |
C2 |
H7 |
109.986 |
| C3 |
C2 |
H8 |
109.986 |
|
Cl4 |
C1 |
H5 |
107.021 |
| Cl4 |
C1 |
H6 |
107.021 |
|
H5 |
C1 |
H6 |
108.806 |
| H7 |
C2 |
H8 |
107.699 |
|
H9 |
C3 |
H10 |
108.316 |
| H9 |
C3 |
H11 |
108.316 |
|
H10 |
C3 |
H11 |
108.370 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -571.055587 |
| Energy at 298.15K | -571.063229 |
| Nuclear repulsion energy | 159.442185 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3566 |
3170 |
0.99 |
|
|
|
| 2 |
A |
3560 |
3164 |
2.51 |
|
|
|
| 3 |
A |
3529 |
3137 |
0.93 |
|
|
|
| 4 |
A |
3521 |
3130 |
0.46 |
|
|
|
| 5 |
A |
3409 |
3030 |
1.87 |
|
|
|
| 6 |
A |
3405 |
3027 |
1.21 |
|
|
|
| 7 |
A |
3382 |
3006 |
2.21 |
|
|
|
| 8 |
A |
1737 |
1544 |
1.16 |
|
|
|
| 9 |
A |
1730 |
1537 |
2.06 |
|
|
|
| 10 |
A |
1700 |
1511 |
0.16 |
|
|
|
| 11 |
A |
1675 |
1489 |
1.17 |
|
|
|
| 12 |
A |
1622 |
1442 |
0.26 |
|
|
|
| 13 |
A |
1549 |
1377 |
4.18 |
|
|
|
| 14 |
A |
1484 |
1319 |
32.95 |
|
|
|
| 15 |
A |
1421 |
1263 |
4.45 |
|
|
|
| 16 |
A |
1362 |
1211 |
1.02 |
|
|
|
| 17 |
A |
1227 |
1091 |
1.60 |
|
|
|
| 18 |
A |
1198 |
1065 |
1.13 |
|
|
|
| 19 |
A |
1168 |
1039 |
1.49 |
|
|
|
| 20 |
A |
1007 |
895 |
6.36 |
|
|
|
| 21 |
A |
955 |
849 |
4.78 |
|
|
|
| 22 |
A |
887 |
788 |
10.20 |
|
|
|
| 23 |
A |
782 |
695 |
4.33 |
|
|
|
| 24 |
A |
445 |
396 |
1.21 |
|
|
|
| 25 |
A |
294 |
261 |
0.91 |
|
|
|
| 26 |
A |
211 |
187 |
0.94 |
|
|
|
| 27 |
A |
125 |
111 |
1.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23474.1 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 20866.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.188 |
0.902 |
0.317 |
| C2 |
-1.178 |
0.577 |
-0.369 |
| C3 |
-1.825 |
-0.740 |
0.152 |
| Cl4 |
1.474 |
-0.352 |
-0.071 |
| H5 |
0.581 |
1.879 |
-0.029 |
| H6 |
0.084 |
0.927 |
1.421 |
| H7 |
-1.854 |
1.431 |
-0.171 |
| H8 |
-1.026 |
0.517 |
-1.463 |
| H9 |
-2.788 |
-0.917 |
-0.355 |
| H10 |
-1.164 |
-1.601 |
-0.043 |
| H11 |
-2.009 |
-0.681 |
1.238 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5625 | 2.6030 | 1.8379 | 1.1086 | 1.1087 | 2.1652 | 2.1889 | 3.5523 | 2.8668 | 2.8597 |
C2 | 1.5625 | | 1.5574 | 2.8258 | 2.2147 | 2.2180 | 1.1075 | 1.1060 | 2.1970 | 2.2016 | 2.2037 | C3 | 2.6030 | 1.5574 | | 3.3297 | 3.5613 | 2.8348 | 2.1954 | 2.1975 | 1.1029 | 1.1026 | 1.1032 | Cl4 | 1.8379 | 2.8258 | 3.3297 | | 2.4037 | 2.4070 | 3.7773 | 2.9909 | 4.3095 | 2.9188 | 3.7355 | H5 | 1.1086 | 2.2147 | 3.5613 | 2.4037 | | 1.8036 | 2.4800 | 2.5485 | 4.3907 | 3.8925 | 3.8554 | H6 | 1.1087 | 2.2180 | 2.8348 | 2.4070 | 1.8036 | | 2.5584 | 3.1173 | 3.8481 | 3.1766 | 2.6457 | H7 | 2.1652 | 1.1075 | 2.1954 | 3.7773 | 2.4800 | 2.5584 | | 1.7860 | 2.5339 | 3.1118 | 2.5437 | H8 | 2.1889 | 1.1060 | 2.1975 | 2.9909 | 2.5485 | 3.1173 | 1.7860 | | 2.5283 | 2.5534 | 3.1143 | H9 | 3.5523 | 2.1970 | 1.1029 | 4.3095 | 4.3907 | 3.8481 | 2.5339 | 2.5283 | | 1.7899 | 1.7894 | H10 | 2.8668 | 2.2016 | 1.1026 | 2.9188 | 3.8925 | 3.1766 | 3.1118 | 2.5534 | 1.7899 | | 1.7896 | H11 | 2.8597 | 2.2037 | 1.1032 | 3.7355 | 3.8554 | 2.6457 | 2.5437 | 3.1143 | 1.7894 | 1.7896 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.091 |
|
C1 |
C2 |
H7 |
107.143 |
| C1 |
C2 |
H8 |
109.016 |
|
C2 |
C1 |
Cl4 |
112.150 |
| C2 |
C1 |
H5 |
110.868 |
|
C2 |
C1 |
H6 |
111.125 |
| C2 |
C3 |
H9 |
110.176 |
|
C2 |
C3 |
H10 |
110.554 |
| C2 |
C3 |
H11 |
110.684 |
|
C3 |
C2 |
H7 |
109.790 |
| C3 |
C2 |
H8 |
110.035 |
|
Cl4 |
C1 |
H5 |
106.696 |
| Cl4 |
C1 |
H6 |
106.932 |
|
H5 |
C1 |
H6 |
108.867 |
| H7 |
C2 |
H8 |
107.582 |
|
H9 |
C3 |
H10 |
108.499 |
| H9 |
C3 |
H11 |
108.413 |
|
H10 |
C3 |
H11 |
108.447 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
