Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -318.005682 |
| Energy at 298.15K | -318.014197 |
| HF Energy | -317.647379 |
| Nuclear repulsion energy | 226.661821 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3558 |
3162 |
2.22 |
|
|
|
| 2 |
A' |
3413 |
3034 |
2.85 |
|
|
|
| 3 |
A' |
3376 |
3001 |
2.69 |
|
|
|
| 4 |
A' |
3351 |
2979 |
7.36 |
|
|
|
| 5 |
A' |
1742 |
1548 |
0.79 |
|
|
|
| 6 |
A' |
1722 |
1530 |
1.01 |
|
|
|
| 7 |
A' |
1707 |
1517 |
2.27 |
|
|
|
| 8 |
A' |
1624 |
1443 |
1.30 |
|
|
|
| 9 |
A' |
1578 |
1402 |
5.76 |
|
|
|
| 10 |
A' |
1464 |
1302 |
29.59 |
|
|
|
| 11 |
A' |
1368 |
1216 |
26.59 |
|
|
|
| 12 |
A' |
1269 |
1128 |
0.89 |
|
|
|
| 13 |
A' |
1198 |
1065 |
33.37 |
|
|
|
| 14 |
A' |
1166 |
1036 |
5.04 |
|
|
|
| 15 |
A' |
1089 |
968 |
181.42 |
|
|
|
| 16 |
A' |
1010 |
898 |
3.01 |
|
|
|
| 17 |
A' |
637 |
566 |
0.74 |
|
|
|
| 18 |
A' |
415 |
369 |
0.76 |
|
|
|
| 19 |
A' |
362 |
322 |
0.40 |
|
|
|
| 20 |
A' |
158 |
141 |
0.13 |
|
|
|
| 21 |
A" |
3556 |
3161 |
3.82 |
|
|
|
| 22 |
A" |
3526 |
3134 |
2.16 |
|
|
|
| 23 |
A" |
3463 |
3078 |
7.45 |
|
|
|
| 24 |
A" |
1734 |
1542 |
1.94 |
|
|
|
| 25 |
A" |
1451 |
1289 |
0.01 |
|
|
|
| 26 |
A" |
1390 |
1236 |
0.54 |
|
|
|
| 27 |
A" |
1268 |
1127 |
2.17 |
|
|
|
| 28 |
A" |
989 |
879 |
1.60 |
|
|
|
| 29 |
A" |
822 |
731 |
1.46 |
|
|
|
| 30 |
A" |
236 |
210 |
0.00 |
|
|
|
| 31 |
A" |
177 |
158 |
1.20 |
|
|
|
| 32 |
A" |
91 |
81 |
0.20 |
|
|
|
| 33 |
A" |
54 |
48 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25480.9 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 22650.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.945 |
2.273 |
0.000 |
| C2 |
-1.552 |
0.765 |
0.000 |
| C3 |
0.000 |
0.575 |
0.000 |
| O4 |
0.265 |
-0.880 |
0.000 |
| N5 |
1.732 |
-1.001 |
0.000 |
| O6 |
1.990 |
-2.278 |
0.000 |
| H7 |
-3.042 |
2.384 |
0.000 |
| H8 |
-1.545 |
2.781 |
0.894 |
| H9 |
-1.545 |
2.781 |
-0.894 |
| H10 |
-1.975 |
0.266 |
-0.892 |
| H11 |
-1.975 |
0.266 |
0.892 |
| H12 |
0.447 |
1.057 |
0.900 |
| H13 |
0.447 |
1.057 |
-0.900 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5584 | 2.5814 | 3.8502 | 4.9228 | 6.0163 | 1.1031 | 1.1034 | 1.1034 | 2.1966 | 2.1966 | 2.8293 | 2.8293 |
C2 | 1.5584 | | 1.5636 | 2.4512 | 3.7285 | 4.6699 | 2.2007 | 2.2056 | 2.2056 | 1.1063 | 1.1063 | 2.2113 | 2.2113 | C3 | 2.5814 | 1.5636 | | 1.4790 | 2.3416 | 3.4790 | 3.5394 | 2.8376 | 2.8376 | 2.1889 | 2.1889 | 1.1142 | 1.1142 | O4 | 3.8502 | 2.4512 | 1.4790 | | 1.4715 | 2.2204 | 4.6470 | 4.1808 | 4.1808 | 2.6698 | 2.6698 | 2.1437 | 2.1437 | N5 | 4.9228 | 3.7285 | 2.3416 | 1.4715 | | 1.3035 | 5.8523 | 5.0832 | 5.0832 | 4.0175 | 4.0175 | 2.5879 | 2.5879 | O6 | 6.0163 | 4.6699 | 3.4790 | 2.2204 | 1.3035 | | 6.8603 | 6.2365 | 6.2365 | 4.7948 | 4.7948 | 3.7841 | 3.7841 | H7 | 1.1031 | 2.2007 | 3.5394 | 4.6470 | 5.8523 | 6.8603 | | 1.7884 | 1.7884 | 2.5344 | 2.5344 | 3.8394 | 3.8394 | H8 | 1.1034 | 2.2056 | 2.8376 | 4.1808 | 5.0832 | 6.2365 | 1.7884 | | 1.7886 | 3.1148 | 2.5515 | 2.6339 | 3.1867 | H9 | 1.1034 | 2.2056 | 2.8376 | 4.1808 | 5.0832 | 6.2365 | 1.7884 | 1.7886 | | 2.5515 | 3.1148 | 3.1867 | 2.6339 | H10 | 2.1966 | 1.1063 | 2.1889 | 2.6698 | 4.0175 | 4.7948 | 2.5344 | 3.1148 | 2.5515 | | 1.7847 | 3.1146 | 2.5476 | H11 | 2.1966 | 1.1063 | 2.1889 | 2.6698 | 4.0175 | 4.7948 | 2.5344 | 2.5515 | 3.1148 | 1.7847 | | 2.5476 | 3.1146 | H12 | 2.8293 | 2.2113 | 1.1142 | 2.1437 | 2.5879 | 3.7841 | 3.8394 | 2.6339 | 3.1867 | 3.1146 | 2.5476 | | 1.7990 | H13 | 2.8293 | 2.2113 | 1.1142 | 2.1437 | 2.5879 | 3.7841 | 3.8394 | 3.1867 | 2.6339 | 2.5476 | 3.1146 | 1.7990 | |
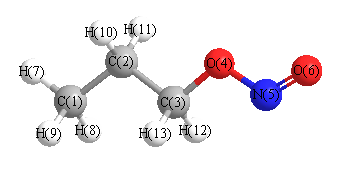 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.556 |
|
C1 |
C2 |
H10 |
109.886 |
| C1 |
C2 |
H11 |
109.886 |
|
C2 |
C1 |
H7 |
110.390 |
| C2 |
C1 |
H8 |
110.754 |
|
C2 |
C1 |
H9 |
110.754 |
| C2 |
C3 |
O4 |
107.306 |
|
C2 |
C3 |
H12 |
110.212 |
| C2 |
C3 |
H13 |
110.212 |
|
C3 |
C2 |
H10 |
108.940 |
| C3 |
C2 |
H11 |
108.940 |
|
C3 |
O4 |
N5 |
105.051 |
| O4 |
C3 |
H12 |
110.727 |
|
O4 |
C3 |
H13 |
110.727 |
| O4 |
N5 |
O6 |
106.133 |
|
H7 |
C1 |
H8 |
108.287 |
| H7 |
C1 |
H9 |
108.287 |
|
H8 |
C1 |
H9 |
108.282 |
| H10 |
C2 |
H11 |
107.539 |
|
H12 |
C3 |
H13 |
107.676 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -318.005607 |
| Energy at 298.15K | -318.014196 |
| HF Energy | -317.647287 |
| Nuclear repulsion energy | 230.859403 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3561 |
3165 |
2.29 |
|
|
|
| 2 |
A |
3557 |
3162 |
3.18 |
|
|
|
| 3 |
A |
3523 |
3131 |
1.95 |
|
|
|
| 4 |
A |
3464 |
3079 |
8.20 |
|
|
|
| 5 |
A |
3410 |
3031 |
2.67 |
|
|
|
| 6 |
A |
3378 |
3002 |
2.68 |
|
|
|
| 7 |
A |
3351 |
2979 |
7.29 |
|
|
|
| 8 |
A |
1740 |
1547 |
0.72 |
|
|
|
| 9 |
A |
1731 |
1539 |
1.64 |
|
|
|
| 10 |
A |
1712 |
1522 |
4.14 |
|
|
|
| 11 |
A |
1702 |
1513 |
0.87 |
|
|
|
| 12 |
A |
1622 |
1442 |
0.28 |
|
|
|
| 13 |
A |
1560 |
1386 |
4.48 |
|
|
|
| 14 |
A |
1528 |
1358 |
36.09 |
|
|
|
| 15 |
A |
1438 |
1278 |
0.62 |
|
|
|
| 16 |
A |
1371 |
1219 |
10.13 |
|
|
|
| 17 |
A |
1366 |
1214 |
12.58 |
|
|
|
| 18 |
A |
1280 |
1138 |
4.86 |
|
|
|
| 19 |
A |
1249 |
1110 |
2.26 |
|
|
|
| 20 |
A |
1207 |
1073 |
23.60 |
|
|
|
| 21 |
A |
1117 |
993 |
94.82 |
|
|
|
| 22 |
A |
1084 |
963 |
97.55 |
|
|
|
| 23 |
A |
1017 |
904 |
1.49 |
|
|
|
| 24 |
A |
976 |
868 |
0.33 |
|
|
|
| 25 |
A |
851 |
757 |
3.51 |
|
|
|
| 26 |
A |
599 |
532 |
1.28 |
|
|
|
| 27 |
A |
497 |
442 |
0.08 |
|
|
|
| 28 |
A |
332 |
295 |
0.71 |
|
|
|
| 29 |
A |
260 |
231 |
0.39 |
|
|
|
| 30 |
A |
181 |
161 |
0.40 |
|
|
|
| 31 |
A |
170 |
152 |
0.64 |
|
|
|
| 32 |
A |
99 |
88 |
0.11 |
|
|
|
| 33 |
A |
50 |
45 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25491.1 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 22659.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.264 |
-0.930 |
0.199 |
| C2 |
-1.816 |
0.469 |
-0.324 |
| C3 |
-0.411 |
0.899 |
0.215 |
| O4 |
0.576 |
-0.062 |
-0.311 |
| N5 |
1.838 |
0.250 |
0.335 |
| O6 |
2.689 |
-0.571 |
-0.143 |
| H7 |
-3.262 |
-1.191 |
-0.194 |
| H8 |
-2.316 |
-0.940 |
1.301 |
| H9 |
-1.552 |
-1.712 |
-0.120 |
| H10 |
-1.791 |
0.470 |
-1.430 |
| H11 |
-2.551 |
1.238 |
-0.016 |
| H12 |
-0.161 |
1.931 |
-0.128 |
| H13 |
-0.397 |
0.893 |
1.331 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5586 | 2.6034 | 3.0122 | 4.2701 | 4.9771 | 1.1038 | 1.1040 | 1.1036 | 2.1990 | 2.1978 | 3.5656 | 2.8441 |
C2 | 1.5586 | | 1.5654 | 2.4504 | 3.7198 | 4.6271 | 2.2047 | 2.2076 | 2.2057 | 1.1069 | 1.1075 | 2.2175 | 2.2205 | C3 | 2.6034 | 1.5654 | | 1.4746 | 2.3440 | 3.4494 | 3.5584 | 2.8617 | 2.8688 | 2.1897 | 2.1791 | 1.1160 | 1.1155 | O4 | 3.0122 | 2.4504 | 1.4746 | | 1.4521 | 2.1798 | 4.0018 | 3.4250 | 2.6991 | 2.6712 | 3.3993 | 2.1328 | 2.1339 | N5 | 4.2701 | 3.7198 | 2.3440 | 1.4521 | | 1.2753 | 5.3258 | 4.4277 | 3.9432 | 4.0414 | 4.5128 | 2.6526 | 2.5299 | O6 | 4.9771 | 4.6271 | 3.4494 | 2.1798 | 1.2753 | | 5.9830 | 5.2222 | 4.3918 | 4.7753 | 5.5449 | 3.7920 | 3.7201 | H7 | 1.1038 | 2.2047 | 3.5584 | 4.0018 | 5.3258 | 5.9830 | | 1.7866 | 1.7886 | 2.5400 | 2.5373 | 4.4008 | 3.8565 | H8 | 1.1040 | 2.2076 | 2.8617 | 3.4250 | 4.4277 | 5.2222 | 1.7866 | | 1.7884 | 3.1182 | 2.5563 | 3.8638 | 2.6537 | H9 | 1.1036 | 2.2057 | 2.8688 | 2.6991 | 3.9432 | 4.3918 | 1.7886 | 1.7884 | | 2.5558 | 3.1161 | 3.8993 | 3.1972 | H10 | 2.1990 | 1.1069 | 2.1897 | 2.6712 | 4.0414 | 4.7753 | 2.5400 | 3.1182 | 2.5558 | | 1.7800 | 2.5470 | 3.1214 | H11 | 2.1978 | 1.1075 | 2.1791 | 3.3993 | 4.5128 | 5.5449 | 2.5373 | 2.5563 | 3.1161 | 1.7800 | | 2.4912 | 2.5637 | H12 | 3.5656 | 2.2175 | 1.1160 | 2.1328 | 2.6526 | 3.7920 | 4.4008 | 3.8638 | 3.8993 | 2.5470 | 2.4912 | | 1.8058 | H13 | 2.8441 | 2.2205 | 1.1155 | 2.1339 | 2.5299 | 3.7201 | 3.8565 | 2.6537 | 3.1972 | 3.1214 | 2.5637 | 1.8058 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.888 |
|
C1 |
C2 |
H10 |
110.019 |
| C1 |
C2 |
H11 |
109.896 |
|
C2 |
C1 |
H7 |
110.647 |
| C2 |
C1 |
H8 |
110.861 |
|
C2 |
C1 |
H9 |
110.738 |
| C2 |
C3 |
O4 |
107.382 |
|
C2 |
C3 |
H12 |
110.463 |
| C2 |
C3 |
H13 |
110.726 |
|
C3 |
C2 |
H10 |
108.844 |
| C3 |
C2 |
H11 |
108.001 |
|
C3 |
O4 |
N5 |
106.432 |
| O4 |
C3 |
H12 |
110.062 |
|
O4 |
C3 |
H13 |
110.174 |
| O4 |
N5 |
O6 |
105.934 |
|
H7 |
C1 |
H8 |
108.042 |
| H7 |
C1 |
H9 |
108.245 |
|
H8 |
C1 |
H9 |
108.206 |
| H10 |
C2 |
H11 |
107.000 |
|
H12 |
C3 |
H13 |
108.044 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
