return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (Methyl propyl ether)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-233.532932
Energy at 298.15K 
HF Energy-229.292989
Nuclear repulsion energy184.712661
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3556 3161 2.87      
2 A' 3516 3125 3.33      
3 A' 3413 3034 3.48      
4 A' 3375 3000 3.25      
5 A' 3321 2952 12.36      
6 A' 3309 2941 14.41      
7 A' 1745 1551 0.73      
8 A' 1738 1545 0.97      
9 A' 1731 1539 2.82      
10 A' 1714 1523 0.33      
11 A' 1646 1463 7.97      
12 A' 1625 1444 3.55      
13 A' 1593 1416 14.95      
14 A' 1476 1312 34.25      
15 A' 1333 1185 7.10      
16 A' 1274 1132 8.90      
17 A' 1228 1092 0.43      
18 A' 1172 1042 0.07      
19 A' 1073 954 1.47      
20 A' 1007 895 2.69      
21 A' 483 429 0.21      
22 A' 423 376 0.60      
23 A' 203 180 0.87      
24 A" 3555 3160 4.77      
25 A" 3526 3134 2.51      
26 A" 3463 3078 12.64      
27 A" 3427 3046 18.44      
28 A" 1737 1544 1.68      
29 A" 1707 1517 1.73      
30 A" 1453 1292 0.00      
31 A" 1398 1243 1.19      
32 A" 1277 1135 5.18      
33 A" 1229 1092 2.58      
34 A" 996 886 1.72      
35 A" 827 735 1.29      
36 A" 261 232 1.82      
37 A" 238 212 0.04      
38 A" 120 107 2.13      
39 A" 100 89 0.39      

Unscaled Zero Point Vibrational Energy (zpe) 33631.5 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 29895.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.61531 0.06853 0.06472

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.304 0.803 0.000
O2 -1.329 -0.283 0.000
C3 0.000 0.332 0.000
C4 1.061 -0.818 0.000
C5 2.517 -0.262 0.000
H6 -3.311 0.339 0.000
H7 -2.216 1.454 0.899
H8 -2.216 1.454 -0.899
H9 0.145 0.980 0.897
H10 0.145 0.980 -0.897
H11 0.901 -1.453 -0.892
H12 0.901 -1.453 0.892
H13 2.698 0.359 0.894
H14 2.698 0.359 -0.894
H15 3.247 -1.088 0.000

Atom - Atom Distances (Å)
  C1 O2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.45972.35213.73554.93721.10851.11351.11352.61502.61504.02014.02015.10105.10105.8649
O21.45971.46432.44933.84582.07712.14742.14742.13872.13872.67192.67194.17494.17494.6467
C32.35211.46431.56442.58563.31102.64172.64171.11651.11652.18902.18902.84242.84243.5441
C43.73552.44931.56441.55844.52254.08764.08762.20812.20811.10611.10612.20582.20582.2029
C54.93723.84582.58561.55845.85855.11375.11372.82302.82302.19612.19611.10341.10341.1032
H61.10852.07713.31104.52255.85851.80291.80293.62813.62814.66374.66376.07526.07526.7117
H71.11352.14742.64174.08765.11371.80291.79852.40863.00494.62324.26235.03455.34436.0925
H81.11352.14742.64174.08765.11371.80291.79853.00492.40864.26234.62325.34435.03456.0925
H92.61502.13871.11652.20812.82303.62812.40863.00491.79483.11282.54732.62693.17953.8345
H102.61502.13871.11652.20812.82303.62813.00492.40861.79482.54733.11283.17952.62693.8345
H114.02012.67192.18901.10612.19614.66374.62324.26233.11282.54731.78343.11452.55182.5359
H124.02012.67192.18901.10612.19614.66374.26234.62322.54733.11281.78342.55183.11452.5359
H135.10104.17492.84242.20581.10346.07525.03455.34432.62693.17953.11452.55181.78781.7878
H145.10104.17492.84242.20581.10346.07525.34435.03453.17952.62692.55183.11451.78781.7878
H155.86494.64673.54412.20291.10326.71176.09256.09253.83453.83452.53592.53591.78781.7878

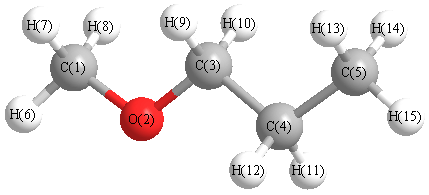
picture of Methyl propyl ether state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C3 107.111 O2 C1 H6 107.168
O2 C1 H7 112.444 O2 C1 H8 112.444
O2 C3 C4 107.886 O2 C3 H9 111.216
O2 C3 H10 111.216 C3 C4 C5 111.781
C3 C4 H11 108.899 C3 C4 H12 108.899
C4 C3 H9 109.773 C4 C3 H10 109.773
C4 C5 H13 110.762 C4 C5 H14 110.762
C4 C5 H15 110.550 C5 C4 H11 109.851
C5 C4 H12 109.851 H6 C1 H7 108.467
H6 C1 H8 108.467 H7 C1 H8 107.723
H9 C3 H10 106.979 H11 C4 H12 107.451
H13 C5 H14 108.211 H13 C5 H15 108.231
H14 C5 H15 108.231
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability