return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (2,3-dimethylbut-1-ene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-231.737442
Energy at 298.15K-231.749371
HF Energy-231.397077
Nuclear repulsion energy239.375657
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3607 3206 0.89      
2 A' 3556 3161 5.43      
3 A' 3551 3157 0.27      
4 A' 3550 3156 7.59      
5 A' 3453 3069 3.67      
6 A' 3428 3047 4.56      
7 A' 3371 2997 2.68      
8 A' 3367 2993 1.93      
9 A' 1830 1627 0.47      
10 A' 1742 1549 0.38      
11 A' 1738 1545 2.51      
12 A' 1728 1536 1.04      
13 A' 1627 1446 1.41      
14 A' 1623 1443 0.01      
15 A' 1600 1422 0.52      
16 A' 1500 1334 8.56      
17 A' 1446 1285 6.85      
18 A' 1325 1178 3.00      
19 A' 1223 1087 11.07      
20 A' 1118 994 1.61      
21 A' 1071 952 0.10      
22 A' 989 879 0.59      
23 A' 786 699 0.59      
24 A' 550 489 0.79      
25 A' 450 400 0.45      
26 A' 341 303 0.11      
27 A' 280 249 0.10      
28 A' 243 216 0.02      
29 A" 3555 3160 0.74      
30 A" 3549 3155 1.09      
31 A" 3537 3144 2.20      
32 A" 3372 2997 3.70      
33 A" 1740 1547 0.01      
34 A" 1731 1539 0.08      
35 A" 1726 1534 1.95      
36 A" 1611 1432 0.35      
37 A" 1518 1350 6.48      
38 A" 1264 1124 1.81      
39 A" 1186 1055 0.87      
40 A" 1082 962 0.21      
41 A" 1042 926 0.63      
42 A" 981 872 13.90      
43 A" 822 731 0.08      
44 A" 570 506 2.26      
45 A" 260 231 0.05      
46 A" 212 188 0.10      
47 A" 148 131 0.05      
48 A" 28 25 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 40513.6 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 36012.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.14248 0.07996 0.07911

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.777 -1.087 0.000
C2 -0.486 -0.684 0.000
C3 -0.127 0.830 0.000
C4 0.667 -1.711 0.000
C5 0.667 1.240 1.287
C6 0.667 1.240 -1.287
H7 -2.050 -2.151 0.000
H8 -2.606 -0.367 0.000
H9 -1.083 1.393 0.000
H10 0.267 -2.739 0.000
H11 1.308 -1.591 -0.891
H12 1.308 -1.591 0.891
H13 0.827 2.332 1.304
H14 0.110 0.958 2.197
H15 1.655 0.750 1.318
H16 0.827 2.332 -1.304
H17 0.110 0.958 -2.197
H18 1.655 0.750 -1.318

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.35252.52982.52303.61263.61261.09831.09832.57532.62833.25093.25094.49153.54624.11014.49153.54624.1101
C21.35251.55621.54462.58672.58672.14442.14332.16072.18872.19922.19923.53862.80732.89463.53862.80732.8946
C32.52981.55622.66231.56721.56723.54782.75271.10933.59072.95172.95172.20582.21382.21762.20582.21382.2176
C42.52301.54462.66233.21993.21992.75313.53873.56321.10311.10451.10454.25143.50192.96144.25143.50192.9614
C53.61262.58671.56723.21992.57464.53263.86692.17804.20143.62912.92921.10381.10361.10292.81623.54012.8287
C63.61262.58671.56723.21992.57464.53263.86692.17804.20142.92923.62912.81623.54012.82871.10381.10361.1029
H71.09832.14443.54782.75314.53264.53261.86913.67342.39093.51993.51995.48434.37784.88715.48434.37784.8871
H81.09832.14332.75273.53873.86693.86691.86912.32683.72614.19694.19694.55693.73654.59774.55693.73654.5977
H92.57532.16071.10933.56322.17802.17803.67342.32684.34663.92573.92572.49572.53793.10562.49572.53793.1056
H102.62832.18873.59071.10314.20144.20142.39093.72614.34661.78821.78825.26604.30363.97965.26604.30363.9796
H113.25092.19922.95171.10453.62912.92923.51994.19693.92571.78821.78284.52094.17973.23713.97373.10412.4043
H123.25092.19922.95171.10452.92923.62913.51994.19693.92571.78821.78283.97373.10412.40434.52094.17973.2371
H134.49153.53862.20584.25141.10382.81625.48434.55692.49575.26604.52093.97371.78891.78602.60723.82853.1719
H143.54622.80732.21383.50191.10363.54014.37783.73652.53794.30364.17973.10411.78891.78953.82854.39463.8451
H154.11012.89462.21762.96141.10292.82874.88714.59773.10563.97963.23712.40431.78601.78953.17193.84512.6355
H164.49153.53862.20584.25142.81621.10385.48434.55692.49575.26603.97374.52092.60723.82853.17191.78891.7860
H173.54622.80732.21383.50193.54011.10364.37783.73652.53794.30363.10414.17973.82854.39463.84511.78891.7895
H184.11012.89462.21762.96142.82871.10294.88714.59773.10563.97962.40433.23713.17193.84512.63551.78601.7895

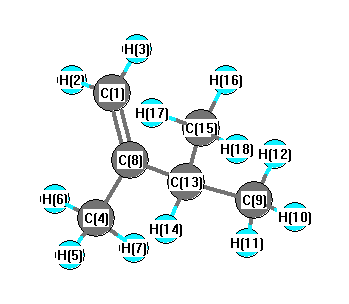
picture of 2,3-dimethylbut-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.698 C1 C2 C4 120.979
C2 C1 H7 121.740 C2 C1 H8 121.637
C2 C3 C5 111.821 C2 C3 C6 111.821
C2 C3 H9 107.123 C2 C4 H10 110.404
C2 C4 H11 111.149 C2 C4 H12 111.149
C3 C2 C4 118.323 C3 C5 H13 110.134
C3 C5 H14 110.773 C3 C5 H15 111.110
C3 C6 H16 110.134 C3 C6 H17 110.773
C3 C6 H18 111.110 C5 C3 C6 110.447
C5 C3 H9 107.698 C6 C3 H9 107.698
H7 C1 H8 116.623 H10 C4 H11 108.200
H10 C4 H12 108.200 H11 C4 H12 107.617
H13 C5 H14 108.274 H13 C5 H15 108.062
H14 C5 H15 108.392 H16 C6 H17 108.274
H16 C6 H18 108.062 H17 C6 H18 108.392
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability