Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -608.479818 |
| Energy at 298.15K | -608.486600 |
| HF Energy | -608.239327 |
| Nuclear repulsion energy | 201.745526 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3554 |
3159 |
1.23 |
|
|
|
| 2 |
A' |
3520 |
3129 |
3.20 |
|
|
|
| 3 |
A' |
3496 |
3108 |
4.42 |
|
|
|
| 4 |
A' |
3387 |
3011 |
1.24 |
|
|
|
| 5 |
A' |
3366 |
2992 |
2.27 |
|
|
|
| 6 |
A' |
1860 |
1653 |
0.71 |
|
|
|
| 7 |
A' |
1725 |
1533 |
2.69 |
|
|
|
| 8 |
A' |
1675 |
1489 |
0.87 |
|
|
|
| 9 |
A' |
1614 |
1434 |
3.60 |
|
|
|
| 10 |
A' |
1465 |
1302 |
41.19 |
|
|
|
| 11 |
A' |
1428 |
1270 |
1.87 |
|
|
|
| 12 |
A' |
1411 |
1254 |
0.73 |
|
|
|
| 13 |
A' |
1227 |
1091 |
1.56 |
|
|
|
| 14 |
A' |
1141 |
1014 |
4.94 |
|
|
|
| 15 |
A' |
1009 |
897 |
3.06 |
|
|
|
| 16 |
A' |
826 |
734 |
10.19 |
|
|
|
| 17 |
A' |
620 |
551 |
2.13 |
|
|
|
| 18 |
A' |
347 |
308 |
0.67 |
|
|
|
| 19 |
A' |
174 |
154 |
1.54 |
|
|
|
| 20 |
A" |
3533 |
3140 |
1.33 |
|
|
|
| 21 |
A" |
3498 |
3109 |
0.35 |
|
|
|
| 22 |
A" |
1719 |
1528 |
1.78 |
|
|
|
| 23 |
A" |
1309 |
1164 |
0.13 |
|
|
|
| 24 |
A" |
1184 |
1053 |
0.08 |
|
|
|
| 25 |
A" |
1101 |
979 |
13.02 |
|
|
|
| 26 |
A" |
1023 |
909 |
0.26 |
|
|
|
| 27 |
A" |
759 |
675 |
0.00 |
|
|
|
| 28 |
A" |
244 |
217 |
2.77 |
|
|
|
| 29 |
A" |
172 |
153 |
0.11 |
|
|
|
| 30 |
A" |
70 |
62 |
1.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24227.1 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 21535.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.351 |
0.304 |
0.000 |
| C2 |
0.000 |
1.050 |
0.000 |
| C3 |
1.225 |
0.485 |
0.000 |
| C4 |
2.534 |
1.290 |
0.000 |
| Cl5 |
-1.199 |
-1.525 |
0.000 |
| H6 |
-1.947 |
0.574 |
0.898 |
| H7 |
-1.947 |
0.574 |
-0.898 |
| H8 |
-0.107 |
2.147 |
0.000 |
| H9 |
1.317 |
-0.612 |
0.000 |
| H10 |
2.331 |
2.375 |
0.000 |
| H11 |
3.140 |
1.047 |
0.892 |
| H12 |
3.140 |
1.047 |
-0.892 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5435 | 2.5831 | 4.0088 | 1.8354 | 1.1109 | 1.1109 | 2.2237 | 2.8210 | 4.2247 | 4.6392 | 4.6392 |
C2 | 1.5435 | | 1.3491 | 2.5456 | 2.8404 | 2.1959 | 2.1959 | 1.1024 | 2.1203 | 2.6813 | 3.2644 | 3.2644 | C3 | 2.5831 | 1.3491 | | 1.5365 | 3.1495 | 3.2979 | 3.2979 | 2.1298 | 1.1013 | 2.1890 | 2.1857 | 2.1857 | C4 | 4.0088 | 2.5456 | 1.5365 | | 4.6758 | 4.6257 | 4.6257 | 2.7768 | 2.2585 | 1.1032 | 1.1052 | 1.1052 | Cl5 | 1.8354 | 2.8404 | 3.1495 | 4.6758 | | 2.4029 | 2.4029 | 3.8309 | 2.6763 | 5.2601 | 5.1226 | 5.1226 | H6 | 1.1109 | 2.1959 | 3.2979 | 4.6257 | 2.4029 | | 1.7961 | 2.5812 | 3.5867 | 4.7272 | 5.1088 | 5.4132 | H7 | 1.1109 | 2.1959 | 3.2979 | 4.6257 | 2.4029 | 1.7961 | | 2.5812 | 3.5867 | 4.7272 | 5.4132 | 5.1088 | H8 | 2.2237 | 1.1024 | 2.1298 | 2.7768 | 3.8309 | 2.5812 | 2.5812 | | 3.1047 | 2.4489 | 3.5426 | 3.5426 | H9 | 2.8210 | 2.1203 | 1.1013 | 2.2585 | 2.6763 | 3.5867 | 3.5867 | 3.1047 | | 3.1542 | 2.6217 | 2.6217 | H10 | 4.2247 | 2.6813 | 2.1890 | 1.1032 | 5.2601 | 4.7272 | 4.7272 | 2.4489 | 3.1542 | | 1.7921 | 1.7921 | H11 | 4.6392 | 3.2644 | 2.1857 | 1.1052 | 5.1226 | 5.1088 | 5.4132 | 3.5426 | 2.6217 | 1.7921 | | 1.7834 | H12 | 4.6392 | 3.2644 | 2.1857 | 1.1052 | 5.1226 | 5.4132 | 5.1088 | 3.5426 | 2.6217 | 1.7921 | 1.7834 | |
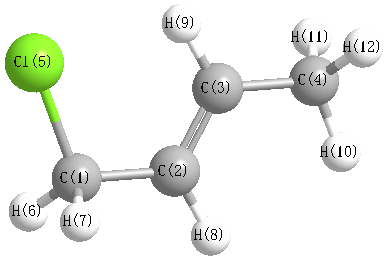 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
126.374 |
|
C1 |
C2 |
H8 |
113.319 |
| C2 |
C1 |
Cl5 |
114.133 |
|
C2 |
C1 |
H6 |
110.575 |
| C2 |
C1 |
H7 |
110.575 |
|
C2 |
C3 |
C4 |
123.682 |
| C2 |
C3 |
H9 |
119.489 |
|
C3 |
C2 |
H8 |
120.307 |
| C3 |
C4 |
H10 |
110.981 |
|
C3 |
C4 |
H11 |
110.603 |
| C3 |
C4 |
H12 |
110.603 |
|
C4 |
C3 |
H9 |
116.829 |
| Cl5 |
C1 |
H6 |
106.694 |
|
Cl5 |
C1 |
H7 |
106.694 |
| H6 |
C1 |
H7 |
107.880 |
|
H10 |
C4 |
H11 |
108.489 |
| H10 |
C4 |
H12 |
108.489 |
|
H11 |
C4 |
H12 |
107.572 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -608.481848 |
| Energy at 298.15K | -608.488660 |
| HF Energy | -608.241756 |
| Nuclear repulsion energy | 197.525997 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3556 |
3160 |
1.07 |
|
|
|
| 2 |
A |
3533 |
3140 |
1.14 |
|
|
|
| 3 |
A |
3524 |
3132 |
0.46 |
|
|
|
| 4 |
A |
3515 |
3124 |
2.49 |
|
|
|
| 5 |
A |
3498 |
3109 |
7.14 |
|
|
|
| 6 |
A |
3399 |
3021 |
1.04 |
|
|
|
| 7 |
A |
3366 |
2992 |
1.75 |
|
|
|
| 8 |
A |
1844 |
1640 |
2.45 |
|
|
|
| 9 |
A |
1722 |
1531 |
2.69 |
|
|
|
| 10 |
A |
1717 |
1526 |
2.18 |
|
|
|
| 11 |
A |
1666 |
1481 |
0.50 |
|
|
|
| 12 |
A |
1612 |
1433 |
4.55 |
|
|
|
| 13 |
A |
1468 |
1305 |
6.08 |
|
|
|
| 14 |
A |
1435 |
1275 |
5.02 |
|
|
|
| 15 |
A |
1405 |
1249 |
39.80 |
|
|
|
| 16 |
A |
1314 |
1168 |
0.32 |
|
|
|
| 17 |
A |
1223 |
1087 |
1.64 |
|
|
|
| 18 |
A |
1184 |
1053 |
0.17 |
|
|
|
| 19 |
A |
1173 |
1043 |
4.50 |
|
|
|
| 20 |
A |
1105 |
982 |
10.87 |
|
|
|
| 21 |
A |
1039 |
924 |
2.72 |
|
|
|
| 22 |
A |
949 |
844 |
2.84 |
|
|
|
| 23 |
A |
895 |
795 |
23.46 |
|
|
|
| 24 |
A |
789 |
701 |
6.40 |
|
|
|
| 25 |
A |
513 |
456 |
1.86 |
|
|
|
| 26 |
A |
372 |
330 |
1.64 |
|
|
|
| 27 |
A |
280 |
249 |
1.10 |
|
|
|
| 28 |
A |
187 |
166 |
0.79 |
|
|
|
| 29 |
A |
152 |
135 |
1.16 |
|
|
|
| 30 |
A |
77 |
69 |
0.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24255.1 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 21560.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.804 |
0.859 |
0.068 |
| C2 |
0.542 |
0.239 |
0.464 |
| C3 |
1.611 |
0.208 |
-0.364 |
| C4 |
2.972 |
-0.396 |
0.011 |
| Cl5 |
-2.124 |
-0.424 |
-0.083 |
| H6 |
-1.163 |
1.580 |
0.831 |
| H7 |
-0.738 |
1.369 |
-0.914 |
| H8 |
0.602 |
-0.196 |
1.474 |
| H9 |
1.526 |
0.644 |
-1.374 |
| H10 |
2.951 |
-0.805 |
1.036 |
| H11 |
3.240 |
-1.211 |
-0.686 |
| H12 |
3.768 |
0.369 |
-0.046 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5340 | 2.5379 | 3.9796 | 1.8465 | 1.1098 | 1.1087 | 2.2509 | 2.7488 | 4.2190 | 4.6052 | 4.5995 |
C2 | 1.5340 | | 1.3526 | 2.5522 | 2.8010 | 2.2005 | 2.1941 | 1.1012 | 2.1241 | 2.6864 | 3.2719 | 3.2685 | C3 | 2.5379 | 1.3526 | | 1.5363 | 3.7981 | 3.3175 | 2.6765 | 2.1356 | 1.1029 | 2.1867 | 2.1847 | 2.1865 | C4 | 3.9796 | 2.5522 | 1.5363 | | 5.0969 | 4.6562 | 4.2112 | 2.7924 | 2.2564 | 1.1031 | 1.1053 | 1.1053 | Cl5 | 1.8465 | 2.8010 | 3.7981 | 5.0969 | | 2.4030 | 2.4136 | 3.1476 | 4.0165 | 5.2102 | 5.4550 | 5.9449 | H6 | 1.1098 | 2.2005 | 3.3175 | 4.6562 | 2.4030 | | 1.8085 | 2.5857 | 3.6015 | 4.7597 | 5.4298 | 5.1530 | H7 | 1.1087 | 2.1941 | 2.6765 | 4.2112 | 2.4136 | 1.8085 | | 3.1539 | 2.4212 | 4.7041 | 4.7465 | 4.6962 | H8 | 2.2509 | 1.1012 | 2.1356 | 2.7924 | 3.1476 | 2.5857 | 3.1539 | | 3.1099 | 2.4654 | 3.5577 | 3.5569 | H9 | 2.7488 | 2.1241 | 1.1029 | 2.2564 | 4.0165 | 3.6015 | 2.4212 | 3.1099 | | 3.1518 | 2.6176 | 2.6200 | H10 | 4.2190 | 2.6864 | 2.1867 | 1.1031 | 5.2102 | 4.7597 | 4.7041 | 2.4654 | 3.1518 | | 1.7926 | 1.7929 | H11 | 4.6052 | 3.2719 | 2.1847 | 1.1053 | 5.4550 | 5.4298 | 4.7465 | 3.5577 | 2.6176 | 1.7926 | | 1.7842 | H12 | 4.5995 | 3.2685 | 2.1865 | 1.1053 | 5.9449 | 5.1530 | 4.6962 | 3.5569 | 2.6200 | 1.7929 | 1.7842 | |
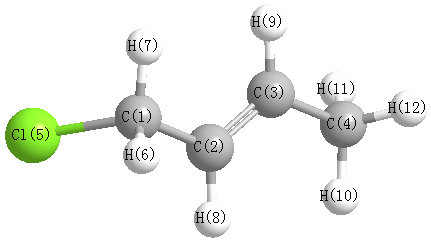 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
122.971 |
|
C1 |
C2 |
H8 |
116.374 |
| C2 |
C1 |
Cl5 |
111.571 |
|
C2 |
C1 |
H6 |
111.678 |
| C2 |
C1 |
H7 |
111.233 |
|
C2 |
C3 |
C4 |
124.007 |
| C2 |
C3 |
H9 |
119.433 |
|
C3 |
C2 |
H8 |
120.654 |
| C3 |
C4 |
H10 |
110.821 |
|
C3 |
C4 |
H11 |
110.534 |
| C3 |
C4 |
H12 |
110.673 |
|
C4 |
C3 |
H9 |
116.559 |
| Cl5 |
C1 |
H6 |
106.050 |
|
Cl5 |
C1 |
H7 |
106.847 |
| H6 |
C1 |
H7 |
109.212 |
|
H10 |
C4 |
H11 |
108.530 |
| H10 |
C4 |
H12 |
108.560 |
|
H11 |
C4 |
H12 |
107.628 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
