return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,5-Hexadiene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1Ag
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-230.520617
Energy at 298.15K-230.530050
HF Energy-230.166469
Nuclear repulsion energy208.895385
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3613 3212 0.31      
2 A 3517 3126 5.38      
3 A 3503 3114 0.24      
4 A 3457 3073 3.20      
5 A 3391 3015 0.27      
6 A 1824 1621 0.17      
7 A 1700 1512 0.60      
8 A 1615 1436 0.07      
9 A 1540 1369 1.14      
10 A 1429 1271 0.54      
11 A 1365 1214 0.60      
12 A 1186 1054 0.26      
13 A 1159 1030 0.00      
14 A 1132 1006 2.48      
15 A 1027 913 0.02      
16 A 995 885 4.88      
17 A 852 757 1.27      
18 A 665 591 2.44      
19 A 431 383 0.03      
20 A 362 322 0.02      
21 A 90 80 0.01      
22 A 68 60 0.01      
23 B 3613 3212 0.71      
24 B 3508 3119 2.98      
25 B 3501 3112 3.16      
26 B 3457 3073 9.36      
27 B 3396 3019 7.78      
28 B 1826 1623 0.17      
29 B 1719 1528 0.27      
30 B 1609 1430 2.90      
31 B 1458 1296 0.03      
32 B 1439 1279 0.72      
33 B 1408 1252 21.89      
34 B 1287 1144 0.13      
35 B 1137 1011 5.59      
36 B 1053 936 0.76      
37 B 1009 896 0.39      
38 B 995 885 19.14      
39 B 696 618 5.95      
40 B 444 395 0.63      
41 B 222 197 0.04      
42 B 102 90 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 34399.1 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 30577.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.39701 0.04546 0.04403

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.436 0.654 -0.327
C2 0.436 -0.654 -0.327
C3 0.436 1.924 -0.347
C4 -0.436 -1.924 -0.347
C5 0.455 2.846 0.636
C6 -0.455 -2.846 0.636
H7 -1.096 0.642 -1.218
H8 -1.085 0.665 0.569
H9 1.096 -0.642 -1.218
H10 1.085 -0.665 0.569
H11 1.085 2.052 -1.229
H12 -1.085 -2.052 -1.229
H13 1.099 3.734 0.582
H14 -0.177 2.748 1.529
H15 -1.099 -3.734 0.582
H16 0.177 -2.748 1.529

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.57261.54052.57792.55473.63051.10841.10652.19572.20432.25452.92563.55952.81084.53033.9244
C21.57262.57791.54053.63052.55472.19572.20431.10841.10652.92562.25454.53033.92443.55952.8108
C31.54052.57793.94501.34824.95092.17892.17652.78882.82181.10224.34752.14022.13945.93555.0415
C42.57791.54053.94504.95091.34822.78882.82182.17892.17654.34751.10225.93555.04152.14022.1394
C52.55473.63051.34824.95095.76463.27102.67074.00213.56812.12225.46301.09821.09856.76145.6723
C63.63052.55474.95091.34825.76464.00213.56813.27102.67075.46302.12226.76145.67231.09821.0985
H71.10842.19572.17892.78883.27104.00211.78722.54013.10792.59732.69464.19733.58194.73224.5458
H81.10652.20432.17652.82182.67073.56811.78723.10792.54583.14113.25823.76672.46744.39943.7640
H92.19571.10842.78882.17894.00213.27102.54013.10791.78722.69462.59734.73224.54584.19733.5819
H102.20431.10652.82182.17653.56812.67073.10792.54581.78723.25823.14114.39943.76403.76672.4674
H112.25452.92561.10224.34752.12225.46302.59733.14112.69463.25824.64352.47133.11186.44465.6107
H122.92562.25454.34751.10225.46302.12222.69463.25822.59733.14114.64356.44465.61072.47133.1118
H133.55954.53032.14025.93551.09826.76144.19733.76674.73224.39942.47136.44461.86967.78496.6160
H142.81083.92442.13945.04151.09855.67233.58192.46744.54583.76403.11185.61071.86966.61605.5082
H154.53033.55955.93552.14026.76141.09824.73224.39944.19733.76676.44462.47137.78496.61601.8696
H163.92442.81085.04152.13945.67231.09854.54583.76403.58192.46745.61073.11186.61605.50821.8696

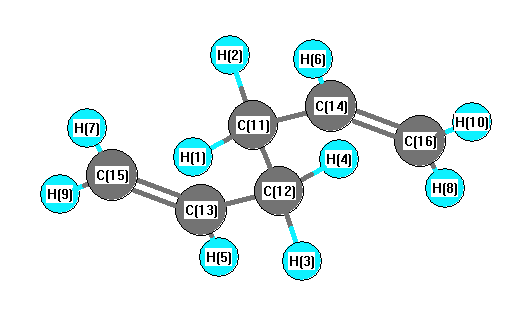
picture of 1,5-Hexadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 111.798 C1 C2 H9 108.729
C1 C2 H10 109.494 C1 C3 C5 124.219
C1 C3 H11 116.110 C2 C1 C3 111.798
C2 C1 H7 108.729 C2 C1 H8 109.494
C2 C4 C6 124.219 C2 C4 H12 116.110
C3 C1 H7 109.605 C3 C1 H8 109.531
C3 C5 H13 121.722 C3 C5 H14 121.615
C4 C2 H9 109.605 C4 C2 H10 109.531
C4 C6 H15 121.722 C4 C6 H16 121.615
C5 C3 H11 119.667 C6 C4 H12 119.667
H7 C1 H8 107.586 H9 C2 H10 107.586
H13 C5 H14 116.662 H15 C6 H16 116.662
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability