Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -187.325948 |
| Energy at 298.15K | |
| HF Energy | -187.055282 |
| Nuclear repulsion energy | 86.143687 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3370 |
2996 |
0.00 |
97.95 |
0.14 |
0.24 |
| 2 |
A1 |
2352 |
2091 |
274.23 |
0.86 |
0.22 |
0.36 |
| 3 |
A1 |
1811 |
1610 |
33.91 |
8.83 |
0.60 |
0.75 |
| 4 |
A1 |
1634 |
1452 |
1.81 |
32.02 |
0.61 |
0.76 |
| 5 |
A1 |
925 |
823 |
0.05 |
10.81 |
0.29 |
0.45 |
| 6 |
B1 |
1006 |
894 |
7.53 |
0.24 |
0.75 |
0.86 |
| 7 |
B1 |
616 |
548 |
5.64 |
0.57 |
0.75 |
0.86 |
| 8 |
B1 |
224 |
200 |
0.02 |
1.37 |
0.75 |
0.86 |
| 9 |
B2 |
3502 |
3113 |
2.18 |
61.97 |
0.75 |
0.86 |
| 10 |
B2 |
1162 |
1033 |
2.46 |
0.00 |
0.75 |
0.86 |
| 11 |
B2 |
354 |
314 |
5.50 |
0.29 |
0.75 |
0.86 |
| 12 |
B2 |
207i |
184i |
12.46 |
0.62 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8374.8 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 7444.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.910 |
| C2 |
0.000 |
0.000 |
-0.575 |
| C3 |
0.000 |
0.000 |
0.717 |
| O4 |
0.000 |
0.000 |
1.954 |
| H5 |
0.000 |
0.932 |
-2.511 |
| H6 |
0.000 |
-0.932 |
-2.511 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3340 | 2.6265 | 3.8632 | 1.1088 | 1.1088 |
C2 | 1.3340 | | 1.2924 | 2.5292 | 2.1477 | 2.1477 | C3 | 2.6265 | 1.2924 | | 1.2368 | 3.3593 | 3.3593 | O4 | 3.8632 | 2.5292 | 1.2368 | | 4.5605 | 4.5605 | H5 | 1.1088 | 2.1477 | 3.3593 | 4.5605 | | 1.8636 | H6 | 1.1088 | 2.1477 | 3.3593 | 4.5605 | 1.8636 | |
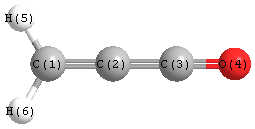 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
122.824 |
| C2 |
C1 |
H6 |
122.824 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
114.353 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -187.330009 |
| Energy at 298.15K | |
| HF Energy | -187.046600 |
| Nuclear repulsion energy | 86.125781 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3536 |
3143 |
1.18 |
53.41 |
0.68 |
0.81 |
| 2 |
A' |
3392 |
3015 |
1.24 |
70.25 |
0.14 |
0.24 |
| 3 |
A' |
2188 |
1945 |
147.35 |
2.29 |
0.56 |
0.71 |
| 4 |
A' |
1749 |
1555 |
55.93 |
10.03 |
0.66 |
0.79 |
| 5 |
A' |
1603 |
1425 |
7.44 |
21.74 |
0.60 |
0.75 |
| 6 |
A' |
1138 |
1011 |
20.02 |
2.96 |
0.07 |
0.13 |
| 7 |
A' |
931 |
827 |
6.55 |
11.77 |
0.18 |
0.30 |
| 8 |
A' |
456 |
405 |
6.14 |
2.48 |
0.67 |
0.80 |
| 9 |
A' |
233 |
208 |
10.68 |
1.25 |
0.68 |
0.81 |
| 10 |
A" |
996 |
885 |
9.74 |
0.41 |
0.75 |
0.86 |
| 11 |
A" |
759 |
674 |
1.18 |
1.62 |
0.75 |
0.86 |
| 12 |
A" |
262 |
233 |
1.58 |
2.05 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8620.5 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 7662.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.310 |
-1.810 |
0.000 |
| C2 |
-0.349 |
-0.628 |
0.000 |
| C3 |
0.000 |
0.680 |
0.000 |
| O4 |
-0.117 |
1.902 |
0.000 |
| H5 |
1.413 |
-1.907 |
0.000 |
| H6 |
-0.241 |
-2.767 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3532 | 2.5085 | 3.7363 | 1.1074 | 1.1046 |
C2 | 1.3532 | | 1.3533 | 2.5407 | 2.1776 | 2.1418 | C3 | 2.5085 | 1.3533 | | 1.2282 | 2.9475 | 3.4550 | O4 | 3.7363 | 2.5407 | 1.2282 | | 4.1051 | 4.6708 | H5 | 1.1074 | 2.1776 | 2.9475 | 4.1051 | | 1.8643 | H6 | 1.1046 | 2.1418 | 3.4550 | 4.6708 | 1.8643 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
135.894 |
|
C2 |
C1 |
H5 |
124.202 |
| C2 |
C1 |
H6 |
120.920 |
|
C2 |
C3 |
O4 |
159.580 |
| H5 |
C1 |
H6 |
114.878 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
