Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -273.334032 |
| Energy at 298.15K | |
| HF Energy | -273.195936 |
| Nuclear repulsion energy | 122.267992 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3340 |
2969 |
0.00 |
|
|
|
| 2 |
Ag |
1709 |
1519 |
0.00 |
|
|
|
| 3 |
Ag |
1605 |
1427 |
0.00 |
|
|
|
| 4 |
Ag |
1257 |
1117 |
0.00 |
|
|
|
| 5 |
Ag |
1104 |
982 |
0.00 |
|
|
|
| 6 |
Ag |
461 |
410 |
0.00 |
|
|
|
| 7 |
Au |
3457 |
3073 |
19.30 |
|
|
|
| 8 |
Au |
1335 |
1186 |
9.29 |
|
|
|
| 9 |
Au |
854 |
759 |
0.12 |
|
|
|
| 10 |
Au |
78 |
70 |
3.59 |
|
|
|
| 11 |
Bg |
3445 |
3062 |
0.00 |
|
|
|
| 12 |
Bg |
1425 |
1267 |
0.00 |
|
|
|
| 13 |
Bg |
1222 |
1086 |
0.00 |
|
|
|
| 14 |
Bu |
3344 |
2973 |
17.29 |
|
|
|
| 15 |
Bu |
1723 |
1531 |
0.56 |
|
|
|
| 16 |
Bu |
1478 |
1314 |
30.59 |
|
|
|
| 17 |
Bu |
1267 |
1126 |
10.61 |
|
|
|
| 18 |
Bu |
292 |
259 |
5.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14697.6 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 13064.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.423 |
0.671 |
0.000 |
| C2 |
-0.423 |
-0.671 |
0.000 |
| F3 |
-0.423 |
1.790 |
0.000 |
| F4 |
0.423 |
-1.790 |
0.000 |
| H5 |
1.083 |
0.684 |
0.901 |
| H6 |
1.083 |
0.684 |
-0.901 |
| H7 |
-1.083 |
-0.684 |
0.901 |
| H8 |
-1.083 |
-0.684 |
-0.901 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5862 | 1.4027 | 2.4612 | 1.1167 | 1.1167 | 2.2169 | 2.2169 |
C2 | 1.5862 | | 2.4612 | 1.4027 | 2.2169 | 2.2169 | 1.1167 | 1.1167 | F3 | 1.4027 | 2.4612 | | 3.6788 | 2.0741 | 2.0741 | 2.7146 | 2.7146 | F4 | 2.4612 | 1.4027 | 3.6788 | | 2.7146 | 2.7146 | 2.0741 | 2.0741 | H5 | 1.1167 | 2.2169 | 2.0741 | 2.7146 | | 1.8015 | 2.5617 | 3.1317 | H6 | 1.1167 | 2.2169 | 2.0741 | 2.7146 | 1.8015 | | 3.1317 | 2.5617 | H7 | 2.2169 | 1.1167 | 2.7146 | 2.0741 | 2.5617 | 3.1317 | | 1.8015 | H8 | 2.2169 | 1.1167 | 2.7146 | 2.0741 | 3.1317 | 2.5617 | 1.8015 | |
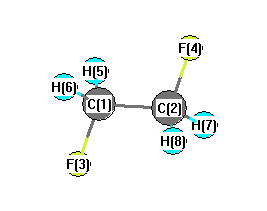 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.709 |
|
C1 |
C2 |
H7 |
108.964 |
| C1 |
C2 |
H8 |
108.964 |
|
C2 |
C1 |
F3 |
110.709 |
| C2 |
C1 |
H5 |
108.964 |
|
C2 |
C1 |
H6 |
108.964 |
| F3 |
C1 |
H5 |
110.302 |
|
F3 |
C1 |
H6 |
110.302 |
| F4 |
C2 |
H7 |
110.302 |
|
F4 |
C2 |
H8 |
110.302 |
| H5 |
C1 |
H6 |
107.524 |
|
H7 |
C2 |
H8 |
107.524 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -273.334603 |
| Energy at 298.15K | |
| HF Energy | -273.196463 |
| Nuclear repulsion energy | 124.104666 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3445 |
3062 |
6.28 |
44.19 |
0.75 |
0.86 |
| 2 |
A |
3339 |
2968 |
9.75 |
57.11 |
0.08 |
0.15 |
| 3 |
A |
1704 |
1515 |
0.61 |
3.77 |
0.73 |
0.84 |
| 4 |
A |
1575 |
1400 |
3.71 |
4.90 |
0.68 |
0.81 |
| 5 |
A |
1396 |
1241 |
2.48 |
15.58 |
0.74 |
0.85 |
| 6 |
A |
1274 |
1133 |
4.49 |
5.24 |
0.68 |
0.81 |
| 7 |
A |
1180 |
1049 |
1.46 |
3.79 |
0.51 |
0.68 |
| 8 |
A |
949 |
843 |
0.75 |
6.59 |
0.24 |
0.39 |
| 9 |
A |
305 |
271 |
0.01 |
0.37 |
0.53 |
0.69 |
| 10 |
A |
107 |
95 |
1.29 |
0.15 |
0.75 |
0.86 |
| 11 |
B |
3450 |
3067 |
12.21 |
10.26 |
0.75 |
0.86 |
| 12 |
B |
3337 |
2967 |
6.51 |
11.67 |
0.75 |
0.86 |
| 13 |
B |
1698 |
1509 |
0.97 |
18.40 |
0.75 |
0.86 |
| 14 |
B |
1561 |
1388 |
25.55 |
2.80 |
0.75 |
0.86 |
| 15 |
B |
1355 |
1204 |
8.12 |
5.81 |
0.75 |
0.86 |
| 16 |
B |
1257 |
1117 |
0.79 |
3.14 |
0.75 |
0.86 |
| 17 |
B |
988 |
878 |
4.83 |
2.97 |
0.75 |
0.86 |
| 18 |
B |
495 |
440 |
5.15 |
0.75 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 14707.0 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 13073.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.267 |
0.745 |
0.508 |
| C2 |
-0.267 |
-0.745 |
0.508 |
| F3 |
-0.267 |
1.490 |
-0.553 |
| F4 |
0.267 |
-1.490 |
-0.553 |
| H5 |
-0.009 |
1.213 |
1.484 |
| H6 |
1.382 |
0.728 |
0.443 |
| H7 |
0.009 |
-1.213 |
1.484 |
| H8 |
-1.382 |
-0.728 |
0.443 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5835 | 1.4024 | 2.4742 | 1.1170 | 1.1170 | 2.2027 | 2.2127 |
C2 | 1.5835 | | 2.4742 | 1.4024 | 2.2027 | 2.2127 | 1.1170 | 1.1170 | F3 | 1.4024 | 2.4742 | | 3.0274 | 2.0721 | 2.0724 | 3.3956 | 2.6745 | F4 | 2.4742 | 1.4024 | 3.0274 | | 3.3956 | 2.6745 | 2.0721 | 2.0724 | H5 | 1.1170 | 2.2027 | 2.0721 | 3.3956 | | 1.8045 | 2.4252 | 2.5951 | H6 | 1.1170 | 2.2127 | 2.0724 | 2.6745 | 1.8045 | | 2.5951 | 3.1246 | H7 | 2.2027 | 1.1170 | 3.3956 | 2.0721 | 2.4252 | 2.5951 | | 1.8045 | H8 | 2.2127 | 1.1170 | 2.6745 | 2.0724 | 2.5951 | 3.1246 | 1.8045 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
111.774 |
|
C1 |
C2 |
H7 |
108.064 |
| C1 |
C2 |
H8 |
108.817 |
|
C2 |
C1 |
F3 |
111.774 |
| C2 |
C1 |
H5 |
108.064 |
|
C2 |
C1 |
H6 |
108.817 |
| F3 |
C1 |
H5 |
110.148 |
|
F3 |
C1 |
H6 |
110.172 |
| F4 |
C2 |
H7 |
110.148 |
|
F4 |
C2 |
H8 |
110.172 |
| H5 |
C1 |
H6 |
107.753 |
|
H7 |
C2 |
H8 |
107.753 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
