Jump to
S1C2
Energy calculated at B2PLYP=FULLultrafine/STO-3G
| | hartrees |
|---|
| Energy at 0K | -274.070594 |
| Energy at 298.15K | |
| HF Energy | -274.021953 |
| Nuclear repulsion energy | 122.142463 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3289 |
3289 |
0.00 |
74.53 |
0.14 |
0.25 |
| 2 |
Ag |
1683 |
1683 |
0.00 |
25.25 |
0.75 |
0.86 |
| 3 |
Ag |
1568 |
1568 |
0.00 |
4.16 |
0.62 |
0.77 |
| 4 |
Ag |
1224 |
1224 |
0.00 |
7.21 |
0.61 |
0.76 |
| 5 |
Ag |
1070 |
1070 |
0.00 |
9.14 |
0.34 |
0.51 |
| 6 |
Ag |
451 |
451 |
0.00 |
2.78 |
0.53 |
0.70 |
| 7 |
Au |
3403 |
3403 |
20.53 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
1311 |
1311 |
7.26 |
0.00 |
0.00 |
0.00 |
| 9 |
Au |
843 |
843 |
0.01 |
0.00 |
0.00 |
0.00 |
| 10 |
Au |
76 |
76 |
3.44 |
0.00 |
0.00 |
0.00 |
| 11 |
Bg |
3388 |
3388 |
0.00 |
59.82 |
0.75 |
0.86 |
| 12 |
Bg |
1399 |
1399 |
0.00 |
21.78 |
0.75 |
0.86 |
| 13 |
Bg |
1203 |
1203 |
0.00 |
4.46 |
0.75 |
0.86 |
| 14 |
Bu |
3295 |
3295 |
20.30 |
0.00 |
0.00 |
0.00 |
| 15 |
Bu |
1696 |
1696 |
0.25 |
0.00 |
0.00 |
0.00 |
| 16 |
Bu |
1455 |
1455 |
25.15 |
0.00 |
0.00 |
0.00 |
| 17 |
Bu |
1235 |
1235 |
19.55 |
0.00 |
0.00 |
0.00 |
| 18 |
Bu |
282 |
282 |
4.92 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 14434.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 14434.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/STO-3G
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.423 |
0.672 |
0.000 |
| C2 |
-0.423 |
-0.672 |
0.000 |
| F3 |
-0.423 |
1.793 |
0.000 |
| F4 |
0.423 |
-1.793 |
0.000 |
| H5 |
1.082 |
0.681 |
0.900 |
| H6 |
1.082 |
0.681 |
-0.900 |
| H7 |
-1.082 |
-0.681 |
0.900 |
| H8 |
-1.082 |
-0.681 |
-0.900 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5880 | 1.4051 | 2.4648 | 1.1153 | 1.1153 | 2.2148 | 2.2148 |
C2 | 1.5880 | | 2.4648 | 1.4051 | 2.2148 | 2.2148 | 1.1153 | 1.1153 | F3 | 1.4051 | 2.4648 | | 3.6848 | 2.0771 | 2.0771 | 2.7135 | 2.7135 | F4 | 2.4648 | 1.4051 | 3.6848 | | 2.7135 | 2.7135 | 2.0771 | 2.0771 | H5 | 1.1153 | 2.2148 | 2.0771 | 2.7135 | | 1.7998 | 2.5569 | 3.1269 | H6 | 1.1153 | 2.2148 | 2.0771 | 2.7135 | 1.7998 | | 3.1269 | 2.5569 | H7 | 2.2148 | 1.1153 | 2.7135 | 2.0771 | 2.5569 | 3.1269 | | 1.7998 | H8 | 2.2148 | 1.1153 | 2.7135 | 2.0771 | 3.1269 | 2.5569 | 1.7998 | |
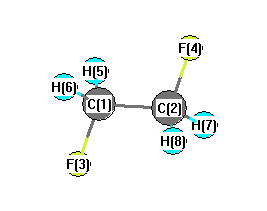 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.721 |
|
C1 |
C2 |
H7 |
108.767 |
| C1 |
C2 |
H8 |
108.767 |
|
C2 |
C1 |
F3 |
110.721 |
| C2 |
C1 |
H5 |
108.767 |
|
C2 |
C1 |
H6 |
108.767 |
| F3 |
C1 |
H5 |
110.462 |
|
F3 |
C1 |
H6 |
110.462 |
| F4 |
C2 |
H7 |
110.462 |
|
F4 |
C2 |
H8 |
110.462 |
| H5 |
C1 |
H6 |
107.578 |
|
H7 |
C2 |
H8 |
107.578 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/STO-3G
| | hartrees |
|---|
| Energy at 0K | -274.071181 |
| Energy at 298.15K | |
| HF Energy | -274.022531 |
| Nuclear repulsion energy | 123.964324 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3388 |
3388 |
6.95 |
49.75 |
0.75 |
0.86 |
| 2 |
A |
3287 |
3287 |
12.37 |
65.28 |
0.07 |
0.14 |
| 3 |
A |
1677 |
1677 |
0.34 |
3.60 |
0.74 |
0.85 |
| 4 |
A |
1539 |
1539 |
4.47 |
5.67 |
0.71 |
0.83 |
| 5 |
A |
1371 |
1371 |
2.79 |
15.67 |
0.74 |
0.85 |
| 6 |
A |
1240 |
1240 |
8.12 |
4.29 |
0.67 |
0.80 |
| 7 |
A |
1154 |
1154 |
2.07 |
4.06 |
0.53 |
0.70 |
| 8 |
A |
924 |
924 |
0.51 |
7.32 |
0.25 |
0.40 |
| 9 |
A |
298 |
298 |
0.04 |
0.43 |
0.53 |
0.69 |
| 10 |
A |
104 |
104 |
1.27 |
0.16 |
0.75 |
0.86 |
| 11 |
B |
3394 |
3394 |
13.37 |
10.81 |
0.75 |
0.86 |
| 12 |
B |
3285 |
3285 |
6.84 |
12.86 |
0.75 |
0.86 |
| 13 |
B |
1671 |
1671 |
1.34 |
19.23 |
0.75 |
0.86 |
| 14 |
B |
1530 |
1530 |
23.43 |
3.11 |
0.75 |
0.86 |
| 15 |
B |
1331 |
1331 |
7.08 |
5.88 |
0.75 |
0.86 |
| 16 |
B |
1229 |
1229 |
2.98 |
2.75 |
0.75 |
0.86 |
| 17 |
B |
973 |
973 |
4.64 |
3.08 |
0.75 |
0.86 |
| 18 |
B |
484 |
484 |
4.57 |
0.79 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 14438.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 14438.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/STO-3G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.269 |
0.745 |
0.507 |
| C2 |
-0.269 |
-0.745 |
0.507 |
| F3 |
-0.269 |
1.495 |
-0.551 |
| F4 |
0.269 |
-1.495 |
-0.551 |
| H5 |
0.000 |
1.208 |
1.486 |
| H6 |
1.383 |
0.721 |
0.437 |
| H7 |
-0.000 |
-1.208 |
1.486 |
| H8 |
-1.383 |
-0.721 |
0.437 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5849 | 1.4045 | 2.4777 | 1.1158 | 1.1156 | 2.2011 | 2.2103 |
C2 | 1.5849 | | 2.4777 | 1.4045 | 2.2011 | 2.2103 | 1.1158 | 1.1156 | F3 | 1.4045 | 2.4777 | | 3.0386 | 2.0750 | 2.0748 | 3.3953 | 2.6699 | F4 | 2.4777 | 1.4045 | 3.0386 | | 3.3953 | 2.6699 | 2.0750 | 2.0748 | H5 | 1.1158 | 2.2011 | 2.0750 | 3.3953 | | 1.8025 | 2.4152 | 2.5949 | H6 | 1.1156 | 2.2103 | 2.0748 | 2.6699 | 1.8025 | | 2.5949 | 3.1190 | H7 | 2.2011 | 1.1158 | 3.3953 | 2.0750 | 2.4152 | 2.5949 | | 1.8025 | H8 | 2.2103 | 1.1156 | 2.6699 | 2.0748 | 2.5949 | 3.1190 | 1.8025 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
111.818 |
|
C1 |
C2 |
H7 |
107.912 |
| C1 |
C2 |
H8 |
108.614 |
|
C2 |
C1 |
F3 |
111.818 |
| C2 |
C1 |
H5 |
107.912 |
|
C2 |
C1 |
H6 |
108.614 |
| F3 |
C1 |
H5 |
110.313 |
|
F3 |
C1 |
H6 |
110.306 |
| F4 |
C2 |
H7 |
110.313 |
|
F4 |
C2 |
H8 |
110.306 |
| H5 |
C1 |
H6 |
107.753 |
|
H7 |
C2 |
H8 |
107.753 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
