return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SSH (Hydrogen methyl disulfide)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-826.180624
Energy at 298.15K 
HF Energy-826.076107
Nuclear repulsion energy147.783225
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3535 3142 1.07 36.70 0.71 0.83
2 A 3518 3127 5.49 23.95 0.74 0.85
3 A 3346 2974 3.54 38.01 0.01 0.02
4 A 3039 2702 33.42 71.99 0.30 0.46
5 A 1736 1544 3.40 13.69 0.73 0.84
6 A 1721 1530 2.65 18.57 0.75 0.86
7 A 1588 1412 13.75 4.45 0.68 0.81
8 A 1098 976 7.56 4.63 0.61 0.76
9 A 1092 971 1.12 4.58 0.73 0.85
10 A 1052 935 15.30 14.43 0.74 0.85
11 A 868 772 0.75 11.46 0.36 0.53
12 A 636 566 0.13 15.41 0.29 0.44
13 A 278 247 3.66 2.85 0.67 0.80
14 A 242 215 6.30 15.08 0.74 0.85
15 A 162 144 0.24 0.13 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 11955.9 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 10627.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.52332 0.14752 0.12063

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.607 0.714 -0.006
S2 -0.493 -0.733 0.012
S3 1.351 0.248 -0.086
H4 1.496 0.418 1.250
H5 -1.482 1.329 -0.915
H6 -2.648 0.346 0.018
H7 -1.459 1.370 0.872

Atom - Atom Distances (Å)
  C1 S2 S3 H4 H5 H6 H7
C11.82602.99613.36101.10521.10391.1058
S21.82602.09132.61032.46772.40962.4686
S32.99612.09131.35493.14404.00163.1741
H43.36102.61031.35493.79364.32413.1279
H51.10522.46773.14403.79361.78741.7879
H61.10392.40964.00164.32411.78741.7866
H71.10582.46863.17413.12791.78791.7866

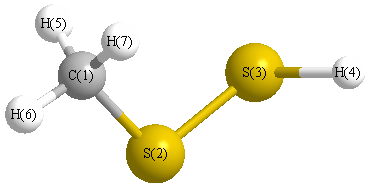
picture of Hydrogen methyl disulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 S3 99.563 S2 C1 H5 112.330
S2 C1 H6 108.118 S2 C1 H7 112.370
S2 S3 H4 96.122 H5 C1 H6 108.016
H5 C1 H7 107.926 H6 C1 H7 107.908
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability