return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CSNH2 (Ethanethioamide)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-524.800541
Energy at 298.15K 
HF Energy-524.589194
Nuclear repulsion energy151.418213
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3842 3415 2.12      
2 A 3637 3233 2.63      
3 A 3554 3159 0.74      
4 A 3529 3137 1.92      
5 A 3360 2986 1.25      
6 A 1854 1648 9.32      
7 A 1724 1532 0.06      
8 A 1719 1528 1.85      
9 A 1596 1419 0.02      
10 A 1542 1370 43.49      
11 A 1386 1232 53.85      
12 A 1180 1049 4.87      
13 A 1138 1012 6.74      
14 A 1105 982 2.53      
15 A 844 750 171.97      
16 A 735 654 105.21      
17 A 470 417 14.17      
18 A 429 381 20.10      
19 A 377 335 9.41      
20 A 373 332 30.71      
21 A 120 107 0.30      

Unscaled Zero Point Vibrational Energy (zpe) 17256.5 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 15339.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.30238 0.15946 0.10715

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.239 0.039 0.006
S2 -1.382 -0.123 0.007
C3 1.269 -1.131 -0.001
N4 0.917 1.333 -0.121
H5 1.943 -1.067 0.874
H6 0.751 -2.104 0.028
H7 1.893 -1.089 -0.911
H8 1.773 1.354 0.491
H9 0.280 2.096 0.227

Atom - Atom Distances (Å)
  C1 S2 C3 N4 H5 H6 H7 H8 H9
C11.62911.55861.46632.20882.20372.20212.07742.0690
S21.62912.83582.72423.56292.91093.53573.51672.7813
C31.55862.83582.49211.10561.10301.10492.58273.3825
N41.46632.72422.49212.79383.44472.72831.05231.0526
H52.20883.56291.10562.79381.79181.78592.45753.6315
H62.20372.91091.10303.44471.79181.79393.63574.2311
H72.20213.53571.10492.72831.78591.79392.81923.7469
H82.07743.51672.58271.05232.45753.63572.81921.6872
H92.06902.78133.38251.05263.63154.23113.74691.6872

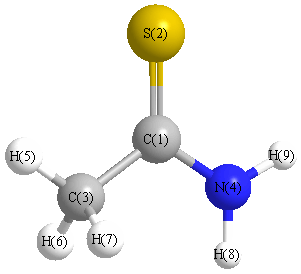
picture of Ethanethioamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H5 110.859 C1 C3 H6 110.613
C1 C3 H7 110.376 C1 N4 H8 110.055
C1 N4 H9 109.350 S2 C1 C3 125.633
S2 C1 N4 123.218 C3 C1 N4 110.903
H5 C3 H6 108.442 H5 C3 H7 107.783
H6 C3 H7 108.682 H8 N4 H9 106.560
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability