return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H5NO3 (Nitric acid, ethyl ester)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-353.198189
Energy at 298.15K-353.205430
HF Energy-352.824326
Nuclear repulsion energy239.351474
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3564 3168 0.52      
2 A' 3383 3008 0.77      
3 A' 3359 2986 3.82      
4 A' 2065 1835 60.21      
5 A' 1731 1539 2.03      
6 A' 1705 1516 5.76      
7 A' 1610 1431 0.09      
8 A' 1513 1345 20.52      
9 A' 1295 1152 156.80      
10 A' 1272 1131 17.51      
11 A' 1174 1044 4.33      
12 A' 1035 920 9.43      
13 A' 921 819 71.00      
14 A' 623 554 7.88      
15 A' 486 432 1.07      
16 A' 365 324 0.87      
17 A' 218 194 0.19      
18 A" 3568 3171 1.01      
19 A" 3469 3084 6.50      
20 A" 1717 1526 1.56      
21 A" 1395 1240 0.04      
22 A" 1256 1117 6.38      
23 A" 895 795 0.23      
24 A" 549 488 2.86      
25 A" 216 192 0.26      
26 A" 95 84 0.76      
27 A" 86 76 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 19782.6 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 17584.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.29359 0.06868 0.05688

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.122 -0.337 0.000
O2 0.000 0.676 0.000
O3 2.300 0.245 0.000
O4 0.816 -1.608 0.000
C5 -1.271 -0.074 0.000
C6 -2.403 0.998 0.000
H7 -1.351 -0.727 0.899
H8 -1.351 -0.727 -0.899
H9 -3.386 0.499 0.000
H10 -2.329 1.636 0.896
H11 -2.329 1.636 -0.896

Atom - Atom Distances (Å)
  N1 O2 O3 O4 C5 C6 H7 H8 H9 H10 H11
N11.51141.31361.30802.40733.76902.66022.66024.58514.07534.0753
O21.51142.34012.42531.47562.42402.14482.14483.39072.67412.6741
O31.31362.34012.37423.58514.76263.88353.88355.69194.91634.9163
O41.30802.42532.37422.58974.14142.50592.50594.70074.60674.6067
C52.40731.47563.58512.58971.55921.11371.11372.19162.20232.2023
C63.76902.42404.76264.14141.55922.21132.21131.10311.10291.1029
H72.66022.14483.88352.50591.11372.21131.79792.54032.55783.1250
H82.66022.14483.88352.50591.11372.21131.79792.54033.12502.5578
H94.58513.39075.69194.70072.19161.10312.54032.54031.79311.7931
H104.07532.67414.91634.60672.20231.10292.55783.12501.79311.7927
H114.07532.67414.91634.60672.20231.10293.12502.55781.79311.7927

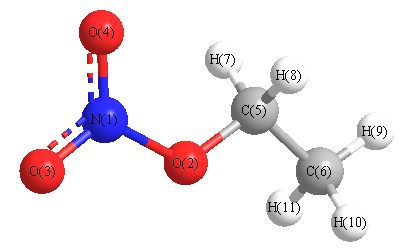
picture of Nitric acid, ethyl ester state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 O2 C5 107.396 O2 N1 O3 111.675
O2 N1 O4 118.506 O2 C5 C6 105.989
O2 C5 H7 111.085 O2 C5 H8 111.085
O3 N1 O4 129.819 C5 C6 H9 109.624
C5 C6 H10 110.473 C5 C6 H11 110.473
C6 C5 H7 110.539 C6 C5 H8 110.539
H7 C5 H8 107.639 H9 C6 H10 108.746
H9 C6 H11 108.746 H10 C6 H11 108.735
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability