Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Geometric Data calculated at MP2=FULL/STO-3G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -546.468636 |
| Energy at 298.15K | -546.473891 |
| HF Energy | -546.191821 |
| Nuclear repulsion energy | 198.962124 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3615 |
3213 |
0.16 |
|
|
|
| 2 |
A |
3612 |
3210 |
0.12 |
|
|
|
| 3 |
A |
3495 |
3107 |
0.38 |
|
|
|
| 4 |
A |
3493 |
3105 |
3.07 |
|
|
|
| 5 |
A |
3460 |
3075 |
13.25 |
|
|
|
| 6 |
A |
3454 |
3071 |
13.34 |
|
|
|
| 7 |
A |
1783 |
1585 |
15.99 |
|
|
|
| 8 |
A |
1774 |
1577 |
5.68 |
|
|
|
| 9 |
A |
1591 |
1414 |
5.99 |
|
|
|
| 10 |
A |
1574 |
1399 |
23.47 |
|
|
|
| 11 |
A |
1431 |
1272 |
3.17 |
|
|
|
| 12 |
A |
1420 |
1263 |
13.96 |
|
|
|
| 13 |
A |
1160 |
1031 |
7.52 |
|
|
|
| 14 |
A |
1135 |
1009 |
2.21 |
|
|
|
| 15 |
A |
1124 |
1000 |
9.77 |
|
|
|
| 16 |
A |
1116 |
992 |
10.82 |
|
|
|
| 17 |
A |
962 |
855 |
12.99 |
|
|
|
| 18 |
A |
951 |
845 |
14.55 |
|
|
|
| 19 |
A |
839 |
746 |
11.17 |
|
|
|
| 20 |
A |
793 |
705 |
1.18 |
|
|
|
| 21 |
A |
643 |
572 |
3.38 |
|
|
|
| 22 |
A |
627 |
558 |
2.38 |
|
|
|
| 23 |
A |
495 |
440 |
0.85 |
|
|
|
| 24 |
A |
386 |
343 |
0.14 |
|
|
|
| 25 |
A |
245 |
218 |
0.58 |
|
|
|
| 26 |
A |
125 |
111 |
0.97 |
|
|
|
| 27 |
A |
75 |
67 |
0.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20689.0 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 18390.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.043 |
-0.968 |
0.001 |
| C2 |
1.070 |
0.473 |
0.240 |
| C3 |
-1.586 |
-0.241 |
0.020 |
| C4 |
2.367 |
0.523 |
-0.135 |
| C5 |
-1.944 |
1.054 |
-0.125 |
| H6 |
0.627 |
1.325 |
0.780 |
| H7 |
-2.377 |
-0.997 |
0.148 |
| H8 |
2.988 |
1.400 |
0.091 |
| H9 |
2.856 |
-0.304 |
-0.667 |
| H10 |
-1.222 |
1.865 |
-0.279 |
| H11 |
-3.002 |
1.344 |
-0.089 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7855 | 1.7837 | 2.7651 | 2.8378 | 2.4910 | 2.4244 | 3.7802 | 2.9665 | 3.1149 | 3.8242 |
C2 | 1.7855 | | 2.7586 | 1.3513 | 3.0914 | 1.1018 | 3.7483 | 2.1356 | 2.1484 | 2.7313 | 4.1776 | C3 | 1.7837 | 2.7586 | | 4.0289 | 1.3512 | 2.8148 | 1.1022 | 4.8594 | 4.4947 | 2.1575 | 2.1280 | C4 | 2.7651 | 1.3513 | 4.0289 | | 4.3438 | 2.1232 | 4.9899 | 1.0978 | 1.0984 | 3.8339 | 5.4320 | C5 | 2.8378 | 3.0914 | 1.3512 | 4.3438 | | 2.7392 | 2.1140 | 4.9488 | 5.0178 | 1.0969 | 1.0976 | H6 | 2.4910 | 1.1018 | 2.8148 | 2.1232 | 2.7392 | | 3.8487 | 2.4605 | 3.1169 | 2.1982 | 3.7322 | H7 | 2.4244 | 3.7483 | 1.1022 | 4.9899 | 2.1140 | 3.8487 | | 5.8763 | 5.3411 | 3.1157 | 2.4343 | H8 | 3.7802 | 2.1356 | 4.8594 | 1.0978 | 4.9488 | 2.4605 | 5.8763 | | 1.8700 | 4.2512 | 5.9932 | H9 | 2.9665 | 2.1484 | 4.4947 | 1.0984 | 5.0178 | 3.1169 | 5.3411 | 1.8700 | | 4.6346 | 6.1128 | H10 | 3.1149 | 2.7313 | 2.1575 | 3.8339 | 1.0969 | 2.1982 | 3.1157 | 4.2512 | 4.6346 | | 1.8652 | H11 | 3.8242 | 4.1776 | 2.1280 | 5.4320 | 1.0976 | 3.7322 | 2.4343 | 5.9932 | 6.1128 | 1.8652 | |
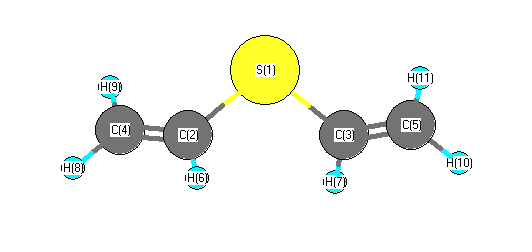 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
123.052 |
|
S1 |
C2 |
H6 |
117.278 |
| S1 |
C3 |
C5 |
129.191 |
|
S1 |
C3 |
H7 |
112.136 |
| C2 |
S1 |
C3 |
101.228 |
|
C2 |
C4 |
H8 |
121.027 |
| C2 |
C4 |
H9 |
122.224 |
|
C3 |
C5 |
H10 |
123.262 |
| C3 |
C5 |
H11 |
120.327 |
|
C4 |
C2 |
H6 |
119.542 |
| C5 |
C3 |
H7 |
118.657 |
|
H8 |
C4 |
H9 |
116.746 |
| H10 |
C5 |
H11 |
116.410 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
