Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -493.129125 |
| Energy at 298.15K | |
| HF Energy | -493.081886 |
| Nuclear repulsion energy | 43.647659 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3440 |
3058 |
0.58 |
|
|
|
| 2 |
A' |
1566 |
1392 |
25.51 |
|
|
|
| 3 |
A' |
928 |
825 |
28.61 |
|
|
|
| 4 |
A' |
766 |
681 |
8.43 |
|
|
|
| 5 |
A" |
3630 |
3227 |
0.07 |
|
|
|
| 6 |
A" |
1083 |
963 |
3.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5707.0 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 5072.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
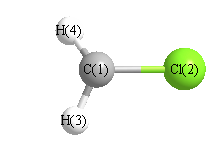
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.026 |
1.175 |
0.000 |
| Cl2 |
-0.026 |
-0.607 |
0.000 |
| H3 |
0.294 |
1.637 |
0.948 |
| H4 |
0.294 |
1.637 |
-0.948 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.7827 | 1.1012 | 1.1012 |
Cl2 | 1.7827 | | 2.4567 | 2.4567 | H3 | 1.1012 | 2.4567 | | 1.8952 | H4 | 1.1012 | 2.4567 | 1.8952 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
114.768 |
|
Br2 |
C1 |
H4 |
114.768 |
| H3 |
C1 |
H4 |
118.747 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -493.127166 |
| Energy at 298.15K | |
| HF Energy | -493.080585 |
| Nuclear repulsion energy | 43.757726 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3474 |
3088 |
0.97 |
|
|
|
| 2 |
A1 |
1544 |
1372 |
35.98 |
|
|
|
| 3 |
A1 |
938 |
834 |
25.15 |
|
|
|
| 4 |
B1 |
616i |
548i |
32.68 |
|
|
|
| 5 |
B2 |
3680 |
3271 |
1.77 |
|
|
|
| 6 |
B2 |
1039 |
924 |
3.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5029.7 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 4470.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.164 |
| Cl2 |
0.000 |
0.000 |
0.609 |
| H3 |
0.000 |
0.967 |
-1.679 |
| H4 |
0.000 |
-0.967 |
-1.679 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.7729 | 1.0958 | 1.0958 |
Cl2 | 1.7729 | | 2.4838 | 2.4838 | H3 | 1.0958 | 2.4838 | | 1.9346 | H4 | 1.0958 | 2.4838 | 1.9346 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
118.020 |
|
Br2 |
C1 |
H4 |
118.020 |
| H3 |
C1 |
H4 |
123.960 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
