Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -244.125272 |
| Energy at 298.15K | |
| HF Energy | -243.843157 |
| Nuclear repulsion energy | 176.656423 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3534 |
3141 |
1.92 |
|
|
|
| 2 |
A' |
3533 |
3140 |
0.27 |
|
|
|
| 3 |
A' |
3335 |
2964 |
14.32 |
|
|
|
| 4 |
A' |
3333 |
2962 |
17.89 |
|
|
|
| 5 |
A' |
3304 |
2937 |
33.81 |
|
|
|
| 6 |
A' |
1852 |
1646 |
153.53 |
|
|
|
| 7 |
A' |
1738 |
1545 |
3.85 |
|
|
|
| 8 |
A' |
1729 |
1537 |
8.48 |
|
|
|
| 9 |
A' |
1632 |
1450 |
25.21 |
|
|
|
| 10 |
A' |
1598 |
1420 |
16.53 |
|
|
|
| 11 |
A' |
1530 |
1360 |
43.29 |
|
|
|
| 12 |
A' |
1520 |
1351 |
21.05 |
|
|
|
| 13 |
A' |
1407 |
1251 |
20.20 |
|
|
|
| 14 |
A' |
1178 |
1047 |
2.14 |
|
|
|
| 15 |
A' |
1156 |
1028 |
112.72 |
|
|
|
| 16 |
A' |
918 |
816 |
5.62 |
|
|
|
| 17 |
A' |
663 |
589 |
2.10 |
|
|
|
| 18 |
A' |
386 |
343 |
0.49 |
|
|
|
| 19 |
A' |
273 |
243 |
4.99 |
|
|
|
| 20 |
A" |
3491 |
3104 |
4.81 |
|
|
|
| 21 |
A" |
3484 |
3097 |
9.89 |
|
|
|
| 22 |
A" |
1702 |
1513 |
3.46 |
|
|
|
| 23 |
A" |
1697 |
1508 |
3.15 |
|
|
|
| 24 |
A" |
1263 |
1122 |
9.13 |
|
|
|
| 25 |
A" |
1226 |
1090 |
0.03 |
|
|
|
| 26 |
A" |
963 |
856 |
0.97 |
|
|
|
| 27 |
A" |
275 |
245 |
8.42 |
|
|
|
| 28 |
A" |
186 |
166 |
0.20 |
|
|
|
| 29 |
A" |
179 |
159 |
3.68 |
|
|
|
| 30 |
A" |
159i |
141i |
9.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24461.8 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 21744.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.734 |
-0.855 |
0.000 |
| O2 |
0.208 |
-1.998 |
0.000 |
| N3 |
0.000 |
0.370 |
0.000 |
| C4 |
-1.481 |
0.333 |
0.000 |
| C5 |
0.669 |
1.690 |
0.000 |
| H6 |
1.836 |
-0.643 |
0.000 |
| H7 |
-1.786 |
-0.731 |
0.000 |
| H8 |
-1.903 |
0.830 |
0.899 |
| H9 |
-1.903 |
0.830 |
-0.899 |
| H10 |
1.765 |
1.539 |
0.000 |
| H11 |
0.396 |
2.284 |
-0.898 |
| H12 |
0.396 |
2.284 |
0.898 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.2581 | 1.4282 | 2.5136 | 2.5458 | 1.1225 | 2.5232 | 3.2557 | 3.2557 | 2.6067 | 3.2830 | 3.2830 |
O2 | 1.2581 | | 2.3772 | 2.8787 | 3.7166 | 2.1181 | 2.3627 | 3.6416 | 3.6416 | 3.8646 | 4.3797 | 4.3797 | N3 | 1.4282 | 2.3772 | | 1.4815 | 1.4798 | 2.0974 | 2.0982 | 2.1541 | 2.1541 | 2.1170 | 2.1513 | 2.1513 | C4 | 2.5136 | 2.8787 | 1.4815 | | 2.5425 | 3.4581 | 1.1068 | 1.1105 | 1.1105 | 3.4627 | 2.8525 | 2.8525 | C5 | 2.5458 | 3.7166 | 1.4798 | 2.5425 | | 2.6089 | 3.4479 | 2.8569 | 2.8569 | 1.1061 | 1.1112 | 1.1112 | H6 | 1.1225 | 2.1181 | 2.0974 | 3.4581 | 2.6089 | | 3.6235 | 4.1181 | 4.1181 | 2.1836 | 3.3844 | 3.3844 | H7 | 2.5232 | 2.3627 | 2.0982 | 1.1068 | 3.4479 | 3.6235 | | 1.8050 | 1.8050 | 4.2146 | 3.8287 | 3.8287 | H8 | 3.2557 | 3.6416 | 2.1541 | 1.1105 | 2.8569 | 4.1181 | 1.8050 | | 1.7984 | 3.8422 | 3.2600 | 2.7199 | H9 | 3.2557 | 3.6416 | 2.1541 | 1.1105 | 2.8569 | 4.1181 | 1.8050 | 1.7984 | | 3.8422 | 2.7199 | 3.2600 | H10 | 2.6067 | 3.8646 | 2.1170 | 3.4627 | 1.1061 | 2.1836 | 4.2146 | 3.8422 | 3.8422 | | 1.7991 | 1.7991 | H11 | 3.2830 | 4.3797 | 2.1513 | 2.8525 | 1.1112 | 3.3844 | 3.8287 | 3.2600 | 2.7199 | 1.7991 | | 1.7960 | H12 | 3.2830 | 4.3797 | 2.1513 | 2.8525 | 1.1112 | 3.3844 | 3.8287 | 2.7199 | 3.2600 | 1.7991 | 1.7960 | |
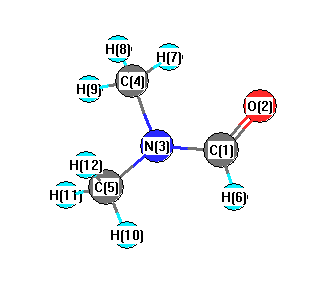 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
119.497 |
|
C1 |
N3 |
C5 |
122.190 |
| O2 |
C1 |
N3 |
124.364 |
|
O2 |
C1 |
H6 |
125.585 |
| N3 |
C1 |
H6 |
110.052 |
|
N3 |
C4 |
H7 |
107.430 |
| N3 |
C4 |
H8 |
111.615 |
|
N3 |
C4 |
H9 |
111.615 |
| N3 |
C5 |
H10 |
109.046 |
|
N3 |
C5 |
H11 |
111.469 |
| N3 |
C5 |
H12 |
111.469 |
|
C4 |
N3 |
C5 |
118.313 |
| H7 |
C4 |
H8 |
108.993 |
|
H7 |
C4 |
H9 |
108.993 |
| H8 |
C4 |
H9 |
108.140 |
|
H10 |
C5 |
H11 |
108.465 |
| H10 |
C5 |
H12 |
108.465 |
|
H11 |
C5 |
H12 |
107.835 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -244.127586 |
| Energy at 298.15K | -244.135628 |
| HF Energy | -243.843869 |
| Nuclear repulsion energy | 176.734744 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3550 |
3155 |
0.40 |
|
|
|
| 2 |
A |
3544 |
3150 |
0.99 |
|
|
|
| 3 |
A |
3504 |
3115 |
3.88 |
|
|
|
| 4 |
A |
3499 |
3110 |
8.37 |
|
|
|
| 5 |
A |
3340 |
2969 |
9.37 |
|
|
|
| 6 |
A |
3335 |
2965 |
13.39 |
|
|
|
| 7 |
A |
3310 |
2943 |
32.14 |
|
|
|
| 8 |
A |
1810 |
1609 |
93.82 |
|
|
|
| 9 |
A |
1730 |
1538 |
2.14 |
|
|
|
| 10 |
A |
1723 |
1532 |
2.36 |
|
|
|
| 11 |
A |
1705 |
1515 |
9.30 |
|
|
|
| 12 |
A |
1700 |
1511 |
5.55 |
|
|
|
| 13 |
A |
1627 |
1446 |
15.18 |
|
|
|
| 14 |
A |
1602 |
1424 |
7.68 |
|
|
|
| 15 |
A |
1510 |
1342 |
5.81 |
|
|
|
| 16 |
A |
1453 |
1292 |
35.57 |
|
|
|
| 17 |
A |
1361 |
1210 |
15.33 |
|
|
|
| 18 |
A |
1252 |
1113 |
15.96 |
|
|
|
| 19 |
A |
1206 |
1072 |
5.78 |
|
|
|
| 20 |
A |
1158 |
1029 |
1.58 |
|
|
|
| 21 |
A |
1127 |
1002 |
90.78 |
|
|
|
| 22 |
A |
971 |
863 |
0.82 |
|
|
|
| 23 |
A |
893 |
794 |
19.22 |
|
|
|
| 24 |
A |
649 |
577 |
1.54 |
|
|
|
| 25 |
A |
401 |
356 |
0.38 |
|
|
|
| 26 |
A |
315 |
280 |
18.42 |
|
|
|
| 27 |
A |
285 |
253 |
5.15 |
|
|
|
| 28 |
A |
226 |
201 |
1.63 |
|
|
|
| 29 |
A |
197 |
175 |
2.25 |
|
|
|
| 30 |
A |
130 |
116 |
1.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24555.7 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 21827.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.877 |
-0.683 |
-0.057 |
| O2 |
-1.973 |
-0.090 |
0.113 |
| N3 |
0.389 |
-0.014 |
-0.345 |
| C4 |
0.407 |
1.439 |
0.033 |
| C5 |
1.608 |
-0.752 |
0.121 |
| H6 |
-0.751 |
-1.797 |
-0.106 |
| H7 |
1.222 |
1.952 |
-0.512 |
| H8 |
-0.563 |
1.885 |
-0.255 |
| H9 |
0.558 |
1.598 |
1.122 |
| H10 |
2.502 |
-0.335 |
-0.382 |
| H11 |
1.764 |
-0.683 |
1.220 |
| H12 |
1.513 |
-1.819 |
-0.155 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.2576 | 1.4607 | 2.4816 | 2.4925 | 1.1225 | 3.3997 | 2.5949 | 2.9413 | 3.4124 | 2.9342 | 2.6489 |
O2 | 1.2576 | | 2.4071 | 2.8297 | 3.6417 | 2.1111 | 3.8432 | 2.4547 | 3.2048 | 4.5088 | 3.9429 | 3.9011 | N3 | 1.4607 | 2.4071 | | 1.5009 | 1.4997 | 2.1300 | 2.1416 | 2.1259 | 2.1861 | 2.1376 | 2.1888 | 2.1358 | C4 | 2.4816 | 2.8297 | 1.5009 | | 2.5002 | 3.4396 | 1.1067 | 1.1057 | 1.1116 | 2.7757 | 2.7849 | 3.4459 | C5 | 2.4925 | 3.6417 | 1.4997 | 2.5002 | | 2.5899 | 2.8041 | 3.4366 | 2.7620 | 1.1071 | 1.1123 | 1.1059 | H6 | 1.1225 | 2.1111 | 2.1300 | 3.4396 | 2.5899 | | 4.2561 | 3.6902 | 3.8402 | 3.5770 | 3.0541 | 2.2651 | H7 | 3.3997 | 3.8432 | 2.1416 | 1.1067 | 2.8041 | 4.2561 | | 1.8047 | 1.7996 | 2.6235 | 3.1997 | 3.7994 | H8 | 2.5949 | 2.4547 | 2.1259 | 1.1057 | 3.4366 | 3.6902 | 1.8047 | | 1.7989 | 3.7862 | 3.7669 | 4.2479 | H9 | 2.9413 | 3.2048 | 2.1861 | 1.1116 | 2.7620 | 3.8402 | 1.7996 | 1.7989 | | 3.1264 | 2.5824 | 3.7709 | H10 | 3.4124 | 4.5088 | 2.1376 | 2.7757 | 1.1071 | 3.5770 | 2.6235 | 3.7862 | 3.1264 | | 1.7969 | 1.7978 | H11 | 2.9342 | 3.9429 | 2.1888 | 2.7849 | 1.1123 | 3.0541 | 3.1997 | 3.7669 | 2.5824 | 1.7969 | | 1.8007 | H12 | 2.6489 | 3.9011 | 2.1358 | 3.4459 | 1.1059 | 2.2651 | 3.7994 | 4.2479 | 3.7709 | 1.7978 | 1.8007 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
113.838 |
|
C1 |
N3 |
C5 |
114.680 |
| O2 |
C1 |
N3 |
124.462 |
|
O2 |
C1 |
H6 |
124.900 |
| N3 |
C1 |
H6 |
110.401 |
|
N3 |
C4 |
H7 |
109.502 |
| N3 |
C4 |
H8 |
108.338 |
|
N3 |
C4 |
H9 |
112.757 |
| N3 |
C5 |
H10 |
109.241 |
|
N3 |
C5 |
H11 |
113.017 |
| N3 |
C5 |
H12 |
109.171 |
|
C4 |
N3 |
C5 |
112.864 |
| H7 |
C4 |
H8 |
109.318 |
|
H7 |
C4 |
H9 |
108.431 |
| H8 |
C4 |
H9 |
108.446 |
|
H10 |
C5 |
H11 |
108.119 |
| H10 |
C5 |
H12 |
108.663 |
|
H11 |
C5 |
H12 |
108.540 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
