return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SCH3 (Dimethyl sulfide)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-471.616575
Energy at 298.15K 
HF Energy-471.480537
Nuclear repulsion energy110.125953
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3540 3146 2.76 42.06 0.70 0.82
2 A1 3346 2974 3.95 69.93 0.01 0.01
3 A1 1743 1549 0.03 27.74 0.74 0.85
4 A1 1610 1431 0.96 3.22 0.68 0.81
5 A1 1171 1041 11.07 6.66 0.63 0.77
6 A1 858 763 0.24 14.38 0.17 0.29
7 A1 317 282 0.23 1.04 0.57 0.72
8 A2 3516 3125 0.00 4.41 0.75 0.86
9 A2 1726 1534 0.00 35.07 0.75 0.86
10 A2 1084 964 0.00 4.47 0.75 0.86
11 A2 181 161 0.00 0.07 0.75 0.86
12 B1 3513 3123 13.64 42.74 0.75 0.86
13 B1 1726 1534 4.57 0.50 0.75 0.86
14 B1 1112 988 0.22 4.14 0.75 0.86
15 B1 182 162 0.47 0.00 0.75 0.86
16 B2 3539 3146 0.24 20.83 0.75 0.86
17 B2 3348 2976 5.03 0.60 0.75 0.86
18 B2 1741 1548 6.11 0.11 0.75 0.86
19 B2 1585 1409 17.56 5.01 0.75 0.86
20 B2 1047 930 0.01 1.00 0.75 0.86
21 B2 912 811 0.21 8.47 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 18898.6 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 16799.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.56553 0.25579 0.18882

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 0.681
C2 0.000 1.363 -0.522
C3 0.000 -1.363 -0.522
H4 0.000 2.319 0.028
H5 0.000 -2.319 0.028
H6 0.893 1.345 -1.173
H7 -0.893 1.345 -1.173
H8 -0.893 -1.345 -1.173
H9 0.893 -1.345 -1.173

Atom - Atom Distances (Å)
  S1 C2 C3 H4 H5 H6 H7 H8 H9
S11.81781.81782.40942.40942.45882.45882.45882.4588
C21.81782.72511.10323.72241.10551.10552.92442.9244
C31.81782.72513.72241.10322.92442.92441.10551.1055
H42.40941.10323.72244.63821.78571.78573.95783.9578
H52.40943.72241.10324.63823.95783.95781.78571.7857
H62.45881.10552.92441.78573.95781.78643.22892.6897
H72.45881.10552.92441.78573.95781.78642.68973.2289
H82.45882.92441.10553.95781.78573.22892.68971.7864
H92.45882.92441.10553.95781.78572.68973.22891.7864

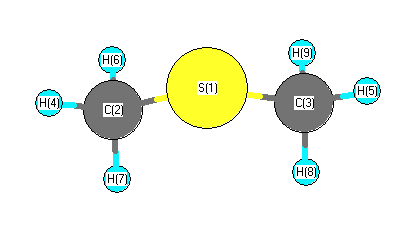
picture of Dimethyl sulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C2 H4 108.671 S1 C2 H6 112.207
S1 C2 H7 112.207 S1 C3 H5 108.671
S1 C3 H8 112.207 S1 C3 H9 112.207
C2 S1 C3 97.102 H4 C2 H6 107.898
H4 C2 H7 107.898 H5 C3 H8 107.898
H5 C3 H9 107.898 H6 C2 H7 107.787
H8 C3 H9 107.787
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability