return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHClCH3 (Propane, 2-chloro-)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-571.058591
Energy at 298.15K-571.066099
HF Energy-570.886964
Nuclear repulsion energy162.447016
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3569 3172 0.82 29.64 0.74 0.85
2 A' 3559 3163 0.54 50.56 0.75 0.85
3 A' 3441 3059 2.00 34.64 0.39 0.56
4 A' 3379 3004 0.81 71.11 0.01 0.02
5 A' 1724 1532 1.71 6.20 0.75 0.86
6 A' 1716 1525 1.99 21.50 0.75 0.86
7 A' 1612 1433 0.22 3.70 0.49 0.66
8 A' 1440 1280 44.63 6.40 0.57 0.73
9 A' 1319 1173 12.03 3.26 0.73 0.84
10 A' 1200 1067 10.68 5.38 0.62 0.76
11 A' 1006 894 4.39 10.14 0.73 0.84
12 A' 750 666 10.48 14.30 0.22 0.36
13 A' 437 389 1.07 1.23 0.29 0.45
14 A' 340 303 0.81 0.76 0.59 0.74
15 A' 244 216 0.09 0.03 0.70 0.82
16 A" 3568 3171 0.15 13.43 0.75 0.86
17 A" 3558 3163 0.20 6.55 0.75 0.86
18 A" 3380 3004 0.64 1.49 0.75 0.86
19 A" 1716 1525 0.33 13.43 0.75 0.86
20 A" 1712 1522 0.16 15.05 0.75 0.86
21 A" 1602 1424 0.08 6.00 0.75 0.86
22 A" 1505 1338 8.51 5.18 0.75 0.86
23 A" 1259 1119 4.37 6.98 0.75 0.86
24 A" 1070 951 0.28 3.60 0.75 0.86
25 A" 1044 928 0.73 0.03 0.75 0.86
26 A" 325 289 2.67 1.88 0.75 0.86
27 A" 225 200 0.00 0.01 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 23348.4 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 20754.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.25790 0.14471 0.10178

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 -0.900 1.049 0.000
C2 0.576 -0.065 0.000
C3 0.576 -0.932 1.299
C4 0.576 -0.932 -1.299
H5 1.457 0.612 0.000
H6 1.482 -1.563 1.316
H7 1.482 -1.563 -1.316
H8 0.574 -0.288 2.195
H9 0.574 -0.288 -2.195
H10 -0.313 -1.585 1.328
H11 -0.313 -1.585 -1.328

Atom - Atom Distances (Å)
  Cl1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11
Cl11.84942.79092.79092.39703.77163.77162.96292.96293.00723.0072
C21.84941.56141.56141.11112.18942.18942.20602.20602.20462.2046
C32.79091.56142.59802.20151.10402.83831.10283.55251.10282.8488
C42.79091.56142.59802.20152.83831.10403.55251.10282.84881.1028
H52.39701.11112.20152.20152.54192.54192.53122.53123.11743.1174
H63.77162.18941.10402.83832.54192.63161.79443.84341.79443.1951
H73.77162.18942.83831.10402.54192.63163.84341.79443.19511.7944
H82.96292.20601.10283.55252.53121.79443.84344.38951.79373.8568
H92.96292.20603.55251.10282.53123.84341.79444.38953.85681.7937
H103.00722.20461.10282.84883.11741.79443.19511.79373.85682.6557
H113.00722.20462.84881.10283.11743.19511.79443.85681.79372.6557

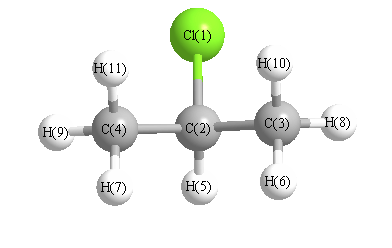
picture of Propane, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C2 C3 109.527 Cl1 C2 C4 109.527
Cl1 C3 H5 55.898 C2 C3 H6 109.252
C2 C3 H8 110.612 C2 C3 H10 110.503
C2 C4 H7 109.252 C2 C4 H9 110.612
C2 C4 H11 110.503 C3 C2 C4 112.597
C3 C2 H5 109.773 C4 C2 H5 109.773
H6 C3 H8 108.805 H6 C3 H10 108.799
H7 C4 H9 108.805 H7 C4 H11 108.799
H8 C3 H10 108.829 H9 C4 H11 108.829
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability