return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(CH3)3SH (2-Propanethiol, 2-methyl-)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-548.876858
Energy at 298.15K-548.887397
HF Energy-548.631728
Nuclear repulsion energy241.106979
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3549 3155 4.38      
2 A' 3549 3155 2.67      
3 A' 3543 3149 6.33      
4 A' 3368 2994 3.80      
5 A' 3363 2989 3.05      
6 A' 3081 2739 27.31      
7 A' 1741 1548 0.99      
8 A' 1736 1543 1.78      
9 A' 1730 1538 0.12      
10 A' 1628 1447 0.81      
11 A' 1610 1431 0.38      
12 A' 1414 1257 4.15      
13 A' 1389 1235 30.20      
14 A' 1185 1054 3.68      
15 A' 1075 956 5.68      
16 A' 1037 922 1.68      
17 A' 953 847 0.03      
18 A' 686 610 0.11      
19 A' 408 363 0.49      
20 A' 383 340 0.03      
21 A' 301 267 0.46      
22 A' 267 237 0.08      
23 A" 3550 3156 3.96      
24 A" 3548 3154 0.90      
25 A" 3542 3148 0.04      
26 A" 3364 2990 3.83      
27 A" 1737 1544 0.90      
28 A" 1730 1538 0.00      
29 A" 1728 1536 0.00      
30 A" 1608 1429 0.33      
31 A" 1385 1231 14.21      
32 A" 1160 1031 0.22      
33 A" 1085 964 0.00      
34 A" 1046 929 0.24      
35 A" 411 365 0.41      
36 A" 311 276 1.53      
37 A" 262 233 0.49      
38 A" 251 223 5.82      
39 A" 221 196 3.43      

Unscaled Zero Point Vibrational Energy (zpe) 32466.9 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 28859.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.14445 0.09668 0.09572

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.318 -0.000 0.000
S2 -1.524 0.082 0.000
C3 0.850 1.477 0.000
C4 0.850 -0.741 1.280
C5 0.850 -0.741 -1.280
H6 -1.685 -1.258 0.000
H7 1.954 1.487 0.000
H8 0.499 2.020 -0.895
H9 0.499 2.020 0.895
H10 1.954 -0.754 1.287
H11 1.954 -0.754 -1.287
H12 0.503 -0.235 2.198
H13 0.494 -1.786 1.306
H14 0.503 -0.235 -2.198
H15 0.494 -1.786 -1.306

Atom - Atom Distances (Å)
  C1 S2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.84391.57011.57201.57202.36532.21042.21682.21682.21382.21382.21812.21932.21812.2193
S21.84392.75332.82012.82011.35013.75042.94052.94053.80163.80163.00683.04433.00683.0443
C31.57012.75332.56102.56103.72921.10361.10371.10372.80242.80242.80743.53262.80743.5326
C41.57202.82012.56102.56082.88672.79643.53222.80971.10412.79461.10371.10423.53202.8123
C51.57202.82012.56102.56082.88672.79642.80973.53222.79461.10413.53202.81231.10371.1042
H62.36531.35013.72922.88672.88674.55774.03924.03923.89263.89263.26582.59453.26582.5945
H72.21043.75041.10362.79642.79644.55771.78891.78892.58422.58423.14623.81403.14623.8140
H82.21682.94051.10373.53222.80974.03921.78891.78933.81733.15703.82734.39632.60443.8279
H92.21682.94051.10372.80973.53224.03921.78891.78933.15703.81732.60443.82793.82734.3963
H102.21383.80162.80241.10412.79463.89262.58423.81733.15702.57371.79011.78803.81023.1496
H112.21383.80162.80242.79461.10413.89262.58423.15703.81732.57373.81023.14961.79011.7880
H122.21813.00682.80741.10373.53203.26583.14623.82732.60441.79013.81021.78904.39583.8319
H132.21933.04433.53261.10422.81232.59453.81404.39633.82791.78803.14961.78903.83192.6124
H142.21813.00682.80743.53201.10373.26583.14622.60443.82733.81021.79014.39583.83191.7890
H152.21933.04433.53262.81231.10422.59453.81403.82794.39633.14961.78803.83192.61241.7890

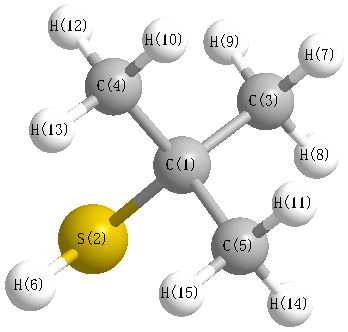
picture of 2-Propanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 H6 94.280 C1 C3 H7 110.307
C1 C3 H8 110.799 C1 C3 H9 110.799
C1 C4 H10 110.413 C1 C4 H12 110.777
C1 C4 H13 110.838 C1 C5 H11 110.413
C1 C5 H14 110.777 C1 C5 H15 110.838
S2 C1 C3 107.232 S2 C1 C4 111.049
S2 C1 C5 111.049 C3 C1 C4 109.191
C3 C1 C5 109.191 C4 C1 C5 109.081
H7 C3 H8 108.276 H7 C3 H9 108.276
H8 C3 H9 108.297 H10 C4 H12 108.353
H10 C4 H13 108.123 H11 C5 H14 108.353
H11 C5 H15 108.123 H12 C4 H13 108.243
H14 C5 H15 108.243
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability