Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -629.881733 |
| Energy at 298.15K | -629.886465 |
| HF Energy | -629.748805 |
| Nuclear repulsion energy | 152.900859 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3409 |
3031 |
0.23 |
|
|
|
| 2 |
A' |
3352 |
2979 |
4.99 |
|
|
|
| 3 |
A' |
1708 |
1518 |
0.01 |
|
|
|
| 4 |
A' |
1674 |
1488 |
1.79 |
|
|
|
| 5 |
A' |
1564 |
1390 |
5.16 |
|
|
|
| 6 |
A' |
1403 |
1247 |
27.32 |
|
|
|
| 7 |
A' |
1256 |
1117 |
7.59 |
|
|
|
| 8 |
A' |
1119 |
994 |
1.09 |
|
|
|
| 9 |
A' |
913 |
812 |
20.44 |
|
|
|
| 10 |
A' |
393 |
349 |
0.80 |
|
|
|
| 11 |
A' |
247 |
219 |
7.15 |
|
|
|
| 12 |
A" |
3533 |
3140 |
0.24 |
|
|
|
| 13 |
A" |
3460 |
3076 |
5.64 |
|
|
|
| 14 |
A" |
1419 |
1262 |
0.13 |
|
|
|
| 15 |
A" |
1303 |
1158 |
5.62 |
|
|
|
| 16 |
A" |
1141 |
1014 |
0.09 |
|
|
|
| 17 |
A" |
834 |
741 |
0.11 |
|
|
|
| 18 |
A" |
100 |
89 |
5.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14413.0 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 12811.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.635 |
0.000 |
| C2 |
1.029 |
-0.559 |
0.000 |
| Cl3 |
-1.717 |
-0.002 |
0.000 |
| F4 |
2.341 |
-0.063 |
0.000 |
| H5 |
0.126 |
1.266 |
0.903 |
| H6 |
0.126 |
1.266 |
-0.903 |
| H7 |
0.851 |
-1.192 |
0.902 |
| H8 |
0.851 |
-1.192 |
-0.902 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5765 | 1.8318 | 2.4426 | 1.1085 | 1.1085 | 2.2086 | 2.2086 |
C2 | 1.5765 | | 2.8020 | 1.4025 | 2.2270 | 2.2270 | 1.1165 | 1.1165 | Cl3 | 1.8318 | 2.8020 | | 4.0583 | 2.4130 | 2.4130 | 2.9706 | 2.9706 | F4 | 2.4426 | 1.4025 | 4.0583 | | 2.7356 | 2.7356 | 2.0758 | 2.0758 | H5 | 1.1085 | 2.2270 | 2.4130 | 2.7356 | | 1.8063 | 2.5624 | 3.1345 | H6 | 1.1085 | 2.2270 | 2.4130 | 2.7356 | 1.8063 | | 3.1345 | 2.5624 | H7 | 2.2086 | 1.1165 | 2.9706 | 2.0758 | 2.5624 | 3.1345 | | 1.8045 | H8 | 2.2086 | 1.1165 | 2.9706 | 2.0758 | 3.1345 | 2.5624 | 1.8045 | |
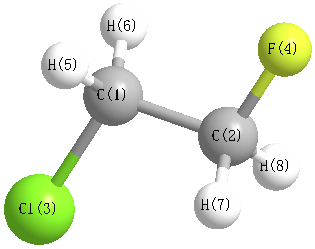 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.022 |
|
C1 |
C2 |
H7 |
108.998 |
| C1 |
C2 |
H8 |
108.998 |
|
C2 |
C1 |
Cl3 |
110.367 |
| C2 |
C1 |
H5 |
110.874 |
|
C2 |
C1 |
H6 |
110.874 |
| Cl3 |
C1 |
H5 |
107.751 |
|
Cl3 |
C1 |
H6 |
107.751 |
| F4 |
C2 |
H7 |
110.476 |
|
F4 |
C2 |
H8 |
110.476 |
| H5 |
C1 |
H6 |
109.117 |
|
H7 |
C2 |
H8 |
107.818 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -629.881642 |
| Energy at 298.15K | -629.886455 |
| HF Energy | -629.748572 |
| Nuclear repulsion energy | 157.035534 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3532 |
3140 |
0.06 |
|
|
|
| 2 |
A |
3449 |
3066 |
4.93 |
|
|
|
| 3 |
A |
3408 |
3029 |
0.04 |
|
|
|
| 4 |
A |
3339 |
2968 |
6.38 |
|
|
|
| 5 |
A |
1697 |
1509 |
0.46 |
|
|
|
| 6 |
A |
1658 |
1473 |
2.06 |
|
|
|
| 7 |
A |
1565 |
1391 |
18.16 |
|
|
|
| 8 |
A |
1470 |
1307 |
30.26 |
|
|
|
| 9 |
A |
1365 |
1214 |
0.50 |
|
|
|
| 10 |
A |
1317 |
1170 |
5.75 |
|
|
|
| 11 |
A |
1250 |
1111 |
4.09 |
|
|
|
| 12 |
A |
1144 |
1017 |
0.15 |
|
|
|
| 13 |
A |
1064 |
946 |
2.78 |
|
|
|
| 14 |
A |
927 |
824 |
1.20 |
|
|
|
| 15 |
A |
802 |
713 |
13.75 |
|
|
|
| 16 |
A |
477 |
424 |
6.20 |
|
|
|
| 17 |
A |
266 |
237 |
0.49 |
|
|
|
| 18 |
A |
99 |
88 |
1.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14413.5 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 12812.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.108 |
0.882 |
-0.293 |
| C2 |
1.249 |
0.417 |
0.353 |
| Cl3 |
-1.462 |
-0.300 |
0.066 |
| F4 |
1.703 |
-0.784 |
-0.205 |
| H5 |
-0.405 |
1.869 |
0.116 |
| H6 |
-0.015 |
0.955 |
-1.395 |
| H7 |
1.996 |
1.231 |
0.183 |
| H8 |
1.108 |
0.306 |
1.455 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5737 | 1.8332 | 2.4632 | 1.1084 | 1.1086 | 2.1855 | 2.2063 |
C2 | 1.5737 | | 2.8194 | 1.4002 | 2.2141 | 2.2234 | 1.1183 | 1.1167 | Cl3 | 1.8332 | 2.8194 | | 3.2142 | 2.4132 | 2.4093 | 3.7843 | 2.9840 | F4 | 2.4632 | 1.4002 | 3.2142 | | 3.4042 | 2.7187 | 2.0735 | 2.0736 | H5 | 1.1084 | 2.2141 | 2.4132 | 3.4042 | | 1.8089 | 2.4855 | 2.5548 | H6 | 1.1086 | 2.2234 | 2.4093 | 2.7187 | 1.8089 | | 2.5711 | 3.1316 | H7 | 2.1855 | 1.1183 | 3.7843 | 2.0735 | 2.4855 | 2.5711 | | 1.8068 | H8 | 2.2063 | 1.1167 | 2.9840 | 2.0736 | 2.5548 | 3.1316 | 1.8068 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
111.709 |
|
C1 |
C2 |
H7 |
107.340 |
| C1 |
C2 |
H8 |
109.000 |
|
C2 |
C1 |
Cl3 |
111.469 |
| C2 |
C1 |
H5 |
110.066 |
|
C2 |
C1 |
H6 |
110.779 |
| Cl3 |
C1 |
H5 |
107.679 |
|
Cl3 |
C1 |
H6 |
107.391 |
| F4 |
C2 |
H7 |
110.335 |
|
F4 |
C2 |
H8 |
110.445 |
| H5 |
C1 |
H6 |
109.358 |
|
H7 |
C2 |
H8 |
107.884 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
