Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Geometric Data calculated at MP2=FULL/STO-3G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -231.735915 |
| Energy at 298.15K | -231.747871 |
| HF Energy | -231.397238 |
| Nuclear repulsion energy | 226.644046 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3563 |
3167 |
2.13 |
|
|
|
| 2 |
A |
3554 |
3159 |
4.98 |
|
|
|
| 3 |
A |
3553 |
3158 |
3.46 |
|
|
|
| 4 |
A |
3528 |
3136 |
2.13 |
|
|
|
| 5 |
A |
3523 |
3131 |
7.05 |
|
|
|
| 6 |
A |
3512 |
3122 |
0.19 |
|
|
|
| 7 |
A |
3502 |
3113 |
3.78 |
|
|
|
| 8 |
A |
3482 |
3096 |
5.01 |
|
|
|
| 9 |
A |
3405 |
3027 |
8.29 |
|
|
|
| 10 |
A |
3398 |
3021 |
1.32 |
|
|
|
| 11 |
A |
3373 |
2998 |
3.76 |
|
|
|
| 12 |
A |
3368 |
2994 |
2.64 |
|
|
|
| 13 |
A |
1848 |
1643 |
0.50 |
|
|
|
| 14 |
A |
1745 |
1551 |
0.30 |
|
|
|
| 15 |
A |
1737 |
1544 |
1.47 |
|
|
|
| 16 |
A |
1731 |
1539 |
0.81 |
|
|
|
| 17 |
A |
1725 |
1534 |
0.45 |
|
|
|
| 18 |
A |
1725 |
1533 |
1.31 |
|
|
|
| 19 |
A |
1710 |
1520 |
0.36 |
|
|
|
| 20 |
A |
1625 |
1444 |
0.38 |
|
|
|
| 21 |
A |
1624 |
1443 |
0.03 |
|
|
|
| 22 |
A |
1586 |
1410 |
8.91 |
|
|
|
| 23 |
A |
1554 |
1381 |
5.49 |
|
|
|
| 24 |
A |
1471 |
1307 |
0.04 |
|
|
|
| 25 |
A |
1448 |
1287 |
22.31 |
|
|
|
| 26 |
A |
1416 |
1259 |
0.07 |
|
|
|
| 27 |
A |
1384 |
1230 |
0.07 |
|
|
|
| 28 |
A |
1294 |
1150 |
1.11 |
|
|
|
| 29 |
A |
1239 |
1102 |
0.33 |
|
|
|
| 30 |
A |
1187 |
1055 |
0.02 |
|
|
|
| 31 |
A |
1174 |
1043 |
0.08 |
|
|
|
| 32 |
A |
1170 |
1040 |
0.78 |
|
|
|
| 33 |
A |
1101 |
979 |
0.71 |
|
|
|
| 34 |
A |
1091 |
970 |
0.08 |
|
|
|
| 35 |
A |
1009 |
897 |
2.21 |
|
|
|
| 36 |
A |
999 |
888 |
1.05 |
|
|
|
| 37 |
A |
959 |
852 |
1.42 |
|
|
|
| 38 |
A |
817 |
727 |
1.84 |
|
|
|
| 39 |
A |
764 |
679 |
10.88 |
|
|
|
| 40 |
A |
605 |
538 |
3.93 |
|
|
|
| 41 |
A |
497 |
441 |
0.32 |
|
|
|
| 42 |
A |
382 |
340 |
0.03 |
|
|
|
| 43 |
A |
298 |
265 |
0.01 |
|
|
|
| 44 |
A |
237 |
211 |
0.00 |
|
|
|
| 45 |
A |
208 |
184 |
0.02 |
|
|
|
| 46 |
A |
103 |
91 |
0.07 |
|
|
|
| 47 |
A |
89 |
79 |
0.00 |
|
|
|
| 48 |
A |
38 |
34 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40674.6 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 36155.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.469 |
0.998 |
-0.055 |
| C2 |
2.024 |
-0.470 |
-0.160 |
| C3 |
0.801 |
-0.978 |
0.118 |
| C4 |
-0.447 |
-0.209 |
0.593 |
| C5 |
-1.583 |
-0.219 |
-0.489 |
| C6 |
-2.866 |
0.519 |
0.001 |
| H7 |
1.654 |
1.654 |
0.287 |
| H8 |
3.313 |
1.096 |
0.653 |
| H9 |
2.818 |
1.365 |
-1.038 |
| H10 |
2.807 |
-1.167 |
-0.500 |
| H11 |
0.651 |
-2.061 |
-0.020 |
| H12 |
-0.191 |
0.837 |
0.841 |
| H13 |
-0.834 |
-0.681 |
1.517 |
| H14 |
-1.211 |
0.263 |
-1.413 |
| H15 |
-1.835 |
-1.266 |
-0.744 |
| H16 |
-3.649 |
0.497 |
-0.776 |
| H17 |
-3.269 |
0.040 |
0.910 |
| H18 |
-2.644 |
1.573 |
0.238 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5371 | 2.5919 | 3.2215 | 4.2530 | 5.3572 | 1.1015 | 1.1053 | 1.1053 | 2.2361 | 3.5590 | 2.8119 | 4.0250 | 3.9914 | 4.9116 | 6.1808 | 5.8970 | 5.1539 |
C2 | 1.5371 | | 1.3536 | 2.5958 | 3.6303 | 4.9915 | 2.2021 | 2.1844 | 2.1835 | 1.1024 | 2.1067 | 2.7597 | 3.3204 | 3.5460 | 3.9830 | 5.7875 | 5.4237 | 5.1109 | C3 | 2.5919 | 1.3536 | | 1.5403 | 2.5737 | 3.9620 | 2.7725 | 3.3010 | 3.3009 | 2.1077 | 1.1025 | 2.1911 | 2.1722 | 2.8162 | 2.7874 | 4.7720 | 4.2686 | 4.2881 | C4 | 3.2215 | 2.5958 | 1.5403 | | 1.5686 | 2.5947 | 2.8244 | 3.9795 | 3.9741 | 3.5634 | 2.2390 | 1.1049 | 1.1082 | 2.1979 | 2.1978 | 3.5533 | 2.8507 | 2.8512 | C5 | 4.2530 | 3.6303 | 2.5737 | 1.5686 | | 1.5593 | 3.8195 | 5.1957 | 4.7095 | 4.4909 | 2.9335 | 2.1954 | 2.1909 | 1.1067 | 1.1067 | 2.2055 | 2.2059 | 2.2061 | C6 | 5.3572 | 4.9915 | 3.9620 | 2.5947 | 1.5593 | | 4.6693 | 6.2396 | 5.8401 | 5.9394 | 4.3622 | 2.8216 | 2.8051 | 2.1919 | 2.1918 | 1.1035 | 1.1034 | 1.1034 | H7 | 1.1015 | 2.2021 | 2.7725 | 2.8244 | 3.8195 | 4.6693 | | 1.7879 | 1.7873 | 3.1484 | 3.8612 | 2.0926 | 3.6270 | 3.6108 | 4.6651 | 5.5309 | 5.2180 | 4.2990 | H8 | 1.1053 | 2.1844 | 3.3010 | 3.9795 | 5.1957 | 6.2396 | 1.7879 | | 1.7816 | 2.5897 | 4.1837 | 3.5182 | 4.5929 | 5.0425 | 5.8328 | 7.1317 | 6.6704 | 5.9900 | H9 | 1.1053 | 2.1835 | 3.3009 | 3.9741 | 4.7095 | 5.8401 | 1.7873 | 1.7816 | | 2.5891 | 4.1802 | 3.5865 | 4.9042 | 4.1944 | 5.3535 | 6.5303 | 6.5269 | 5.6130 | H10 | 2.2361 | 1.1024 | 2.1077 | 3.5634 | 4.4909 | 5.9394 | 3.1484 | 2.5897 | 2.5891 | | 2.3826 | 3.8476 | 4.1905 | 4.3618 | 4.6489 | 6.6725 | 6.3526 | 6.1455 | H11 | 3.5590 | 2.1067 | 1.1025 | 2.2390 | 2.9335 | 4.3622 | 3.8612 | 4.1837 | 4.1802 | 2.3826 | | 3.1388 | 2.5445 | 3.2884 | 2.7085 | 5.0605 | 4.5436 | 4.9128 | H12 | 2.8119 | 2.7597 | 2.1911 | 1.1049 | 2.1954 | 2.8216 | 2.0926 | 3.5182 | 3.5865 | 3.8476 | 3.1388 | | 1.7818 | 2.5399 | 3.1040 | 3.8323 | 3.1799 | 2.6309 | H13 | 4.0250 | 3.3204 | 2.1722 | 1.1082 | 2.1909 | 2.8051 | 3.6270 | 4.5929 | 4.9042 | 4.1905 | 2.5445 | 1.7818 | | 3.1020 | 2.5411 | 3.8176 | 2.6109 | 3.1615 | H14 | 3.9914 | 3.5460 | 2.8162 | 2.1979 | 1.1067 | 2.1919 | 3.6108 | 5.0425 | 4.1944 | 4.3618 | 3.2884 | 2.5399 | 3.1020 | | 1.7819 | 2.5302 | 3.1113 | 2.5487 | H15 | 4.9116 | 3.9830 | 2.7874 | 2.1978 | 1.1067 | 2.1918 | 4.6651 | 5.8328 | 5.3535 | 4.6489 | 2.7085 | 3.1040 | 2.5411 | 1.7819 | | 2.5300 | 2.5486 | 3.1113 | H16 | 6.1808 | 5.7875 | 4.7720 | 3.5533 | 2.2055 | 1.1035 | 5.5309 | 7.1317 | 6.5303 | 6.6725 | 5.0605 | 3.8323 | 3.8176 | 2.5302 | 2.5300 | | 1.7881 | 1.7879 | H17 | 5.8970 | 5.4237 | 4.2686 | 2.8507 | 2.2059 | 1.1034 | 5.2180 | 6.6704 | 6.5269 | 6.3526 | 4.5436 | 3.1799 | 2.6109 | 3.1113 | 2.5486 | 1.7881 | | 1.7872 | H18 | 5.1539 | 5.1109 | 4.2881 | 2.8512 | 2.2061 | 1.1034 | 4.2990 | 5.9900 | 5.6130 | 6.1455 | 4.9128 | 2.6309 | 3.1615 | 2.5487 | 3.1113 | 1.7879 | 1.7872 | |
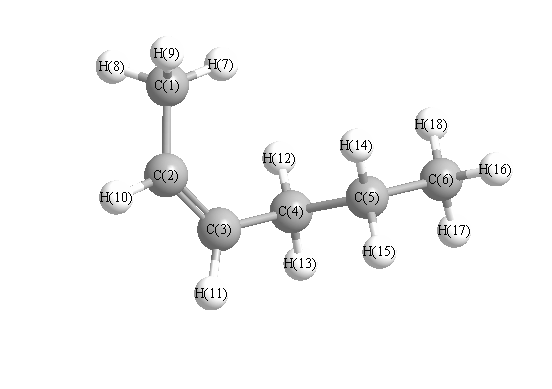 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.325 |
|
C1 |
C2 |
H10 |
114.808 |
| C2 |
C1 |
H7 |
112.079 |
|
C2 |
C1 |
H8 |
110.446 |
| C2 |
C1 |
H9 |
110.371 |
|
C2 |
C3 |
C4 |
127.414 |
| C2 |
C3 |
H11 |
117.772 |
|
C3 |
C2 |
H10 |
117.868 |
| C3 |
C4 |
C5 |
111.753 |
|
C3 |
C4 |
H12 |
110.778 |
| C3 |
C4 |
H13 |
109.113 |
|
C4 |
C3 |
H11 |
114.806 |
| C4 |
C5 |
C6 |
112.105 |
|
C4 |
C5 |
H14 |
109.266 |
| C4 |
C5 |
H15 |
109.253 |
|
C5 |
C4 |
H12 |
109.179 |
| C5 |
C4 |
H13 |
108.642 |
|
C5 |
C6 |
H16 |
110.675 |
| C5 |
C6 |
H17 |
110.716 |
|
C5 |
C6 |
H18 |
110.727 |
| C6 |
C5 |
H14 |
109.437 |
|
C6 |
C5 |
H15 |
109.424 |
| H7 |
C1 |
H8 |
108.223 |
|
H7 |
C1 |
H9 |
108.170 |
| H8 |
C1 |
H9 |
107.397 |
|
H12 |
C4 |
H13 |
107.248 |
| H14 |
C5 |
H16 |
93.681 |
|
H16 |
C6 |
H17 |
108.241 |
| H16 |
C6 |
H18 |
108.223 |
|
H17 |
C6 |
H18 |
108.163 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
