return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (1-Propanol, 2-methyl-)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-229.542611
Energy at 298.15K-229.553623
Nuclear repulsion energy188.580023
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3880 3449 40.20      
2 A 3553 3158 3.45      
3 A 3551 3157 5.44      
4 A 3551 3156 1.15      
5 A 3547 3153 3.70      
6 A 3472 3087 7.25      
7 A 3436 3054 3.62      
8 A 3371 2996 3.99      
9 A 3369 2995 2.36      
10 A 3342 2970 15.47      
11 A 1744 1550 2.46      
12 A 1741 1548 1.07      
13 A 1737 1544 0.74      
14 A 1734 1541 0.34      
15 A 1709 1519 1.03      
16 A 1639 1457 13.98      
17 A 1628 1447 0.26      
18 A 1619 1439 1.24      
19 A 1571 1397 2.38      
20 A 1537 1367 6.53      
21 A 1471 1307 15.43      
22 A 1391 1237 4.09      
23 A 1326 1178 2.40      
24 A 1275 1133 1.16      
25 A 1243 1105 0.25      
26 A 1194 1062 16.12      
27 A 1085 964 0.41      
28 A 1054 936 2.11      
29 A 1041 926 0.71      
30 A 992 882 3.52      
31 A 887 789 4.14      
32 A 499 443 3.97      
33 A 444 395 9.54      
34 A 372 331 19.63      
35 A 356 316 40.95      
36 A 272 242 0.40      
37 A 236 210 1.35      
38 A 222 197 0.10      
39 A 96 86 2.70      

Unscaled Zero Point Vibrational Energy (zpe) 33591.8 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 29859.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.24119 0.11031 0.08345

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.458 0.002 -0.354
C2 -0.826 -0.669 0.276
C3 1.725 -0.836 0.013
C4 0.624 1.485 0.113
O5 -2.092 -0.100 -0.175
H6 0.337 -0.005 -1.457
H7 1.626 -1.873 -0.351
H8 2.628 -0.394 -0.441
H9 1.868 -0.865 1.107
H10 -0.239 2.101 -0.191
H11 0.720 1.540 1.212
H12 1.530 1.930 -0.333
H13 -0.851 -1.738 -0.028
H14 -0.734 -0.641 1.389
H15 -2.037 0.843 0.200

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.58001.56281.56382.55821.10902.20912.20792.20852.21752.21032.20582.20202.20732.6904
C21.58002.56962.60211.45982.18972.80213.53842.82622.87012.85383.56031.11131.11641.9391
C31.56282.56962.57063.89162.18561.10341.10341.10373.53892.84452.79402.72982.82424.1237
C41.56382.60212.57063.15812.18393.53472.80272.83881.10361.10381.10313.54762.82722.7392
O52.55821.45983.89163.15812.74764.12274.73684.23252.87693.53794.15482.06022.14041.0160
H61.10902.18972.18562.18392.74762.52442.53643.10772.52373.10722.53622.54063.10573.0161
H72.20912.80211.10343.53474.12272.52441.78871.78864.39263.86123.80392.50173.17974.5930
H82.20793.53841.10342.80274.73682.53641.78871.78783.80923.18052.57273.75283.83594.8689
H92.20852.82621.10372.83884.23253.10771.78861.78783.86342.66733.16223.07352.62734.3582
H102.21752.87013.53891.10362.87692.52374.39263.80923.86341.78971.78303.89093.20332.2289
H112.21032.85382.84451.10383.53793.10723.86123.18052.66731.78971.78723.84032.62723.0183
H122.20583.56032.79401.10314.15482.53623.80392.57273.16221.78301.78724.38363.83403.7668
H132.20201.11132.72983.54762.06022.54062.50173.75283.07353.89093.84034.38361.79492.8494
H142.20731.11642.82422.82722.14043.10573.17973.83592.62733.20332.62723.83401.79492.3053
H152.69041.93914.12372.73921.01603.01614.59304.86894.35822.22893.01833.76682.84942.3053

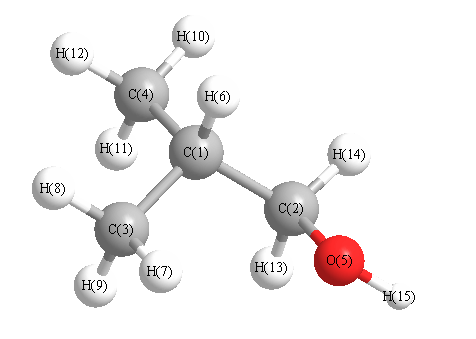
picture of 1-Propanol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O5 114.554 C1 C2 H13 108.557
C1 C2 H14 108.675 C1 C3 H7 110.719
C1 C3 H8 110.629 C1 C3 H9 110.651
C1 C4 H10 111.304 C1 C4 H11 110.720
C1 C4 H12 110.410 C2 C1 C3 109.691
C2 C1 C4 111.729 C2 C1 H6 107.750
C2 O5 H15 101.607 C3 C1 C4 110.605
C3 C1 H6 108.577 C4 C1 H6 108.386
O5 C2 H13 105.706 O5 C2 H14 111.678
H7 C3 H8 108.294 H7 C3 H9 108.263
H8 C3 H9 108.193 H10 C4 H11 108.342
H10 C4 H12 107.801 H11 C4 H12 108.151
H13 C2 H14 107.362
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability