Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -213.294778 |
| Energy at 298.15K | |
| HF Energy | -213.102175 |
| Nuclear repulsion energy | 114.187221 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3619 |
3217 |
0.00 |
27.54 |
0.72 |
0.83 |
| 2 |
A' |
3515 |
3124 |
1.46 |
47.72 |
0.27 |
0.42 |
| 3 |
A' |
3461 |
3077 |
6.93 |
26.50 |
0.15 |
0.26 |
| 4 |
A' |
3321 |
2952 |
10.51 |
46.70 |
0.13 |
0.23 |
| 5 |
A' |
1822 |
1620 |
0.38 |
7.25 |
0.75 |
0.86 |
| 6 |
A' |
1712 |
1522 |
0.58 |
16.48 |
0.74 |
0.85 |
| 7 |
A' |
1584 |
1408 |
0.30 |
6.44 |
0.64 |
0.78 |
| 8 |
A' |
1559 |
1386 |
23.81 |
4.37 |
0.61 |
0.76 |
| 9 |
A' |
1416 |
1258 |
2.04 |
12.23 |
0.61 |
0.76 |
| 10 |
A' |
1280 |
1137 |
5.47 |
2.98 |
0.72 |
0.84 |
| 11 |
A' |
1106 |
983 |
2.70 |
4.75 |
0.72 |
0.84 |
| 12 |
A' |
974 |
865 |
1.96 |
5.18 |
0.27 |
0.43 |
| 13 |
A' |
617 |
548 |
1.36 |
1.31 |
0.75 |
0.86 |
| 14 |
A' |
243 |
216 |
0.63 |
0.55 |
0.51 |
0.67 |
| 15 |
A" |
3419 |
3039 |
10.48 |
38.40 |
0.75 |
0.86 |
| 16 |
A" |
1362 |
1210 |
1.86 |
8.99 |
0.75 |
0.86 |
| 17 |
A" |
1131 |
1005 |
5.67 |
0.05 |
0.75 |
0.86 |
| 18 |
A" |
1117 |
993 |
3.20 |
0.41 |
0.75 |
0.86 |
| 19 |
A" |
999 |
888 |
12.83 |
0.00 |
0.75 |
0.86 |
| 20 |
A" |
585 |
520 |
2.69 |
6.41 |
0.75 |
0.86 |
| 21 |
A" |
125 |
111 |
0.76 |
2.53 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17482.5 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 15540.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
-0.988 |
0.000 |
| C2 |
-0.962 |
0.231 |
0.000 |
| C3 |
-0.532 |
1.507 |
0.000 |
| F4 |
1.346 |
-0.593 |
0.000 |
| H5 |
-1.232 |
2.353 |
0.000 |
| H6 |
0.541 |
1.742 |
0.000 |
| H7 |
-2.037 |
-0.008 |
0.000 |
| H8 |
-0.211 |
-1.622 |
0.898 |
| H9 |
-0.211 |
-1.622 |
-0.898 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5526 | 2.5509 | 1.4030 | 3.5607 | 2.7832 | 2.2601 | 1.1191 | 1.1191 |
C2 | 1.5526 | | 1.3461 | 2.4504 | 2.1391 | 2.1309 | 1.1013 | 2.1912 | 2.1912 | C3 | 2.5509 | 1.3461 | | 2.8172 | 1.0978 | 1.0985 | 2.1349 | 3.2706 | 3.2706 | F4 | 1.4030 | 2.4504 | 2.8172 | | 3.9143 | 2.4699 | 3.4330 | 2.0714 | 2.0714 | H5 | 3.5607 | 2.1391 | 1.0978 | 3.9143 | | 1.8747 | 2.4944 | 4.2006 | 4.2006 | H6 | 2.7832 | 2.1309 | 1.0985 | 2.4699 | 1.8747 | | 3.1154 | 3.5620 | 3.5620 | H7 | 2.2601 | 1.1013 | 2.1349 | 3.4330 | 2.4944 | 3.1154 | | 2.5965 | 2.5965 | H8 | 1.1191 | 2.1912 | 3.2706 | 2.0714 | 4.2006 | 3.5620 | 2.5965 | | 1.7957 | H9 | 1.1191 | 2.1912 | 3.2706 | 2.0714 | 4.2006 | 3.5620 | 2.5965 | 1.7957 | |
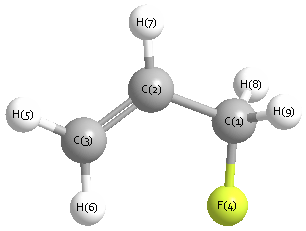 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.133 |
|
C1 |
C2 |
H7 |
115.734 |
| C2 |
C1 |
F4 |
111.904 |
|
C2 |
C1 |
H8 |
109.126 |
| C2 |
C1 |
H9 |
109.126 |
|
C2 |
C3 |
H5 |
121.824 |
| C2 |
C3 |
H6 |
120.972 |
|
C3 |
C2 |
H7 |
121.133 |
| F4 |
C1 |
H8 |
109.920 |
|
F4 |
C1 |
H9 |
109.920 |
| H5 |
C3 |
H6 |
117.204 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -213.294195 |
| Energy at 298.15K | |
| HF Energy | -213.101498 |
| Nuclear repulsion energy | 111.818242 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3613 |
3212 |
0.19 |
28.93 |
0.72 |
0.84 |
| 2 |
A |
3513 |
3123 |
1.70 |
43.56 |
0.26 |
0.42 |
| 3 |
A |
3457 |
3073 |
6.08 |
28.67 |
0.14 |
0.24 |
| 4 |
A |
3434 |
3052 |
9.41 |
29.58 |
0.75 |
0.86 |
| 5 |
A |
3325 |
2956 |
10.55 |
37.12 |
0.15 |
0.26 |
| 6 |
A |
1818 |
1616 |
0.58 |
7.82 |
0.74 |
0.85 |
| 7 |
A |
1705 |
1516 |
0.38 |
12.73 |
0.75 |
0.86 |
| 8 |
A |
1611 |
1432 |
3.10 |
6.83 |
0.50 |
0.67 |
| 9 |
A |
1530 |
1360 |
24.04 |
3.98 |
0.70 |
0.82 |
| 10 |
A |
1419 |
1262 |
3.56 |
9.84 |
0.62 |
0.76 |
| 11 |
A |
1375 |
1223 |
4.07 |
14.14 |
0.75 |
0.86 |
| 12 |
A |
1255 |
1116 |
6.13 |
4.20 |
0.56 |
0.72 |
| 13 |
A |
1246 |
1108 |
2.14 |
3.46 |
0.68 |
0.81 |
| 14 |
A |
1130 |
1004 |
5.84 |
0.12 |
0.72 |
0.84 |
| 15 |
A |
1043 |
927 |
2.64 |
1.21 |
0.39 |
0.56 |
| 16 |
A |
1004 |
893 |
11.53 |
0.03 |
0.29 |
0.44 |
| 17 |
A |
989 |
879 |
2.68 |
2.73 |
0.26 |
0.41 |
| 18 |
A |
685 |
609 |
3.31 |
3.91 |
0.65 |
0.79 |
| 19 |
A |
430 |
383 |
0.19 |
2.15 |
0.54 |
0.70 |
| 20 |
A |
346 |
308 |
1.75 |
3.00 |
0.75 |
0.86 |
| 21 |
A |
87 |
77 |
0.37 |
2.78 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17508.5 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 15563.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.640 |
0.413 |
0.340 |
| C2 |
0.681 |
-0.396 |
0.255 |
| C3 |
1.833 |
0.109 |
-0.229 |
| F4 |
-1.681 |
-0.216 |
-0.367 |
| H5 |
2.755 |
-0.486 |
-0.265 |
| H6 |
1.897 |
1.142 |
-0.599 |
| H7 |
0.638 |
-1.435 |
0.619 |
| H8 |
-0.938 |
0.524 |
1.413 |
| H9 |
-0.467 |
1.440 |
-0.066 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5513 | 2.5551 | 1.4069 | 3.5636 | 2.8012 | 2.2646 | 1.1192 | 1.1172 |
C2 | 1.5513 | | 1.3477 | 2.4486 | 2.1403 | 2.1388 | 1.1017 | 2.1930 | 2.1891 | C3 | 2.5551 | 1.3477 | | 3.5310 | 1.0984 | 1.0990 | 2.1286 | 3.2472 | 2.6619 | F4 | 1.4069 | 2.4486 | 3.5310 | | 4.4450 | 3.8333 | 2.7997 | 2.0658 | 2.0740 | H5 | 3.5636 | 2.1403 | 1.0984 | 4.4450 | | 1.8706 | 2.4826 | 4.1804 | 3.7591 | H6 | 2.8012 | 2.1388 | 1.0990 | 3.8333 | 1.8706 | | 3.1161 | 3.5311 | 2.4415 | H7 | 2.2646 | 1.1017 | 2.1286 | 2.7997 | 2.4826 | 3.1161 | | 2.6367 | 3.1554 | H8 | 1.1192 | 2.1930 | 3.2472 | 2.0658 | 4.1804 | 3.5311 | 2.6367 | | 1.8028 | H9 | 1.1172 | 2.1891 | 2.6619 | 2.0740 | 3.7591 | 2.4415 | 3.1554 | 1.8028 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.461 |
|
C1 |
C2 |
H7 |
116.178 |
| C2 |
C1 |
F4 |
111.637 |
|
C2 |
C1 |
H8 |
109.340 |
| C2 |
C1 |
H9 |
109.154 |
|
C2 |
C3 |
H5 |
121.746 |
| C2 |
C3 |
H6 |
121.558 |
|
C3 |
C2 |
H7 |
120.361 |
| F4 |
C1 |
H8 |
109.196 |
|
F4 |
C1 |
H9 |
109.976 |
| H5 |
C3 |
H6 |
116.695 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
