return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6 (Hexa-1,5-diene-3-yne)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-228.140251
Energy at 298.15K-228.143771
HF Energy-227.743692
Nuclear repulsion energy171.727493
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3617 3215 0.00      
2 Ag 3498 3110 0.00      
3 Ag 3458 3074 0.00      
4 Ag 2382 2117 0.00      
5 Ag 1765 1569 0.00      
6 Ag 1570 1396 0.00      
7 Ag 1422 1264 0.00      
8 Ag 1162 1033 0.00      
9 Ag 739 657 0.00      
10 Ag 546 485 0.00      
11 Ag 227 202 0.00      
12 Au 1115 991 12.66      
13 Au 986 876 23.15      
14 Au 723 643 3.62      
15 Au 184 163 1.73      
16 Au 33 29 0.05      
17 Bg 1113 990 0.00      
18 Bg 985 876 0.00      
19 Bg 738 656 0.00      
20 Bg 365 324 0.00      
21 Bu 3617 3215 0.02      
22 Bu 3499 3110 6.01      
23 Bu 3458 3074 31.21      
24 Bu 1787 1589 10.54      
25 Bu 1587 1411 20.79      
26 Bu 1418 1261 13.51      
27 Bu 1267 1127 16.50      
28 Bu 1072 953 1.86      
29 Bu 542 481 0.99      
30 Bu 111 99 0.79      

Unscaled Zero Point Vibrational Energy (zpe) 22492.0 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 19993.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.84112 0.04362 0.04147

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.615 0.000
C2 0.000 -0.615 0.000
C3 0.000 2.083 0.000
C4 0.000 -2.083 0.000
C5 1.126 2.838 0.000
C6 -1.126 -2.838 0.000
H7 -0.987 2.576 0.000
H8 0.987 -2.576 0.000
H9 2.128 2.385 0.000
H10 1.072 3.936 0.000
H11 -2.128 -2.385 0.000
H12 -1.072 -3.936 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
C11.22911.46882.69802.49253.63172.19613.34012.76823.49003.67784.6749
C21.22912.69801.46883.63172.49253.34012.19613.67784.67492.76823.4900
C31.46882.69804.16681.35595.04871.10364.76312.14912.14034.94936.1139
C42.69801.46884.16685.04871.35594.76311.10364.94936.11392.14912.1403
C52.49253.63171.35595.04876.10682.13005.41601.09901.09906.15407.1217
C63.63172.49255.04871.35596.10685.41602.13006.15407.12171.09901.0990
H72.19613.34011.10364.76312.13005.41605.51793.12112.46805.09086.5125
H83.34012.19614.76311.10365.41602.13005.51795.09086.51253.12112.4680
H92.76823.67782.14914.94931.09906.15403.12115.09081.87576.39277.0848
H103.49004.67492.14036.11391.09907.12172.46806.51251.87577.08488.1584
H113.67782.76824.94932.14916.15401.09905.09083.12116.39277.08481.8757
H124.67493.49006.11392.14037.12171.09906.51252.46807.08488.15841.8757

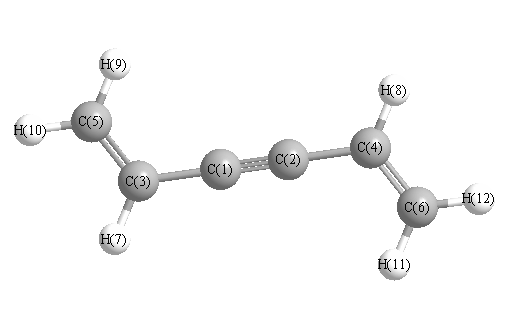
picture of Hexa-1,5-diene-3-yne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 180.000 C1 C3 C5 123.818
C1 C3 H7 116.524 C2 C1 C3 180.000
C2 C4 C6 123.818 C2 C4 H8 116.524
C3 C5 H9 121.848 C3 C5 H10 120.997
C4 C6 H11 121.848 C4 C6 H12 120.997
C5 C3 H7 119.658 C6 C4 H8 119.658
H9 C5 H10 117.154 H11 C6 H12 117.154
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability