return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O2 (propenalol)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-262.441655
Energy at 298.15K-262.446748
HF Energy-262.145198
Nuclear repulsion energy163.121702
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3561 3165 8.18      
2 A' 3476 3090 1.00      
3 A' 3365 2991 20.46      
4 A' 3037 2699 117.72      
5 A' 1925 1711 140.62      
6 A' 1755 1560 10.38      
7 A' 1556 1383 25.77      
8 A' 1503 1336 124.97      
9 A' 1469 1306 5.56      
10 A' 1339 1190 57.48      
11 A' 1170 1040 8.07      
12 A' 1052 935 52.77      
13 A' 916 814 0.25      
14 A' 580 516 8.04      
15 A' 364 323 2.37      
16 A" 1212 1078 86.86      
17 A" 1033 918 3.68      
18 A" 960 853 11.55      
19 A" 795 707 7.61      
20 A" 427 379 8.46      
21 A" 340 302 0.49      

Unscaled Zero Point Vibrational Energy (zpe) 15917.1 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 14148.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.30503 0.17906 0.11283

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.156 0.000
C2 1.246 0.361 0.000
C3 -1.192 0.450 0.000
O4 1.221 -0.932 0.000
O5 -1.245 -0.913 0.000
H6 -0.212 -1.149 0.000
H7 0.030 2.249 0.000
H8 2.227 0.898 0.000
H9 -2.178 0.955 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9
C11.47741.38552.41842.41452.31451.09372.24172.1875
C21.47742.43931.29322.79752.09872.24541.11823.4748
C31.38552.43932.78061.36381.87542.17513.44811.1078
O42.41841.29322.78062.46591.44923.39652.08753.8876
O52.41452.79751.36382.46591.05973.40943.91542.0879
H62.31452.09871.87541.44921.05973.40673.18352.8796
H71.09372.24542.17513.39653.40943.40672.57932.5596
H82.24171.11823.44812.08753.91543.18352.57934.4054
H92.18753.47481.10783.88762.08792.87962.55964.4054

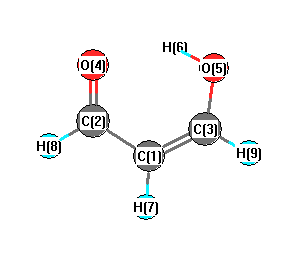
picture of propenalol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O4 121.447 C1 C2 H8 118.807
C1 C3 O5 122.851 C1 C3 H9 122.252
C2 C1 C3 116.830 C2 C1 H7 120.966
C3 C1 H7 122.204 C3 O5 H6 100.650
O4 C2 H8 119.746 O5 C3 H9 114.897
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/STO-3G Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.170      
2 C 0.139      
3 C 0.118      
4 O -0.265      
5 O -0.304      
6 H 0.272      
7 H 0.063      
8 H 0.068      
9 H 0.080      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 100.439
(<r2>)1/2 10.022