return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C10H8 (naphthalene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1Ag
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-379.278749
Energy at 298.15K-379.286692
HF Energy-378.673572
Nuclear repulsion energy450.158947
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3524 3133 0.00      
2 Ag 3501 3112 0.00      
3 Ag 1712 1522 0.00      
4 Ag 1595 1418 0.00      
5 Ag 1461 1299 0.00      
6 Ag 1255 1116 0.00      
7 Ag 1093 971 0.00      
8 Ag 805 715 0.00      
9 Ag 532 473 0.00      
10 Au 1070 951 0.00      
11 Au 921 819 0.00      
12 Au 672 598 0.00      
13 Au 201 179 0.00      
14 B1g 1046 930 0.00      
15 B1g 775 689 0.00      
16 B1g 408 363 0.00      
17 B1u 3515 3125 0.95      
18 B1u 3500 3111 0.92      
19 B1u 1738 1545 5.89      
20 B1u 1516 1347 4.33      
21 B1u 1355 1205 5.87      
22 B1u 1214 1079 1.08      
23 B1u 847 753 0.69      
24 B1u 371 329 0.20      
25 B2g 1072 953 0.00      
26 B2g 944 839 0.00      
27 B2g 830 738 0.00      
28 B2g 496 441 0.00      
29 B2u 3525 3133 4.56      
30 B2u 3503 3114 8.16      
31 B2u 1651 1468 3.21      
32 B2u 1450 1289 1.43      
33 B2u 1286 1143 0.02      
34 B2u 1209 1075 0.02      
35 B2u 1064 946 1.99      
36 B2u 656 583 3.98      
37 B3g 3516 3125 0.00      
38 B3g 3498 3110 0.00      
39 B3g 1778 1581 0.00      
40 B3g 1591 1414 0.00      
41 B3g 1351 1201 0.00      
42 B3g 1247 1108 0.00      
43 B3g 1002 890 0.00      
44 B3g 537 477 0.00      
45 B3u 1055 937 0.52      
46 B3u 850 755 30.32      
47 B3u 508 451 4.33      
48 B3u 176 156 0.68      

Unscaled Zero Point Vibrational Energy (zpe) 34710.1 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 30853.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.10084 0.03973 0.02850

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.275 1.428
C2 0.000 2.480 0.725
C3 0.000 2.480 -0.725
C4 0.000 1.275 -1.428
C5 0.000 -1.275 -1.428
C6 0.000 -2.480 -0.725
C7 0.000 -2.480 0.725
C8 0.000 -1.275 1.428
C9 0.000 0.000 0.723
C10 0.000 0.000 -0.723
H11 0.000 1.271 2.533
H12 0.000 3.443 1.267
H13 0.000 3.443 -1.267
H14 0.000 1.271 -2.533
H15 0.000 -1.271 -2.533
H16 0.000 -3.443 -1.267
H17 0.000 -3.443 1.267
H18 0.000 -1.271 2.533

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 C10 H11 H12 H13 H14 H15 H16 H17 H18
C11.39522.46732.85633.82944.32893.82082.55061.45712.50101.10492.17323.45833.96134.70895.43334.72062.7753
C21.39521.44972.46734.32895.16804.96053.82082.48032.87212.17561.10442.21193.47534.96826.24875.94754.1640
C32.46731.44971.39523.82084.96055.16804.32892.87212.48033.47532.21191.10442.17564.16405.94756.24874.9682
C42.85632.46731.39522.55063.82084.32893.82942.50101.45713.96133.45832.17321.10492.77534.72065.43334.7089
C53.82944.32893.82082.55061.39522.46732.85632.50101.45714.70895.43334.72062.77531.10492.17323.45833.9613
C64.32895.16804.96053.82081.39521.44972.46732.87212.48034.96826.24875.94754.16402.17561.10442.21193.4753
C73.82084.96055.16804.32892.46731.44971.39522.48032.87214.16405.94756.24874.96823.47532.21191.10442.1756
C82.55063.82084.32893.82942.85632.46731.39521.45712.50102.77534.72065.43334.70893.96133.45832.17321.1049
C91.45712.48032.87212.50102.50102.87212.48031.45711.44662.21133.48523.97643.49553.49553.97643.48522.2113
C102.50102.87212.48031.45711.45712.48032.87212.50101.44663.49553.97643.48522.21132.21133.48523.97643.4955
H111.10492.17563.47533.96134.70894.96824.16402.77532.21133.49552.51414.37685.06625.66796.05424.88032.5413
H122.17321.10442.21193.45835.43336.24875.94754.72063.48523.97642.51412.53374.37686.05427.33656.88514.8803
H133.45832.21191.10442.17324.72065.94756.24875.43333.97643.48524.37682.53372.51414.88036.88517.33656.0542
H143.96133.47532.17561.10492.77534.16404.96824.70893.49552.21135.06624.37682.51412.54134.88036.05425.6679
H154.70894.96824.16402.77531.10492.17563.47533.96133.49552.21135.66796.05424.88032.54132.51414.37685.0662
H165.43336.24875.94754.72062.17321.10442.21193.45833.97643.48526.05427.33656.88514.88032.51412.53374.3768
H174.72065.94756.24875.43333.45832.21191.10442.17323.48523.97644.88036.88517.33656.05424.37682.53372.5141
H182.77534.16404.96824.70893.96133.47532.17561.10492.21133.49552.54134.88036.05425.66795.06624.37682.5141

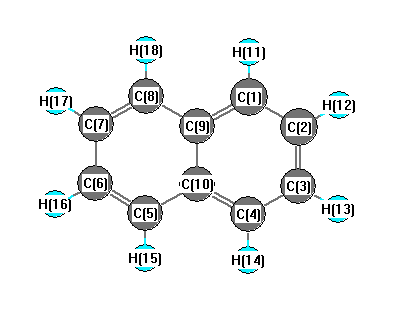
picture of naphthalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.272 C1 C2 H12 120.337
C1 C9 C8 122.139 C1 C9 C10 118.930
C2 C1 C9 120.798 C2 C1 H11 120.513
C2 C3 C4 120.272 C2 C3 H13 119.391
C3 C2 H12 119.391 C3 C4 C10 120.798
C3 C4 H14 120.513 C4 C3 H13 120.337
C4 C10 C5 122.139 C4 C10 C9 118.930
C5 C6 C7 120.272 C5 C6 H16 120.337
C5 C10 C9 118.930 C6 C5 C10 120.798
C6 C5 H15 120.513 C6 C7 C8 120.272
C6 C7 H17 119.391 C7 C6 H16 119.391
C7 C8 C9 120.798 C7 C8 H18 120.513
C8 C7 H17 120.337 C8 C9 C10 118.930
C9 C1 H11 118.690 C9 C8 H18 118.690
C10 C4 H14 118.690 C10 C5 H15 118.690
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability