return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (3-Hexyne)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-230.555146
Energy at 298.15K-230.564014
HF Energy-230.189804
Nuclear repulsion energy203.285669
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3562 3166 0.00      
2 A 3557 3162 0.78      
3 A 3498 3109 0.00      
4 A 3392 3015 3.09      
5 A 3378 3002 3.16      
6 A 2435 2165 0.08      
7 A 1736 1543 0.73      
8 A 1729 1537 0.00      
9 A 1701 1512 0.20      
10 A 1616 1436 0.56      
11 A 1521 1352 1.01      
12 A 1430 1271 0.00      
13 A 1233 1096 0.00      
14 A 1213 1078 0.77      
15 A 1103 980 0.01      
16 A 873 776 0.00      
17 A 734 653 0.00      
18 A 438 389 1.29      
19 A 404 359 0.00      
20 A 221 197 0.00      
21 A 93 83 0.62      
22 A 9 8 0.00      
23 B 3562 3166 4.47      
24 B 3557 3162 4.65      
25 B 3497 3109 2.10      
26 B 3392 3016 5.23      
27 B 3378 3002 1.43      
28 B 1736 1543 0.00      
29 B 1729 1537 2.46      
30 B 1700 1511 3.40      
31 B 1615 1436 0.09      
32 B 1520 1351 65.42      
33 B 1432 1273 0.07      
34 B 1263 1123 8.79      
35 B 1242 1104 0.23      
36 B 1202 1068 0.67      
37 B 1025 911 0.07      
38 B 872 775 4.84      
39 B 577 513 0.52      
40 B 278 247 1.17      
41 B 227 202 0.07      
42 B 157 140 1.34      

Unscaled Zero Point Vibrational Energy (zpe) 34917.8 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 31038.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.30310 0.04497 0.04037

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.000 0.612 -0.576
C2 0.000 -0.612 -0.576
C3 0.000 2.117 -0.564
C4 -0.000 -2.117 -0.564
C5 0.002 2.701 0.888
C6 -0.002 -2.701 0.888
H7 0.891 2.490 -1.110
H8 -0.892 2.490 -1.107
H9 -0.891 -2.490 -1.110
H10 0.892 -2.490 -1.107
H11 0.002 3.804 0.852
H12 -0.892 2.369 1.442
H13 0.898 2.369 1.439
H14 -0.002 -3.804 0.852
H15 0.892 -2.369 1.442
H16 -0.898 -2.369 1.439

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.22311.50562.72872.55103.62142.14612.14613.27043.27043.49752.82022.82044.64093.70783.7079
C21.22312.72871.50563.62142.55103.27043.27042.14612.14614.64093.70783.70793.49752.82022.8204
C31.50562.72874.23421.56505.03221.10891.10894.72364.72362.20302.20992.20996.08844.99414.9943
C42.72871.50564.23425.03221.56504.72364.72361.10891.10896.08844.99414.99432.20302.20992.2099
C52.55103.62141.56505.03225.40202.19642.19645.63295.63151.10371.10311.10316.50535.17685.1785
C63.62142.55105.03221.56505.40205.63295.63152.19642.19646.50535.17685.17851.10371.10311.1031
H72.14613.27041.10894.72362.19645.63291.78315.28824.97912.52363.11512.55206.65275.48755.7706
H82.14613.27041.10894.72362.19645.63151.78314.97915.28932.52352.55213.11516.65165.76915.4852
H93.27042.14614.72361.10895.63292.19645.28824.97911.78316.65275.48755.77062.52363.11512.5520
H103.27042.14614.72361.10895.63152.19644.97915.28931.78316.65165.76915.48522.52352.55213.1151
H113.49754.64092.20306.08841.10376.50532.52362.52356.65276.65161.79091.79097.60836.26436.2656
H122.82023.70782.20994.99411.10315.17683.11512.55215.48755.76911.79091.78956.26435.06164.7373
H132.82043.70792.20994.99431.10315.17852.55203.11515.77065.48521.79091.78956.26564.73735.0663
H144.64093.49756.08842.20306.50531.10376.65276.65162.52362.52357.60836.26436.26561.79091.7909
H153.70782.82024.99412.20995.17681.10315.48755.76913.11512.55216.26435.06164.73731.79091.7895
H163.70792.82044.99432.20995.17851.10315.77065.48522.55203.11516.26564.73735.06631.79091.7895

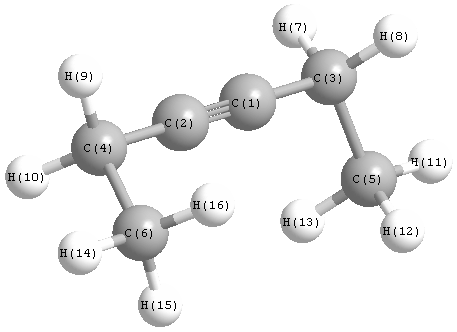
picture of 3-Hexyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 179.562 C1 C3 C5 112.345
C1 C3 H7 109.397 C1 C3 H8 109.397
C2 C1 C3 179.562 C2 C4 C6 112.345
C2 C4 H9 109.397 C2 C4 H10 109.397
C3 C5 H11 110.080 C3 C5 H12 110.653
C3 C5 H13 110.653 C4 C6 H14 110.080
C4 C6 H15 110.653 C4 C6 H16 110.653
C5 C3 H7 109.268 C5 C3 H8 109.269
C6 C4 H9 109.268 C6 C4 H10 109.269
H7 C3 H8 107.020 H9 C4 H10 107.020
H11 C5 H12 108.491 H11 C5 H13 108.491
H12 C5 H13 108.405 H14 C6 H15 108.491
H14 C6 H16 108.491 H15 C6 H15 0.000
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability