return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C9H8 (Indene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-341.801383
Energy at 298.15K-341.809511
HF Energy-341.262991
Nuclear repulsion energy387.893200
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3547 3153 0.02      
2 A' 3524 3133 1.19      
3 A' 3520 3129 7.38      
4 A' 3515 3124 2.44      
5 A' 3508 3118 0.34      
6 A' 3502 3113 0.03      
7 A' 3381 3005 0.18      
8 A' 1759 1564 0.81      
9 A' 1734 1542 0.07      
10 A' 1706 1516 0.24      
11 A' 1673 1487 0.69      
12 A' 1604 1426 14.67      
13 A' 1599 1422 7.41      
14 A' 1497 1331 2.21      
15 A' 1446 1285 0.53      
16 A' 1360 1209 0.26      
17 A' 1345 1196 1.66      
18 A' 1296 1152 0.72      
19 A' 1270 1129 0.63      
20 A' 1228 1091 0.78      
21 A' 1207 1073 0.32      
22 A' 1159 1030 1.02      
23 A' 1094 973 0.57      
24 A' 1045 929 0.56      
25 A' 917 815 0.94      
26 A' 889 790 0.95      
27 A' 771 685 0.93      
28 A' 629 559 0.36      
29 A' 562 500 0.29      
30 A' 399 354 0.13      
31 A" 3485 3098 0.58      
32 A" 1269 1128 0.43      
33 A" 1069 950 0.00      
34 A" 1056 939 0.30      
35 A" 1022 909 0.80      
36 A" 1020 907 0.09      
37 A" 950 844 0.17      
38 A" 835 742 19.74      
39 A" 783 696 1.32      
40 A" 752 668 4.60      
41 A" 588 523 1.29      
42 A" 449 399 2.68      
43 A" 412 366 1.67      
44 A" 223 198 1.21      
45 A" 202 179 0.12      

Unscaled Zero Point Vibrational Energy (zpe) 33399.6 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 29688.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.12205 0.05090 0.03617

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.320 -0.003 0.000
C2 -1.895 -1.356 0.000
C3 -0.515 -1.684 0.000
C4 -1.374 1.056 0.000
C5 0.000 0.734 0.000
C6 0.429 -0.629 0.000
C7 1.933 -0.645 0.000
C8 2.402 0.633 0.000
C9 1.242 1.655 0.000
H10 -3.397 0.228 0.000
H11 -2.647 -2.161 0.000
H12 -0.192 -2.736 0.000
H13 -1.714 2.103 0.000
H14 2.541 -1.561 0.000
H15 3.459 0.938 0.000
H16 1.273 2.312 0.894
H17 1.273 2.312 -0.894

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17
C11.41842.46631.42022.43472.81994.30154.76493.92921.10102.18243.46452.19185.10445.85504.36644.3664
C21.41841.41822.46792.82192.43543.89404.73554.34852.18291.10102.19253.46444.44065.82464.92834.9283
C32.46631.41822.87122.47261.41572.66003.72563.77303.45812.18411.10123.97253.05854.76084.46774.4677
C41.42022.46792.87121.41112.46813.71863.79952.68362.18563.45943.97241.10134.70864.83403.06263.0626
C52.43472.82192.47261.41111.42972.37442.40401.54593.43433.92283.47612.19393.42383.46452.21492.2149
C62.81992.43541.41572.46811.42971.50402.34232.42463.92083.43612.19663.47312.30773.41083.18753.1875
C74.30153.89402.66003.71862.37441.50401.36112.40105.40084.82442.98174.56671.09942.19783.15823.1582
C84.76494.73553.72563.79952.40402.34231.36111.54575.81285.77044.25234.37092.19851.09962.21162.2116
C93.92924.34853.77302.68361.54592.42462.40101.54574.85315.44824.61942.99013.46802.32981.10951.1095
H101.10102.18293.45812.18563.43433.92085.40085.81284.85312.50404.36602.51926.20106.89195.19055.1905
H112.18241.10102.18413.45943.92283.43614.82445.77045.44822.50402.52184.36495.22206.84676.01376.0137
H123.46452.19251.10123.97243.47612.19662.98174.25234.61944.36602.52185.07372.97445.17915.33165.3316
H132.19183.46443.97251.10132.19393.47314.56674.37092.99012.51924.36495.07375.61525.30273.12473.1247
H145.10444.44063.05854.70863.42382.30771.09942.19853.46806.20105.22202.97445.61522.66184.17174.1717
H155.85505.82464.76084.83403.46453.41082.19781.09962.32986.89196.84675.17915.30272.66182.73232.7323
H164.36644.92834.46773.06262.21493.18753.15822.21161.10955.19056.01375.33163.12474.17172.73231.7873
H174.36644.92834.46773.06262.21493.18753.15822.21161.10955.19056.01375.33163.12474.17172.73231.7873

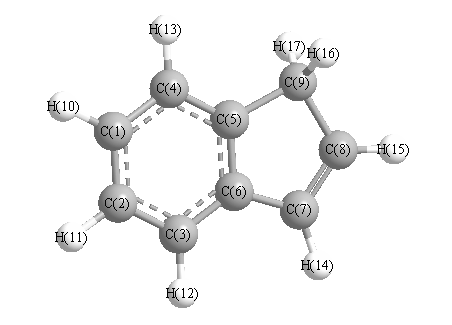
picture of Indene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.791 C1 C2 H11 119.513
C1 C4 C5 118.614 C1 C4 H13 120.210
C2 C1 C4 120.778 C2 C1 H10 119.564
C2 C3 C6 118.499 C2 C3 H12 120.454
C3 C2 H11 119.696 C3 C6 C5 120.680
C3 C6 C7 131.280 C4 C1 H10 119.658
C4 C5 C6 120.638 C4 C5 C9 130.281
C5 C4 H13 121.176 C5 C6 C7 108.040
C5 C9 C8 102.082 C5 C9 H16 112.005
C5 C9 H17 112.005 C6 C3 H12 121.047
C6 C5 C9 109.081 C6 C7 C8 109.575
C6 C7 H14 124.117 C7 C8 C9 111.221
C7 C8 H15 126.218 C8 C7 H14 126.307
C8 C9 H16 111.749 C8 C9 H17 111.749
C9 C8 H15 122.561 H16 C9 H17 107.313
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability