Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -336.403154 |
| Energy at 298.15K | |
| HF Energy | -336.089441 |
| Nuclear repulsion energy | 236.567588 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3369 |
2995 |
8.01 |
|
|
|
| 2 |
A1 |
1892 |
1682 |
150.87 |
|
|
|
| 3 |
A1 |
1719 |
1528 |
5.10 |
|
|
|
| 4 |
A1 |
1481 |
1317 |
3.00 |
|
|
|
| 5 |
A1 |
1253 |
1114 |
69.06 |
|
|
|
| 6 |
A1 |
1053 |
936 |
10.76 |
|
|
|
| 7 |
A1 |
838 |
745 |
24.73 |
|
|
|
| 8 |
A1 |
702 |
624 |
0.11 |
|
|
|
| 9 |
A2 |
3473 |
3087 |
0.00 |
|
|
|
| 10 |
A2 |
1356 |
1205 |
0.00 |
|
|
|
| 11 |
A2 |
1217 |
1082 |
0.00 |
|
|
|
| 12 |
A2 |
90i |
80i |
0.00 |
|
|
|
| 13 |
B1 |
3480 |
3093 |
7.17 |
|
|
|
| 14 |
B1 |
1314 |
1168 |
4.76 |
|
|
|
| 15 |
B1 |
908 |
807 |
1.08 |
|
|
|
| 16 |
B1 |
611 |
543 |
15.04 |
|
|
|
| 17 |
B1 |
102 |
91 |
2.22 |
|
|
|
| 18 |
B2 |
3366 |
2992 |
3.07 |
|
|
|
| 19 |
B2 |
1711 |
1521 |
0.14 |
|
|
|
| 20 |
B2 |
1513 |
1345 |
5.90 |
|
|
|
| 21 |
B2 |
1289 |
1146 |
73.09 |
|
|
|
| 22 |
B2 |
1121 |
996 |
108.56 |
|
|
|
| 23 |
B2 |
788 |
700 |
0.02 |
|
|
|
| 24 |
B2 |
475 |
422 |
1.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17469.9 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 15529.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.873 |
| O2 |
0.000 |
0.000 |
2.117 |
| O3 |
0.000 |
1.205 |
0.066 |
| O4 |
0.000 |
-1.205 |
0.066 |
| C5 |
0.000 |
0.790 |
-1.324 |
| C6 |
0.000 |
-0.790 |
-1.324 |
| H7 |
-0.902 |
1.194 |
-1.835 |
| H8 |
0.902 |
1.194 |
-1.835 |
| H9 |
0.902 |
-1.194 |
-1.835 |
| H10 |
-0.902 |
-1.194 |
-1.835 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.2441 | 1.4506 | 1.4506 | 2.3349 | 2.3349 | 3.0946 | 3.0946 | 3.0946 | 3.0946 |
O2 | 1.2441 | | 2.3793 | 2.3793 | 3.5308 | 3.5308 | 4.2265 | 4.2265 | 4.2265 | 4.2265 | O3 | 1.4506 | 2.3793 | | 2.4103 | 1.4506 | 2.4314 | 2.1044 | 2.1044 | 3.1914 | 3.1914 | O4 | 1.4506 | 2.3793 | 2.4103 | | 2.4314 | 1.4506 | 3.1914 | 3.1914 | 2.1044 | 2.1044 | C5 | 2.3349 | 3.5308 | 1.4506 | 2.4314 | | 1.5796 | 1.1130 | 1.1130 | 2.2385 | 2.2385 | C6 | 2.3349 | 3.5308 | 2.4314 | 1.4506 | 1.5796 | | 2.2385 | 2.2385 | 1.1130 | 1.1130 | H7 | 3.0946 | 4.2265 | 2.1044 | 3.1914 | 1.1130 | 2.2385 | | 1.8043 | 2.9931 | 2.3881 | H8 | 3.0946 | 4.2265 | 2.1044 | 3.1914 | 1.1130 | 2.2385 | 1.8043 | | 2.3881 | 2.9931 | H9 | 3.0946 | 4.2265 | 3.1914 | 2.1044 | 2.2385 | 1.1130 | 2.9931 | 2.3881 | | 1.8043 | H10 | 3.0946 | 4.2265 | 3.1914 | 2.1044 | 2.2385 | 1.1130 | 2.3881 | 2.9931 | 1.8043 | |
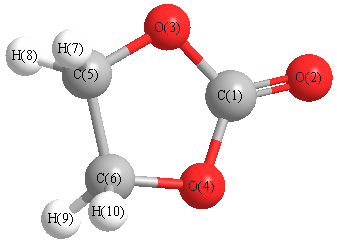 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
107.180 |
|
C1 |
O4 |
C6 |
107.180 |
| O2 |
C1 |
O3 |
123.819 |
|
O2 |
C1 |
O4 |
123.819 |
| O3 |
C1 |
O4 |
112.361 |
|
O3 |
C5 |
C6 |
106.639 |
| O3 |
C5 |
H7 |
109.645 |
|
O3 |
C5 |
H8 |
109.645 |
| O4 |
C6 |
C5 |
106.639 |
|
O4 |
C6 |
H9 |
109.645 |
| O4 |
C6 |
H10 |
109.645 |
|
C5 |
C6 |
H9 |
111.299 |
| C5 |
C6 |
H10 |
111.299 |
|
C6 |
C5 |
H7 |
111.299 |
| C6 |
C5 |
H8 |
111.299 |
|
H7 |
C5 |
H8 |
108.299 |
| H9 |
C6 |
H10 |
108.299 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -336.403166 |
| Energy at 298.15K | |
| HF Energy | -336.089435 |
| Nuclear repulsion energy | 236.629348 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3475 |
3089 |
0.01 |
|
|
|
| 2 |
A |
3369 |
2994 |
7.69 |
|
|
|
| 3 |
A |
1892 |
1682 |
149.68 |
|
|
|
| 4 |
A |
1718 |
1527 |
5.16 |
|
|
|
| 5 |
A |
1484 |
1319 |
3.77 |
|
|
|
| 6 |
A |
1354 |
1204 |
1.25 |
|
|
|
| 7 |
A |
1253 |
1114 |
61.78 |
|
|
|
| 8 |
A |
1215 |
1080 |
6.18 |
|
|
|
| 9 |
A |
1052 |
935 |
9.76 |
|
|
|
| 10 |
A |
838 |
745 |
24.77 |
|
|
|
| 11 |
A |
701 |
624 |
0.10 |
|
|
|
| 12 |
A |
53i |
47i |
0.05 |
|
|
|
| 13 |
B |
3481 |
3094 |
6.89 |
|
|
|
| 14 |
B |
3366 |
2992 |
3.68 |
|
|
|
| 15 |
B |
1710 |
1520 |
0.13 |
|
|
|
| 16 |
B |
1511 |
1343 |
5.92 |
|
|
|
| 17 |
B |
1316 |
1170 |
8.55 |
|
|
|
| 18 |
B |
1284 |
1141 |
68.87 |
|
|
|
| 19 |
B |
1120 |
995 |
107.39 |
|
|
|
| 20 |
B |
918 |
816 |
0.93 |
|
|
|
| 21 |
B |
775 |
689 |
0.07 |
|
|
|
| 22 |
B |
612 |
544 |
15.33 |
|
|
|
| 23 |
B |
475 |
422 |
1.24 |
|
|
|
| 24 |
B |
105 |
94 |
2.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17484.8 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 15542.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.872 |
| O2 |
0.000 |
0.000 |
2.117 |
| O3 |
0.000 |
1.205 |
0.064 |
| O4 |
0.000 |
-1.205 |
0.064 |
| C5 |
0.077 |
0.786 |
-1.323 |
| C6 |
-0.077 |
-0.786 |
-1.323 |
| H7 |
-0.735 |
1.276 |
-1.903 |
| H8 |
1.055 |
1.093 |
-1.758 |
| H9 |
0.735 |
-1.276 |
-1.903 |
| H10 |
-1.055 |
-1.093 |
-1.758 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.2441 | 1.4511 | 1.4511 | 2.3327 | 2.3327 | 3.1421 | 3.0379 | 3.1421 | 3.0379 |
O2 | 1.2441 | | 2.3799 | 2.3799 | 3.5286 | 3.5286 | 4.2811 | 4.1620 | 4.2811 | 4.1620 | O3 | 1.4511 | 2.3799 | | 2.4104 | 1.4511 | 2.4274 | 2.1017 | 2.1089 | 3.2506 | 3.1171 | O4 | 1.4511 | 2.3799 | 2.4104 | | 2.4274 | 1.4511 | 3.2506 | 3.1171 | 2.1017 | 2.1089 | C5 | 2.3327 | 3.5286 | 1.4511 | 2.4274 | | 1.5785 | 1.1122 | 1.1138 | 2.2403 | 2.2359 | C6 | 2.3327 | 3.5286 | 2.4274 | 1.4511 | 1.5785 | | 2.2403 | 2.2359 | 1.1122 | 1.1138 | H7 | 3.1421 | 4.2811 | 2.1017 | 3.2506 | 1.1122 | 2.2403 | | 1.8049 | 2.9446 | 2.3944 | H8 | 3.0379 | 4.1620 | 2.1089 | 3.1171 | 1.1138 | 2.2359 | 1.8049 | | 2.3944 | 3.0377 | H9 | 3.1421 | 4.2811 | 3.2506 | 2.1017 | 2.2403 | 1.1122 | 2.9446 | 2.3944 | | 1.8049 | H10 | 3.0379 | 4.1620 | 3.1171 | 2.1089 | 2.2359 | 1.1138 | 2.3944 | 3.0377 | 1.8049 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
106.984 |
|
C1 |
O4 |
C6 |
106.984 |
| O2 |
C1 |
O3 |
123.844 |
|
O2 |
C1 |
O4 |
123.844 |
| O3 |
C1 |
O4 |
112.312 |
|
O3 |
C5 |
C6 |
106.420 |
| O3 |
C5 |
H7 |
109.449 |
|
O3 |
C5 |
H8 |
109.920 |
| O4 |
C6 |
C5 |
106.420 |
|
O4 |
C6 |
H9 |
109.449 |
| O4 |
C6 |
H10 |
109.920 |
|
C5 |
C6 |
H9 |
111.556 |
| C5 |
C6 |
H10 |
111.118 |
|
C6 |
C5 |
H7 |
111.556 |
| C6 |
C5 |
H8 |
111.118 |
|
H7 |
C5 |
H8 |
108.360 |
| H9 |
C6 |
H10 |
108.360 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
